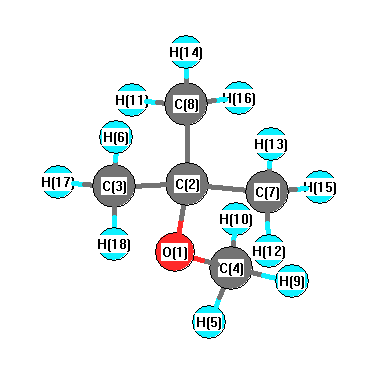
Vibrational Frequencies calculated at LSDA/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3091 |
3040 |
18.75 |
|
|
|
| 2 |
A |
3079 |
3029 |
6.10 |
|
|
|
| 3 |
A |
3067 |
3016 |
27.64 |
|
|
|
| 4 |
A |
3056 |
3005 |
18.48 |
|
|
|
| 5 |
A |
2995 |
2946 |
4.24 |
|
|
|
| 6 |
A |
2986 |
2937 |
10.66 |
|
|
|
| 7 |
A |
2911 |
2863 |
45.61 |
|
|
|
| 8 |
A |
1521 |
1496 |
27.96 |
|
|
|
| 9 |
A |
1512 |
1487 |
6.37 |
|
|
|
| 10 |
A |
1491 |
1467 |
9.37 |
|
|
|
| 11 |
A |
1478 |
1454 |
0.51 |
|
|
|
| 12 |
A |
1436 |
1412 |
2.55 |
|
|
|
| 13 |
A |
1395 |
1372 |
23.85 |
|
|
|
| 14 |
A |
1369 |
1347 |
40.09 |
|
|
|
| 15 |
A |
1283 |
1262 |
15.37 |
|
|
|
| 16 |
A |
1224 |
1204 |
94.23 |
|
|
|
| 17 |
A |
1154 |
1135 |
14.07 |
|
|
|
| 18 |
A |
1086 |
1068 |
60.54 |
|
|
|
| 19 |
A |
1032 |
1015 |
1.73 |
|
|
|
| 20 |
A |
930 |
915 |
0.83 |
|
|
|
| 21 |
A |
854 |
840 |
5.75 |
|
|
|
| 22 |
A |
733 |
721 |
4.31 |
|
|
|
| 23 |
A |
493 |
485 |
2.75 |
|
|
|
| 24 |
A |
384 |
378 |
1.02 |
|
|
|
| 25 |
A |
361 |
355 |
1.25 |
|
|
|
| 26 |
A |
284 |
280 |
1.02 |
|
|
|
| 27 |
A |
247 |
243 |
0.58 |
|
|
|
| 28 |
A |
3083 |
3032 |
15.36 |
|
|
|
| 29 |
A |
3077 |
3027 |
1.39 |
|
|
|
| 30 |
A |
3062 |
3012 |
0.88 |
|
|
|
| 31 |
A |
2982 |
2934 |
14.81 |
|
|
|
| 32 |
A |
2953 |
2905 |
50.19 |
|
|
|
| 33 |
A |
1494 |
1470 |
0.35 |
|
|
|
| 34 |
A |
1479 |
1455 |
11.59 |
|
|
|
| 35 |
A |
1470 |
1446 |
1.66 |
|
|
|
| 36 |
A |
1453 |
1429 |
0.15 |
|
|
|
| 37 |
A |
1366 |
1343 |
44.08 |
|
|
|
| 38 |
A |
1262 |
1242 |
29.57 |
|
|
|
| 39 |
A |
1125 |
1107 |
0.04 |
|
|
|
| 40 |
A |
1028 |
1011 |
0.84 |
|
|
|
| 41 |
A |
951 |
935 |
0.04 |
|
|
|
| 42 |
A |
919 |
904 |
0.21 |
|
|
|
| 43 |
A |
429 |
422 |
5.68 |
|
|
|
| 44 |
A |
336 |
331 |
2.36 |
|
|
|
| 45 |
A |
277 |
273 |
2.08 |
|
|
|
| 46 |
A |
228 |
225 |
1.07 |
|
|
|
| 47 |
A |
186 |
183 |
1.09 |
|
|
|
| 48 |
A |
48 |
47 |
4.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35329.6 cm
-1
Scaled (by 0.9836) Zero Point Vibrational Energy (zpe) 34750.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at LSDA/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.419 |
|
|
|
| 2 |
C |
0.083 |
|
|
|
| 3 |
C |
-0.580 |
|
|
|
| 4 |
C |
-0.414 |
|
|
|
| 5 |
H |
0.230 |
|
|
|
| 6 |
H |
0.199 |
|
|
|
| 7 |
C |
-0.601 |
|
|
|
| 8 |
C |
-0.601 |
|
|
|
| 9 |
H |
0.193 |
|
|
|
| 10 |
H |
0.193 |
|
|
|
| 11 |
H |
0.222 |
|
|
|
| 12 |
H |
0.222 |
|
|
|
| 13 |
H |
0.210 |
|
|
|
| 14 |
H |
0.210 |
|
|
|
| 15 |
H |
0.205 |
|
|
|
| 16 |
H |
0.205 |
|
|
|
| 17 |
H |
0.221 |
|
|
|
| 18 |
H |
0.221 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.185 |
0.452 |
0.000 |
1.268 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.928 |
-2.022 |
0.000 |
| y |
-2.022 |
-37.623 |
0.000 |
| z |
0.000 |
0.000 |
-39.626 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.304 |
-2.022 |
0.000 |
| y |
-2.022 |
2.154 |
0.000 |
| z |
0.000 |
0.000 |
-0.850 |
|
| Polar |
|---|
| 3z2-r2 | -1.700 |
|---|
| x2-y2 | -2.305 |
|---|
| xy | -2.022 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.910 |
-0.519 |
0.000 |
| y |
-0.519 |
9.095 |
0.000 |
| z |
0.000 |
0.000 |
7.813 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |