Jump to
S1C2
S1C3
Energy calculated at PBE1PBE/STO-3G
| | hartrees |
|---|
| Energy at 0K | -98.922993 |
| Energy at 298.15K | -98.922041 |
| HF Energy | -98.922993 |
| Nuclear repulsion energy | 27.305637 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBE1PBE/STO-3G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
-2.024 |
| C2 |
0.000 |
0.000 |
-0.180 |
| N3 |
0.000 |
0.000 |
1.022 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 1.8434 | 3.0454 |
C2 | 1.8434 | | 1.2021 | N3 | 3.0454 | 1.2021 | |
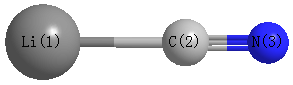 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
180.000 |
|
Li1 |
N3 |
C2 |
0.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/STO-3G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.230 |
|
|
|
| 2 |
C |
-0.014 |
|
|
|
| 3 |
N |
-0.216 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-6.750 |
6.750 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.130 |
0.000 |
0.000 |
| y |
0.000 |
-13.130 |
0.000 |
| z |
0.000 |
0.000 |
-0.182 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.474 |
0.000 |
0.000 |
| y |
0.000 |
-6.474 |
0.000 |
| z |
0.000 |
0.000 |
12.949 |
|
| Polar |
|---|
| 3z2-r2 | 25.897 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.569 |
0.000 |
0.000 |
| y |
0.000 |
1.569 |
0.000 |
| z |
0.000 |
0.000 |
3.849 |
<r2> (average value of r
2) Å
2
| <r2> |
25.294 |
| (<r2>)1/2 |
5.029 |
Jump to
S1C1
S1C3
Energy calculated at PBE1PBE/STO-3G
| | hartrees |
|---|
| Energy at 0K | -98.930447 |
| Energy at 298.15K | -98.929908 |
| HF Energy | -98.930447 |
| Nuclear repulsion energy | 29.493437 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBE1PBE/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
1.249 |
-0.675 |
0.000 |
| C2 |
-0.625 |
-0.404 |
0.000 |
| N3 |
0.000 |
0.636 |
0.000 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 1.8935 | 1.8105 |
C2 | 1.8935 | | 1.2128 | N3 | 1.8105 | 1.2128 | |
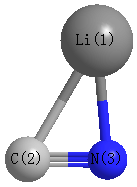 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
67.221 |
|
Li1 |
N3 |
C2 |
74.636 |
| C2 |
Li1 |
N3 |
38.143 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/STO-3G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.219 |
|
|
|
| 2 |
C |
0.017 |
|
|
|
| 3 |
N |
-0.236 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.009 |
-1.826 |
0.000 |
4.405 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-7.182 |
-4.850 |
0.000 |
| y |
-4.850 |
-14.476 |
0.000 |
| z |
0.000 |
0.000 |
-13.676 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.894 |
-4.850 |
0.000 |
| y |
-4.850 |
-4.046 |
0.000 |
| z |
0.000 |
0.000 |
-2.848 |
|
| Polar |
|---|
| 3z2-r2 | -5.696 |
|---|
| x2-y2 | 7.294 |
|---|
| xy | -4.850 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.639 |
-0.380 |
0.000 |
| y |
-0.380 |
2.067 |
0.000 |
| z |
0.000 |
0.000 |
1.652 |
<r2> (average value of r
2) Å
2
| <r2> |
19.554 |
| (<r2>)1/2 |
4.422 |
Jump to
S1C1
S1C2
Energy calculated at PBE1PBE/STO-3G
| | hartrees |
|---|
| Energy at 0K | -98.922762 |
| Energy at 298.15K | -98.921569 |
| HF Energy | -98.922762 |
| Nuclear repulsion energy | 28.132300 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBE1PBE/STO-3G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
1.819 |
| C2 |
0.000 |
0.000 |
-1.077 |
| N3 |
0.000 |
0.000 |
0.144 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.8955 | 1.6748 |
C2 | 2.8955 | | 1.2207 | N3 | 1.6748 | 1.2207 | |
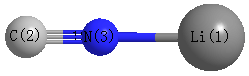 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
0.000 |
|
Li1 |
N3 |
C2 |
180.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/STO-3G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.308 |
|
|
|
| 2 |
C |
-0.005 |
|
|
|
| 3 |
N |
-0.303 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
6.073 |
6.073 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-12.958 |
0.000 |
0.000 |
| y |
0.000 |
-12.958 |
0.000 |
| z |
0.000 |
0.000 |
-3.709 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.625 |
0.000 |
0.000 |
| y |
0.000 |
-4.625 |
0.000 |
| z |
0.000 |
0.000 |
9.249 |
|
| Polar |
|---|
| 3z2-r2 | 18.498 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.403 |
0.000 |
0.000 |
| y |
0.000 |
1.403 |
0.000 |
| z |
0.000 |
0.000 |
4.304 |
<r2> (average value of r
2) Å
2
| <r2> |
23.193 |
| (<r2>)1/2 |
4.816 |
