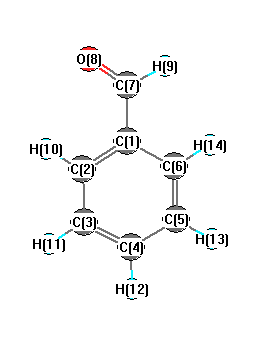
Vibrational Frequencies calculated at PBE1PBE/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3220 |
3089 |
9.26 |
252.30 |
0.13 |
0.23 |
| 2 |
A' |
3214 |
3083 |
9.83 |
59.91 |
0.19 |
0.33 |
| 3 |
A' |
3204 |
3074 |
9.84 |
86.21 |
0.75 |
0.86 |
| 4 |
A' |
3192 |
3063 |
1.09 |
82.65 |
0.71 |
0.83 |
| 5 |
A' |
3181 |
3052 |
4.06 |
48.15 |
0.43 |
0.60 |
| 6 |
A' |
2907 |
2789 |
127.02 |
146.78 |
0.29 |
0.45 |
| 7 |
A' |
1822 |
1748 |
257.68 |
77.47 |
0.37 |
0.54 |
| 8 |
A' |
1672 |
1604 |
28.17 |
76.78 |
0.52 |
0.68 |
| 9 |
A' |
1655 |
1588 |
11.07 |
7.91 |
0.53 |
0.69 |
| 10 |
A' |
1531 |
1468 |
0.66 |
1.37 |
0.28 |
0.44 |
| 11 |
A' |
1494 |
1433 |
14.32 |
1.33 |
0.29 |
0.45 |
| 12 |
A' |
1420 |
1363 |
9.27 |
2.94 |
0.68 |
0.81 |
| 13 |
A' |
1382 |
1326 |
6.22 |
0.87 |
0.71 |
0.83 |
| 14 |
A' |
1335 |
1281 |
15.88 |
1.70 |
0.45 |
0.62 |
| 15 |
A' |
1239 |
1189 |
62.25 |
24.96 |
0.24 |
0.39 |
| 16 |
A' |
1187 |
1139 |
16.09 |
6.33 |
0.17 |
0.28 |
| 17 |
A' |
1183 |
1135 |
2.33 |
6.57 |
0.69 |
0.82 |
| 18 |
A' |
1106 |
1061 |
6.70 |
0.65 |
0.20 |
0.33 |
| 19 |
A' |
1053 |
1010 |
2.68 |
13.36 |
0.09 |
0.17 |
| 20 |
A' |
1021 |
980 |
0.28 |
24.02 |
0.09 |
0.16 |
| 21 |
A' |
847 |
812 |
32.77 |
9.58 |
0.15 |
0.25 |
| 22 |
A' |
664 |
637 |
24.09 |
3.13 |
0.35 |
0.52 |
| 23 |
A' |
628 |
602 |
0.56 |
5.68 |
0.75 |
0.86 |
| 24 |
A' |
444 |
426 |
0.18 |
5.48 |
0.35 |
0.52 |
| 25 |
A' |
217 |
209 |
7.89 |
0.64 |
0.54 |
0.70 |
| 26 |
A" |
1044 |
1002 |
0.53 |
4.72 |
0.75 |
0.86 |
| 27 |
A" |
1021 |
980 |
0.00 |
0.15 |
0.75 |
0.86 |
| 28 |
A" |
1003 |
962 |
0.05 |
0.14 |
0.75 |
0.86 |
| 29 |
A" |
948 |
909 |
1.40 |
0.15 |
0.75 |
0.86 |
| 30 |
A" |
871 |
836 |
0.04 |
0.84 |
0.75 |
0.86 |
| 31 |
A" |
767 |
736 |
48.32 |
0.23 |
0.75 |
0.86 |
| 32 |
A" |
710 |
682 |
34.73 |
0.07 |
0.75 |
0.86 |
| 33 |
A" |
468 |
449 |
8.12 |
0.24 |
0.75 |
0.86 |
| 34 |
A" |
418 |
401 |
0.05 |
0.01 |
0.75 |
0.86 |
| 35 |
A" |
241 |
231 |
8.35 |
1.09 |
0.75 |
0.86 |
| 36 |
A" |
123 |
118 |
3.65 |
2.13 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 24215.5 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 23232.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.224 |
|
|
|
| 2 |
C |
-0.023 |
|
|
|
| 3 |
C |
-0.107 |
|
|
|
| 4 |
C |
-0.082 |
|
|
|
| 5 |
C |
-0.107 |
|
|
|
| 6 |
C |
-0.055 |
|
|
|
| 7 |
C |
0.248 |
|
|
|
| 8 |
O |
-0.295 |
|
|
|
| 9 |
H |
0.074 |
|
|
|
| 10 |
H |
0.122 |
|
|
|
| 11 |
H |
0.115 |
|
|
|
| 12 |
H |
0.115 |
|
|
|
| 13 |
H |
0.112 |
|
|
|
| 14 |
H |
0.108 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.237 |
-2.366 |
0.000 |
3.256 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.008 |
6.029 |
0.000 |
| y |
6.029 |
-46.066 |
0.000 |
| z |
0.000 |
0.000 |
-48.966 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.508 |
6.029 |
0.000 |
| y |
6.029 |
0.421 |
0.000 |
| z |
0.000 |
0.000 |
-3.929 |
|
| Polar |
|---|
| 3z2-r2 | -7.858 |
|---|
| x2-y2 | 2.058 |
|---|
| xy | 6.029 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.935 |
-1.057 |
0.000 |
| y |
-1.057 |
15.168 |
0.000 |
| z |
0.000 |
0.000 |
4.919 |
<r2> (average value of r
2) Å
2
| <r2> |
260.060 |
| (<r2>)1/2 |
16.126 |