Jump to
S1C2
Energy calculated at PBE1PBE/6-311G**
| | hartrees |
|---|
| Energy at 0K | -166.375722 |
| Energy at 298.15K | |
| HF Energy | -166.375722 |
| Nuclear repulsion energy | 49.103972 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
4054 |
3889 |
68.62 |
|
|
|
| 2 |
A1 |
755 |
725 |
8.42 |
|
|
|
| 3 |
A1 |
580 |
557 |
156.27 |
|
|
|
| 4 |
A1 |
319 |
306 |
7.86 |
|
|
|
| 5 |
A2 |
204i |
195i |
0.00 |
|
|
|
| 6 |
B1 |
380 |
365 |
93.89 |
|
|
|
| 7 |
B2 |
4051 |
3887 |
152.58 |
|
|
|
| 8 |
B2 |
1553 |
1490 |
351.23 |
|
|
|
| 9 |
B2 |
452 |
434 |
302.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5970.6 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 5728.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
0.034 |
| O2 |
0.000 |
1.422 |
0.072 |
| O3 |
0.000 |
-1.422 |
0.072 |
| H4 |
0.000 |
2.048 |
-0.643 |
| H5 |
0.000 |
-2.048 |
-0.643 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4226 | 1.4226 | 2.1571 | 2.1571 |
O2 | 1.4226 | | 2.8441 | 0.9506 | 3.5430 | O3 | 1.4226 | 2.8441 | | 3.5430 | 0.9506 | H4 | 2.1571 | 0.9506 | 3.5430 | | 4.0960 | H5 | 2.1571 | 3.5430 | 0.9506 | 4.0960 | |
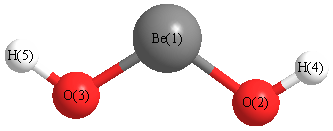 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
129.659 |
|
Be1 |
O3 |
H5 |
129.659 |
| O2 |
Be1 |
O3 |
176.945 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
0.385 |
|
|
|
| 2 |
O |
-0.465 |
|
|
|
| 3 |
O |
-0.465 |
|
|
|
| 4 |
H |
0.272 |
|
|
|
| 5 |
H |
0.272 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.506 |
2.506 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.677 |
0.000 |
0.000 |
| y |
0.000 |
-14.877 |
0.000 |
| z |
0.000 |
0.000 |
-14.254 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.112 |
0.000 |
0.000 |
| y |
0.000 |
0.589 |
0.000 |
| z |
0.000 |
0.000 |
1.523 |
|
| Polar |
|---|
| 3z2-r2 | 3.046 |
|---|
| x2-y2 | -1.800 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.170 |
0.000 |
0.000 |
| y |
0.000 |
3.625 |
0.000 |
| z |
0.000 |
0.000 |
2.357 |
<r2> (average value of r
2) Å
2
| <r2> |
51.196 |
| (<r2>)1/2 |
7.155 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-311G**
| | hartrees |
|---|
| Energy at 0K | -166.376339 |
| Energy at 298.15K | -166.377890 |
| HF Energy | -166.376339 |
| Nuclear repulsion energy | 49.081797 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4036 |
3872 |
42.78 |
|
|
|
| 2 |
A |
750 |
720 |
3.85 |
|
|
|
| 3 |
A |
573 |
550 |
105.20 |
|
|
|
| 4 |
A |
330 |
317 |
29.56 |
|
|
|
| 5 |
A |
234 |
224 |
102.36 |
|
|
|
| 6 |
B |
4034 |
3870 |
163.22 |
|
|
|
| 7 |
B |
1550 |
1488 |
345.22 |
|
|
|
| 8 |
B |
548 |
525 |
326.07 |
|
|
|
| 9 |
B |
329 |
316 |
102.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6192.0 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 5940.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
-0.036 |
| O2 |
0.000 |
1.424 |
-0.055 |
| O3 |
0.000 |
-1.424 |
-0.055 |
| H4 |
0.488 |
2.009 |
0.515 |
| H5 |
-0.488 |
-2.009 |
0.515 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4243 | 1.4243 | 2.1395 | 2.1395 |
O2 | 1.4243 | | 2.8483 | 0.9517 | 3.5141 | O3 | 1.4243 | 2.8483 | | 3.5141 | 0.9517 | H4 | 2.1395 | 0.9517 | 3.5141 | | 4.1345 | H5 | 2.1395 | 3.5141 | 0.9517 | 4.1345 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
127.307 |
|
Be1 |
O3 |
H5 |
127.307 |
| O2 |
Be1 |
O3 |
178.434 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
0.388 |
|
|
|
| 2 |
O |
-0.467 |
|
|
|
| 3 |
O |
-0.467 |
|
|
|
| 4 |
H |
0.273 |
|
|
|
| 5 |
H |
0.273 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.960 |
1.960 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.371 |
4.055 |
0.000 |
| y |
4.055 |
-15.589 |
0.000 |
| z |
0.000 |
0.000 |
-15.147 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.003 |
4.055 |
0.000 |
| y |
4.055 |
-0.330 |
0.000 |
| z |
0.000 |
0.000 |
0.333 |
|
| Polar |
|---|
| 3z2-r2 | 0.665 |
|---|
| x2-y2 | 0.218 |
|---|
| xy | 4.055 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.265 |
0.139 |
0.000 |
| y |
0.139 |
3.611 |
0.000 |
| z |
0.000 |
0.000 |
2.320 |
<r2> (average value of r
2) Å
2
| <r2> |
51.183 |
| (<r2>)1/2 |
7.154 |
