return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CN2 (3H-Diazirin-3-ylidene)

using model chemistry: PBE1PBE/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at PBE1PBE/3-21G*
 hartrees
Energy at 0K-146.383569
Energy at 298.15K-146.383292
HF Energy-146.383569
Nuclear repulsion energy49.705782
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 1285 1234 6.11      
2 A1 902 866 22.16      
3 B2 788 757 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 1488.0 cm-1
Scaled (by 0.9598) Zero Point Vibrational Energy (zpe) 1428.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at PBE1PBE/3-21G*
ABC
1.33057 1.19345 0.62914

See section I.F.4 to change rotational constant units
Geometric Data calculated at PBE1PBE/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.908
N2 0.000 0.673 -0.389
N3 0.000 -0.673 -0.389

Atom - Atom Distances (Å)
  C1 N2 N3
C11.46081.4608
N21.46081.3452
N31.46081.3452

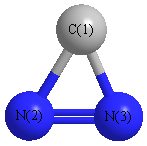
picture of 3H-Diazirin-3-ylidene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 N3 62.585 C1 N3 N2 62.585
N2 C1 N3 54.831
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/3-21G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.315      
2 N -0.158      
3 N -0.158      


Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section VII.A.3)
  x y z Total
  0.000 0.000 -0.107 0.107
CHELPG        
AIM        
ESP        


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -14.916 0.000 0.000
y 0.000 -18.377 0.000
z 0.000 0.000 -19.553
Traceless
 xyz
x 4.049 0.000 0.000
y 0.000 -1.143 0.000
z 0.000 0.000 -2.906
Polar
3z2-r2-5.812
x2-y23.461
xy0.000
xz0.000
yz0.000


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 1.649 0.000 0.000
y 0.000 2.584 0.000
z 0.000 0.000 2.910


<r2> (average value of r2) Å2
<r2> 24.398
(<r2>)1/2 4.939