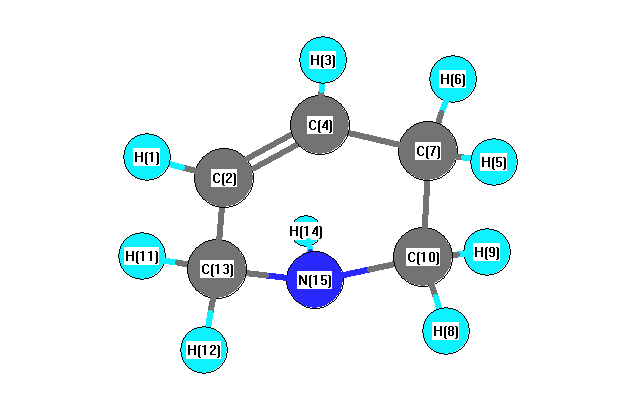
Vibrational Frequencies calculated at B1B95/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3546 |
3397 |
0.55 |
|
|
|
| 2 |
A |
3195 |
3061 |
34.44 |
|
|
|
| 3 |
A |
3170 |
3037 |
9.74 |
|
|
|
| 4 |
A |
3119 |
2988 |
38.87 |
|
|
|
| 5 |
A |
3091 |
2961 |
23.23 |
|
|
|
| 6 |
A |
3080 |
2950 |
47.42 |
|
|
|
| 7 |
A |
3058 |
2929 |
23.61 |
|
|
|
| 8 |
A |
3033 |
2905 |
36.64 |
|
|
|
| 9 |
A |
3019 |
2892 |
43.96 |
|
|
|
| 10 |
A |
1744 |
1671 |
3.44 |
|
|
|
| 11 |
A |
1487 |
1424 |
8.29 |
|
|
|
| 12 |
A |
1471 |
1409 |
3.46 |
|
|
|
| 13 |
A |
1458 |
1396 |
10.50 |
|
|
|
| 14 |
A |
1454 |
1393 |
2.43 |
|
|
|
| 15 |
A |
1414 |
1354 |
0.89 |
|
|
|
| 16 |
A |
1384 |
1326 |
6.06 |
|
|
|
| 17 |
A |
1362 |
1304 |
4.48 |
|
|
|
| 18 |
A |
1334 |
1278 |
0.21 |
|
|
|
| 19 |
A |
1318 |
1263 |
0.57 |
|
|
|
| 20 |
A |
1250 |
1197 |
4.40 |
|
|
|
| 21 |
A |
1222 |
1170 |
17.77 |
|
|
|
| 22 |
A |
1185 |
1135 |
5.91 |
|
|
|
| 23 |
A |
1145 |
1097 |
9.76 |
|
|
|
| 24 |
A |
1098 |
1052 |
1.62 |
|
|
|
| 25 |
A |
1062 |
1017 |
6.13 |
|
|
|
| 26 |
A |
1007 |
965 |
2.86 |
|
|
|
| 27 |
A |
991 |
949 |
0.10 |
|
|
|
| 28 |
A |
973 |
932 |
4.94 |
|
|
|
| 29 |
A |
934 |
895 |
5.41 |
|
|
|
| 30 |
A |
912 |
874 |
13.97 |
|
|
|
| 31 |
A |
866 |
830 |
0.84 |
|
|
|
| 32 |
A |
802 |
768 |
81.05 |
|
|
|
| 33 |
A |
747 |
715 |
43.59 |
|
|
|
| 34 |
A |
644 |
617 |
25.54 |
|
|
|
| 35 |
A |
524 |
502 |
1.25 |
|
|
|
| 36 |
A |
481 |
461 |
0.43 |
|
|
|
| 37 |
A |
399 |
382 |
0.80 |
|
|
|
| 38 |
A |
284 |
272 |
8.03 |
|
|
|
| 39 |
A |
166 |
159 |
2.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29713.2 cm
-1
Scaled (by 0.9579) Zero Point Vibrational Energy (zpe) 28462.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.570 |
|
|
|
| 2 |
C |
0.439 |
|
|
|
| 3 |
H |
-0.592 |
|
|
|
| 4 |
C |
0.192 |
|
|
|
| 5 |
H |
-0.202 |
|
|
|
| 6 |
H |
-0.203 |
|
|
|
| 7 |
C |
0.656 |
|
|
|
| 8 |
H |
-0.344 |
|
|
|
| 9 |
H |
-0.296 |
|
|
|
| 10 |
C |
0.902 |
|
|
|
| 11 |
H |
-0.224 |
|
|
|
| 12 |
H |
-0.215 |
|
|
|
| 13 |
C |
0.782 |
|
|
|
| 14 |
H |
-0.052 |
|
|
|
| 15 |
N |
-0.273 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.524 |
0.695 |
-0.450 |
0.980 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.819 |
-2.001 |
0.216 |
| y |
-2.001 |
-40.506 |
0.856 |
| z |
0.216 |
0.856 |
-37.094 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.981 |
-2.001 |
0.216 |
| y |
-2.001 |
-4.049 |
0.856 |
| z |
0.216 |
0.856 |
1.068 |
|
| Polar |
|---|
| 3z2-r2 | 2.136 |
|---|
| x2-y2 | 4.687 |
|---|
| xy | -2.001 |
|---|
| xz | 0.216 |
|---|
| yz | 0.856 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.750 |
0.021 |
-0.213 |
| y |
0.021 |
10.083 |
0.015 |
| z |
-0.213 |
0.015 |
8.207 |
<r2> (average value of r
2) Å
2
| <r2> |
146.406 |
| (<r2>)1/2 |
12.100 |