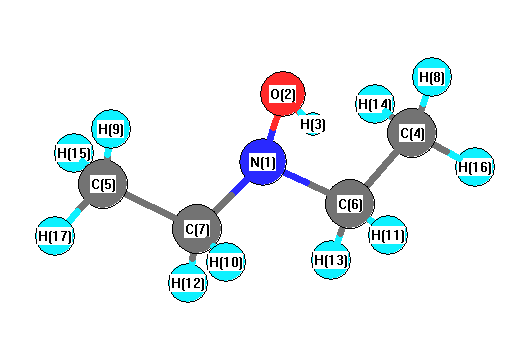
Vibrational Frequencies calculated at B1B95/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3608 |
3451 |
4.84 |
|
|
|
| 2 |
A |
3168 |
3030 |
35.06 |
|
|
|
| 3 |
A |
3152 |
3016 |
32.77 |
|
|
|
| 4 |
A |
3101 |
2967 |
34.20 |
|
|
|
| 5 |
A |
3072 |
2938 |
3.94 |
|
|
|
| 6 |
A |
2915 |
2789 |
175.90 |
|
|
|
| 7 |
A |
1511 |
1445 |
13.05 |
|
|
|
| 8 |
A |
1490 |
1425 |
0.86 |
|
|
|
| 9 |
A |
1485 |
1420 |
23.53 |
|
|
|
| 10 |
A |
1432 |
1370 |
20.72 |
|
|
|
| 11 |
A |
1407 |
1346 |
40.11 |
|
|
|
| 12 |
A |
1385 |
1325 |
0.87 |
|
|
|
| 13 |
A |
1298 |
1242 |
3.59 |
|
|
|
| 14 |
A |
1193 |
1141 |
2.31 |
|
|
|
| 15 |
A |
1165 |
1115 |
15.03 |
|
|
|
| 16 |
A |
1098 |
1051 |
3.14 |
|
|
|
| 17 |
A |
966 |
924 |
20.11 |
|
|
|
| 18 |
A |
826 |
790 |
2.53 |
|
|
|
| 19 |
A |
792 |
758 |
19.86 |
|
|
|
| 20 |
A |
460 |
440 |
5.04 |
|
|
|
| 21 |
A |
427 |
409 |
0.12 |
|
|
|
| 22 |
A |
218 |
209 |
0.33 |
|
|
|
| 23 |
A |
208 |
199 |
2.27 |
|
|
|
| 24 |
A |
114 |
109 |
0.44 |
|
|
|
| 25 |
A |
3167 |
3029 |
0.09 |
|
|
|
| 26 |
A |
3152 |
3015 |
12.52 |
|
|
|
| 27 |
A |
3091 |
2956 |
0.21 |
|
|
|
| 28 |
A |
3071 |
2938 |
49.27 |
|
|
|
| 29 |
A |
2905 |
2779 |
7.65 |
|
|
|
| 30 |
A |
1508 |
1442 |
0.93 |
|
|
|
| 31 |
A |
1486 |
1422 |
1.00 |
|
|
|
| 32 |
A |
1474 |
1410 |
2.45 |
|
|
|
| 33 |
A |
1411 |
1350 |
29.90 |
|
|
|
| 34 |
A |
1377 |
1318 |
21.11 |
|
|
|
| 35 |
A |
1300 |
1243 |
0.80 |
|
|
|
| 36 |
A |
1210 |
1158 |
3.71 |
|
|
|
| 37 |
A |
1101 |
1054 |
4.18 |
|
|
|
| 38 |
A |
1082 |
1035 |
18.41 |
|
|
|
| 39 |
A |
965 |
923 |
0.33 |
|
|
|
| 40 |
A |
805 |
770 |
0.06 |
|
|
|
| 41 |
A |
554 |
530 |
28.72 |
|
|
|
| 42 |
A |
413 |
395 |
106.35 |
|
|
|
| 43 |
A |
345 |
330 |
3.90 |
|
|
|
| 44 |
A |
253 |
242 |
0.00 |
|
|
|
| 45 |
A |
142 |
136 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33651.2 cm
-1
Scaled (by 0.9566) Zero Point Vibrational Energy (zpe) 32190.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/6-31+G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.109 |
|
|
|
| 2 |
O |
-0.293 |
|
|
|
| 3 |
H |
0.314 |
|
|
|
| 4 |
C |
-0.512 |
|
|
|
| 5 |
C |
-0.512 |
|
|
|
| 6 |
C |
-0.237 |
|
|
|
| 7 |
C |
-0.237 |
|
|
|
| 8 |
H |
0.179 |
|
|
|
| 9 |
H |
0.179 |
|
|
|
| 10 |
H |
0.139 |
|
|
|
| 11 |
H |
0.139 |
|
|
|
| 12 |
H |
0.149 |
|
|
|
| 13 |
H |
0.149 |
|
|
|
| 14 |
H |
0.173 |
|
|
|
| 15 |
H |
0.173 |
|
|
|
| 16 |
H |
0.153 |
|
|
|
| 17 |
H |
0.153 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.138 |
2.373 |
0.000 |
2.377 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.347 |
0.692 |
0.000 |
| y |
0.692 |
-41.912 |
0.000 |
| z |
0.000 |
0.000 |
-37.849 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.533 |
0.692 |
0.000 |
| y |
0.692 |
-4.314 |
0.000 |
| z |
0.000 |
0.000 |
1.781 |
|
| Polar |
|---|
| 3z2-r2 | 3.561 |
|---|
| x2-y2 | 4.564 |
|---|
| xy | 0.692 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.957 |
-0.009 |
0.000 |
| y |
-0.009 |
8.877 |
0.000 |
| z |
0.000 |
0.000 |
10.825 |
<r2> (average value of r
2) Å
2
| <r2> |
210.117 |
| (<r2>)1/2 |
14.495 |