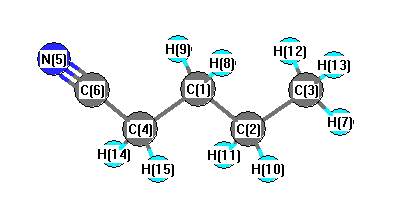
Vibrational Frequencies calculated at B1B95/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3152 |
3006 |
33.37 |
|
|
|
| 2 |
A' |
3073 |
2931 |
39.18 |
|
|
|
| 3 |
A' |
3062 |
2920 |
5.78 |
|
|
|
| 4 |
A' |
3060 |
2918 |
18.74 |
|
|
|
| 5 |
A' |
3044 |
2903 |
22.69 |
|
|
|
| 6 |
A' |
2339 |
2231 |
13.18 |
|
|
|
| 7 |
A' |
1552 |
1480 |
12.38 |
|
|
|
| 8 |
A' |
1540 |
1468 |
1.88 |
|
|
|
| 9 |
A' |
1533 |
1462 |
1.01 |
|
|
|
| 10 |
A' |
1515 |
1444 |
5.17 |
|
|
|
| 11 |
A' |
1457 |
1389 |
6.38 |
|
|
|
| 12 |
A' |
1415 |
1349 |
0.27 |
|
|
|
| 13 |
A' |
1386 |
1322 |
3.47 |
|
|
|
| 14 |
A' |
1317 |
1256 |
0.54 |
|
|
|
| 15 |
A' |
1151 |
1098 |
7.76 |
|
|
|
| 16 |
A' |
1098 |
1047 |
0.41 |
|
|
|
| 17 |
A' |
1052 |
1003 |
0.66 |
|
|
|
| 18 |
A' |
959 |
915 |
0.18 |
|
|
|
| 19 |
A' |
927 |
884 |
4.51 |
|
|
|
| 20 |
A' |
543 |
517 |
2.16 |
|
|
|
| 21 |
A' |
387 |
369 |
0.76 |
|
|
|
| 22 |
A' |
278 |
265 |
0.68 |
|
|
|
| 23 |
A' |
126 |
120 |
5.12 |
|
|
|
| 24 |
A" |
3146 |
3000 |
62.32 |
|
|
|
| 25 |
A" |
3130 |
2985 |
15.42 |
|
|
|
| 26 |
A" |
3105 |
2961 |
0.90 |
|
|
|
| 27 |
A" |
3082 |
2939 |
7.77 |
|
|
|
| 28 |
A" |
1541 |
1470 |
10.77 |
|
|
|
| 29 |
A" |
1350 |
1288 |
0.15 |
|
|
|
| 30 |
A" |
1336 |
1274 |
0.61 |
|
|
|
| 31 |
A" |
1270 |
1211 |
0.10 |
|
|
|
| 32 |
A" |
1170 |
1116 |
0.05 |
|
|
|
| 33 |
A" |
962 |
918 |
1.32 |
|
|
|
| 34 |
A" |
818 |
781 |
0.13 |
|
|
|
| 35 |
A" |
757 |
722 |
6.57 |
|
|
|
| 36 |
A" |
385 |
367 |
0.00 |
|
|
|
| 37 |
A" |
238 |
227 |
0.06 |
|
|
|
| 38 |
A" |
111 |
106 |
0.50 |
|
|
|
| 39 |
A" |
86 |
82 |
3.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29224.7 cm
-1
Scaled (by 0.9537) Zero Point Vibrational Energy (zpe) 27871.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.284 |
|
|
|
| 2 |
C |
-0.293 |
|
|
|
| 3 |
C |
-0.459 |
|
|
|
| 4 |
C |
-0.351 |
|
|
|
| 5 |
N |
-0.289 |
|
|
|
| 6 |
C |
0.130 |
|
|
|
| 7 |
H |
0.159 |
|
|
|
| 8 |
H |
0.173 |
|
|
|
| 9 |
H |
0.173 |
|
|
|
| 10 |
H |
0.154 |
|
|
|
| 11 |
H |
0.154 |
|
|
|
| 12 |
H |
0.157 |
|
|
|
| 13 |
H |
0.157 |
|
|
|
| 14 |
H |
0.209 |
|
|
|
| 15 |
H |
0.209 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.766 |
1.878 |
0.000 |
4.208 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.893 |
-8.165 |
0.000 |
| y |
-8.165 |
-41.733 |
0.000 |
| z |
0.000 |
0.000 |
-36.573 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.740 |
-8.165 |
0.000 |
| y |
-8.165 |
0.500 |
0.000 |
| z |
0.000 |
0.000 |
8.240 |
|
| Polar |
|---|
| 3z2-r2 | 16.480 |
|---|
| x2-y2 | -6.160 |
|---|
| xy | -8.165 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.400 |
1.770 |
0.000 |
| y |
1.770 |
8.314 |
0.000 |
| z |
0.000 |
0.000 |
6.545 |
<r2> (average value of r
2) Å
2
| <r2> |
258.636 |
| (<r2>)1/2 |
16.082 |