Jump to
S1C2
Energy calculated at B2PLYP/6-31G**
| | hartrees |
|---|
| Energy at 0K | -158.263987 |
| Energy at 298.15K | -158.274666 |
| HF Energy | -158.070744 |
| Nuclear repulsion energy | 130.570194 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
126 |
126 |
0.00 |
|
|
|
|
Au |
228 |
228 |
0.00 |
|
|
|
|
Bu |
261 |
261 |
0.04 |
|
|
|
|
Bg |
267 |
267 |
0.00 |
|
|
|
|
Ag |
430 |
430 |
0.00 |
|
|
|
|
Au |
748 |
748 |
3.10 |
|
|
|
|
Bg |
826 |
826 |
0.00 |
|
|
|
|
Ag |
858 |
858 |
0.00 |
|
|
|
|
Au |
979 |
979 |
0.96 |
|
|
|
|
Bu |
996 |
996 |
5.31 |
|
|
|
|
Bu |
1041 |
1041 |
0.22 |
|
|
|
|
Ag |
1089 |
1089 |
0.00 |
|
|
|
|
Ag |
1192 |
1192 |
0.00 |
|
|
|
|
Bg |
1237 |
1237 |
0.00 |
|
|
|
|
Au |
1316 |
1316 |
0.15 |
|
|
|
|
Bu |
1347 |
1347 |
5.26 |
|
|
|
|
Bg |
1357 |
1357 |
0.00 |
|
|
|
|
Ag |
1429 |
1429 |
0.00 |
|
|
|
|
Ag |
1447 |
1447 |
0.00 |
|
|
|
|
Bu |
1449 |
1449 |
2.64 |
|
|
|
|
Ag |
1524 |
1524 |
0.00 |
|
|
|
|
Bu |
1529 |
1529 |
1.33 |
|
|
|
|
Bg |
1537 |
1537 |
0.00 |
|
|
|
|
Au |
1538 |
1538 |
10.72 |
|
|
|
|
Ag |
1544 |
1544 |
0.00 |
|
|
|
|
Bu |
1550 |
1550 |
4.16 |
|
|
|
|
Ag |
3060 |
3060 |
0.00 |
|
|
|
|
Bu |
3068 |
3068 |
45.48 |
|
|
|
|
Bu |
3076 |
3076 |
73.86 |
|
|
|
|
Ag |
3077 |
3077 |
0.00 |
|
|
|
|
Bg |
3091 |
3091 |
0.00 |
|
|
|
|
Au |
3112 |
3112 |
27.77 |
|
|
|
|
Bg |
3151 |
3151 |
0.00 |
|
|
|
|
Au |
3154 |
3154 |
108.05 |
|
|
|
|
Ag |
3156 |
3156 |
0.00 |
|
|
|
|
Bu |
3157 |
3157 |
76.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29472.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 29472.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G**
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.421 |
0.638 |
0.000 |
| C2 |
0.421 |
-0.638 |
0.000 |
| C3 |
0.421 |
1.913 |
0.000 |
| C4 |
-0.421 |
-1.913 |
0.000 |
| H5 |
-1.080 |
0.634 |
0.876 |
| H6 |
-1.080 |
0.634 |
-0.876 |
| H7 |
1.080 |
-0.634 |
0.876 |
| H8 |
1.080 |
-0.634 |
-0.876 |
| H9 |
-0.205 |
2.807 |
0.000 |
| H10 |
1.065 |
1.957 |
0.882 |
| H11 |
1.065 |
1.957 |
-0.882 |
| H12 |
0.205 |
-2.807 |
0.000 |
| H13 |
-1.065 |
-1.957 |
0.882 |
| H14 |
-1.065 |
-1.957 |
-0.882 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5298 | 1.5283 | 2.5517 | 1.0955 | 1.0955 | 2.1537 | 2.1537 | 2.1798 | 2.1743 | 2.1743 | 3.5023 | 2.8160 | 2.8160 |
C2 | 1.5298 | | 2.5517 | 1.5283 | 2.1537 | 2.1537 | 1.0955 | 1.0955 | 3.5023 | 2.8160 | 2.8160 | 2.1798 | 2.1743 | 2.1743 | C3 | 1.5283 | 2.5517 | | 3.9183 | 2.1580 | 2.1580 | 2.7728 | 2.7728 | 1.0919 | 1.0928 | 1.0928 | 4.7256 | 4.2390 | 4.2390 | C4 | 2.5517 | 1.5283 | 3.9183 | | 2.7728 | 2.7728 | 2.1580 | 2.1580 | 4.7256 | 4.2390 | 4.2390 | 1.0919 | 1.0928 | 1.0928 | H5 | 1.0955 | 2.1537 | 2.1580 | 2.7728 | | 1.7515 | 2.5038 | 3.0557 | 2.5011 | 2.5203 | 3.0726 | 3.7763 | 2.5912 | 3.1310 | H6 | 1.0955 | 2.1537 | 2.1580 | 2.7728 | 1.7515 | | 3.0557 | 2.5038 | 2.5011 | 3.0726 | 2.5203 | 3.7763 | 3.1310 | 2.5912 | H7 | 2.1537 | 1.0955 | 2.7728 | 2.1580 | 2.5038 | 3.0557 | | 1.7515 | 3.7763 | 2.5912 | 3.1310 | 2.5011 | 2.5203 | 3.0726 | H8 | 2.1537 | 1.0955 | 2.7728 | 2.1580 | 3.0557 | 2.5038 | 1.7515 | | 3.7763 | 3.1310 | 2.5912 | 2.5011 | 3.0726 | 2.5203 | H9 | 2.1798 | 3.5023 | 1.0919 | 4.7256 | 2.5011 | 2.5011 | 3.7763 | 3.7763 | | 1.7649 | 1.7649 | 5.6298 | 4.9213 | 4.9213 | H10 | 2.1743 | 2.8160 | 1.0928 | 4.2390 | 2.5203 | 3.0726 | 2.5912 | 3.1310 | 1.7649 | | 1.7637 | 4.9213 | 4.4568 | 4.7931 | H11 | 2.1743 | 2.8160 | 1.0928 | 4.2390 | 3.0726 | 2.5203 | 3.1310 | 2.5912 | 1.7649 | 1.7637 | | 4.9213 | 4.7931 | 4.4568 | H12 | 3.5023 | 2.1798 | 4.7256 | 1.0919 | 3.7763 | 3.7763 | 2.5011 | 2.5011 | 5.6298 | 4.9213 | 4.9213 | | 1.7649 | 1.7649 | H13 | 2.8160 | 2.1743 | 4.2390 | 1.0928 | 2.5912 | 3.1310 | 2.5203 | 3.0726 | 4.9213 | 4.4568 | 4.7931 | 1.7649 | | 1.7637 | H14 | 2.8160 | 2.1743 | 4.2390 | 1.0928 | 3.1310 | 2.5912 | 3.0726 | 2.5203 | 4.9213 | 4.7931 | 4.4568 | 1.7649 | 1.7637 | |
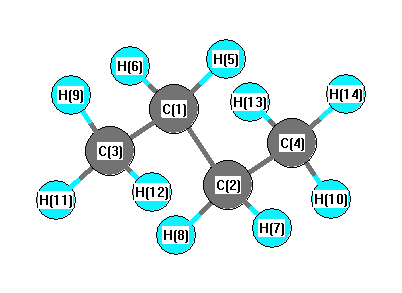 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
113.107 |
|
C1 |
C2 |
H7 |
109.120 |
| C1 |
C2 |
H8 |
109.120 |
|
C1 |
C3 |
H9 |
111.505 |
| C1 |
C3 |
H11 |
111.007 |
|
C1 |
C3 |
H12 |
30.845 |
| C2 |
C1 |
C3 |
113.107 |
|
C2 |
C1 |
H5 |
109.120 |
| C2 |
C1 |
H6 |
109.120 |
|
C2 |
C4 |
H10 |
17.233 |
| C2 |
C4 |
H13 |
111.007 |
|
C2 |
C4 |
H14 |
111.007 |
| C3 |
C1 |
H5 |
109.556 |
|
C3 |
C1 |
H6 |
109.556 |
| C4 |
C2 |
H7 |
109.556 |
|
C4 |
C2 |
H8 |
109.556 |
| H5 |
C1 |
H6 |
106.143 |
|
H7 |
C2 |
H8 |
106.143 |
| H9 |
C3 |
H11 |
107.776 |
|
H9 |
C3 |
H12 |
142.350 |
| H10 |
C4 |
H13 |
94.331 |
|
H10 |
C4 |
H14 |
114.284 |
| H11 |
C3 |
H12 |
93.849 |
|
H13 |
C4 |
H14 |
107.598 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.178 |
|
|
|
| 2 |
C |
-0.178 |
|
|
|
| 3 |
C |
-0.322 |
|
|
|
| 4 |
C |
-0.322 |
|
|
|
| 5 |
H |
0.095 |
|
|
|
| 6 |
H |
0.095 |
|
|
|
| 7 |
H |
0.095 |
|
|
|
| 8 |
H |
0.095 |
|
|
|
| 9 |
H |
0.103 |
|
|
|
| 10 |
H |
0.104 |
|
|
|
| 11 |
H |
0.104 |
|
|
|
| 12 |
H |
0.103 |
|
|
|
| 13 |
H |
0.104 |
|
|
|
| 14 |
H |
0.104 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.482 |
-0.305 |
0.000 |
| y |
-0.305 |
-28.826 |
0.000 |
| z |
0.000 |
0.000 |
-27.909 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.115 |
-0.305 |
0.000 |
| y |
-0.305 |
-0.630 |
0.000 |
| z |
0.000 |
0.000 |
0.746 |
|
| Polar |
|---|
| 3z2-r2 | 1.491 |
|---|
| x2-y2 | 0.343 |
|---|
| xy | -0.305 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.203 |
0.149 |
0.000 |
| y |
0.149 |
7.332 |
0.000 |
| z |
0.000 |
0.000 |
6.056 |
<r2> (average value of r
2) Å
2
| <r2> |
118.981 |
| (<r2>)1/2 |
10.908 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G**
| | hartrees |
|---|
| Energy at 0K | -158.262782 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3167 |
3167 |
37.06 |
|
|
|
| 2 |
A |
3153 |
3153 |
1.00 |
|
|
|
| 3 |
A |
3104 |
3104 |
27.32 |
|
|
|
| 4 |
A |
3081 |
3081 |
21.55 |
|
|
|
| 5 |
A |
3066 |
3066 |
24.06 |
|
|
|
| 6 |
A |
1550 |
1550 |
1.00 |
|
|
|
| 7 |
A |
1544 |
1544 |
3.06 |
|
|
|
| 8 |
A |
1525 |
1525 |
0.04 |
|
|
|
| 9 |
A |
1449 |
1449 |
1.89 |
|
|
|
| 10 |
A |
1410 |
1410 |
0.84 |
|
|
|
| 11 |
A |
1338 |
1338 |
0.20 |
|
|
|
| 12 |
A |
1222 |
1222 |
0.05 |
|
|
|
| 13 |
A |
1109 |
1109 |
0.04 |
|
|
|
| 14 |
A |
1012 |
1012 |
0.20 |
|
|
|
| 15 |
A |
850 |
850 |
0.04 |
|
|
|
| 16 |
A |
809 |
809 |
1.37 |
|
|
|
| 17 |
A |
327 |
327 |
0.03 |
|
|
|
| 18 |
A |
272 |
272 |
0.01 |
|
|
|
| 19 |
A |
114 |
114 |
0.01 |
|
|
|
| 20 |
B |
3160 |
3160 |
52.77 |
|
|
|
| 21 |
B |
3154 |
3154 |
83.45 |
|
|
|
| 22 |
B |
3109 |
3109 |
32.50 |
|
|
|
| 23 |
B |
3078 |
3078 |
22.94 |
|
|
|
| 24 |
B |
3064 |
3064 |
28.87 |
|
|
|
| 25 |
B |
1544 |
1544 |
6.75 |
|
|
|
| 26 |
B |
1536 |
1536 |
6.71 |
|
|
|
| 27 |
B |
1521 |
1521 |
0.22 |
|
|
|
| 28 |
B |
1450 |
1450 |
5.50 |
|
|
|
| 29 |
B |
1398 |
1398 |
4.76 |
|
|
|
| 30 |
B |
1315 |
1315 |
0.55 |
|
|
|
| 31 |
B |
1177 |
1177 |
2.53 |
|
|
|
| 32 |
B |
987 |
987 |
0.12 |
|
|
|
| 33 |
B |
983 |
983 |
3.12 |
|
|
|
| 34 |
B |
761 |
761 |
3.50 |
|
|
|
| 35 |
B |
439 |
439 |
0.38 |
|
|
|
| 36 |
B |
221 |
221 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29498.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 29498.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G**
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
