Jump to
S1C2
Energy calculated at B2PLYP/6-31G**
| | hartrees |
|---|
| Energy at 0K | -247.018718 |
| Energy at 298.15K | -247.024282 |
| HF Energy | -246.769126 |
| Nuclear repulsion energy | 156.758453 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3199 |
3199 |
18.24 |
|
|
|
| 2 |
A' |
3037 |
3037 |
52.48 |
|
|
|
| 3 |
A' |
3022 |
3022 |
23.37 |
|
|
|
| 4 |
A' |
2316 |
2316 |
1.76 |
|
|
|
| 5 |
A' |
1552 |
1552 |
2.69 |
|
|
|
| 6 |
A' |
1533 |
1533 |
7.05 |
|
|
|
| 7 |
A' |
1507 |
1507 |
1.36 |
|
|
|
| 8 |
A' |
1435 |
1435 |
47.20 |
|
|
|
| 9 |
A' |
1241 |
1241 |
46.96 |
|
|
|
| 10 |
A' |
1174 |
1174 |
131.49 |
|
|
|
| 11 |
A' |
1004 |
1004 |
32.03 |
|
|
|
| 12 |
A' |
961 |
961 |
7.42 |
|
|
|
| 13 |
A' |
544 |
544 |
1.39 |
|
|
|
| 14 |
A' |
380 |
380 |
2.24 |
|
|
|
| 15 |
A' |
180 |
180 |
2.31 |
|
|
|
| 16 |
A" |
3097 |
3097 |
50.92 |
|
|
|
| 17 |
A" |
3057 |
3057 |
33.45 |
|
|
|
| 18 |
A" |
1525 |
1525 |
6.74 |
|
|
|
| 19 |
A" |
1275 |
1275 |
3.32 |
|
|
|
| 20 |
A" |
1198 |
1198 |
3.44 |
|
|
|
| 21 |
A" |
1051 |
1051 |
1.85 |
|
|
|
| 22 |
A" |
371 |
371 |
1.93 |
|
|
|
| 23 |
A" |
231 |
231 |
4.43 |
|
|
|
| 24 |
A" |
87 |
87 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17487.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17487.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.134 |
-0.357 |
0.000 |
| O2 |
-0.737 |
-0.606 |
0.000 |
| C3 |
0.000 |
0.593 |
0.000 |
| C4 |
1.433 |
0.269 |
0.000 |
| N5 |
2.578 |
0.047 |
0.000 |
| H6 |
-2.623 |
-1.328 |
0.000 |
| H7 |
-2.442 |
0.203 |
0.891 |
| H8 |
-2.442 |
0.203 |
-0.891 |
| H9 |
-0.220 |
1.206 |
-0.886 |
| H10 |
-0.220 |
1.206 |
0.886 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4184 | 2.3355 | 3.6211 | 4.7292 | 1.0877 | 1.0968 | 1.0968 | 2.6246 | 2.6246 |
O2 | 1.4184 | | 1.4082 | 2.3404 | 3.3794 | 2.0190 | 2.0869 | 2.0869 | 2.0827 | 2.0827 | C3 | 2.3355 | 1.4082 | | 1.4691 | 2.6357 | 3.2514 | 2.6284 | 2.6284 | 1.0994 | 1.0994 | C4 | 3.6211 | 2.3404 | 1.4691 | | 1.1669 | 4.3592 | 3.9765 | 3.9765 | 2.0964 | 2.0964 | N5 | 4.7292 | 3.3794 | 2.6357 | 1.1669 | | 5.3799 | 5.1011 | 5.1011 | 3.1561 | 3.1561 | H6 | 1.0877 | 2.0190 | 3.2514 | 4.3592 | 5.3799 | | 1.7811 | 1.7811 | 3.6029 | 3.6029 | H7 | 1.0968 | 2.0869 | 2.6284 | 3.9765 | 5.1011 | 1.7811 | | 1.7826 | 3.0166 | 2.4375 | H8 | 1.0968 | 2.0869 | 2.6284 | 3.9765 | 5.1011 | 1.7811 | 1.7826 | | 2.4375 | 3.0166 | H9 | 2.6246 | 2.0827 | 1.0994 | 2.0964 | 3.1561 | 3.6029 | 3.0166 | 2.4375 | | 1.7716 | H10 | 2.6246 | 2.0827 | 1.0994 | 2.0964 | 3.1561 | 3.6029 | 2.4375 | 3.0166 | 1.7716 | |
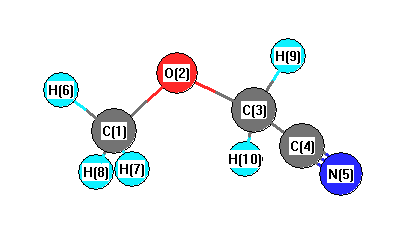 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.435 |
|
O2 |
C1 |
H6 |
106.598 |
| O2 |
C1 |
H7 |
111.500 |
|
O2 |
C1 |
H8 |
111.500 |
| O2 |
C3 |
C4 |
108.842 |
|
O2 |
C3 |
H9 |
111.722 |
| O2 |
C3 |
H10 |
111.722 |
|
C3 |
C4 |
N5 |
178.279 |
| C4 |
C3 |
H9 |
108.550 |
|
C4 |
C3 |
H10 |
108.550 |
| H6 |
C1 |
H7 |
109.243 |
|
H6 |
C1 |
H8 |
109.243 |
| H7 |
C1 |
H8 |
108.703 |
|
H9 |
C3 |
H10 |
107.358 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.063 |
|
|
|
| 2 |
O |
-0.488 |
|
|
|
| 3 |
C |
0.054 |
|
|
|
| 4 |
C |
0.298 |
|
|
|
| 5 |
N |
-0.421 |
|
|
|
| 6 |
H |
0.138 |
|
|
|
| 7 |
H |
0.103 |
|
|
|
| 8 |
H |
0.103 |
|
|
|
| 9 |
H |
0.138 |
|
|
|
| 10 |
H |
0.138 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.275 |
1.790 |
0.000 |
4.635 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.711 |
0.743 |
0.000 |
| y |
0.743 |
-29.068 |
0.000 |
| z |
0.000 |
0.000 |
-29.146 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.604 |
0.743 |
0.000 |
| y |
0.743 |
3.360 |
0.000 |
| z |
0.000 |
0.000 |
3.243 |
|
| Polar |
|---|
| 3z2-r2 | 6.487 |
|---|
| x2-y2 | -6.643 |
|---|
| xy | 0.743 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.864 |
0.005 |
0.000 |
| y |
0.005 |
4.599 |
0.000 |
| z |
0.000 |
0.000 |
4.317 |
<r2> (average value of r
2) Å
2
| <r2> |
143.159 |
| (<r2>)1/2 |
11.965 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G**
| | hartrees |
|---|
| Energy at 0K | -247.021222 |
| Energy at 298.15K | -247.026888 |
| HF Energy | -246.771277 |
| Nuclear repulsion energy | 159.603936 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3200 |
3200 |
18.83 |
|
|
|
| 2 |
A |
3156 |
3156 |
6.06 |
|
|
|
| 3 |
A |
3119 |
3119 |
37.51 |
|
|
|
| 4 |
A |
3049 |
3049 |
42.00 |
|
|
|
| 5 |
A |
3045 |
3045 |
27.77 |
|
|
|
| 6 |
A |
2299 |
2299 |
1.88 |
|
|
|
| 7 |
A |
1545 |
1545 |
8.83 |
|
|
|
| 8 |
A |
1527 |
1527 |
5.93 |
|
|
|
| 9 |
A |
1515 |
1515 |
2.77 |
|
|
|
| 10 |
A |
1504 |
1504 |
0.43 |
|
|
|
| 11 |
A |
1409 |
1409 |
19.33 |
|
|
|
| 12 |
A |
1326 |
1326 |
6.55 |
|
|
|
| 13 |
A |
1233 |
1233 |
36.67 |
|
|
|
| 14 |
A |
1200 |
1200 |
8.62 |
|
|
|
| 15 |
A |
1169 |
1169 |
110.00 |
|
|
|
| 16 |
A |
1046 |
1046 |
6.99 |
|
|
|
| 17 |
A |
953 |
953 |
31.83 |
|
|
|
| 18 |
A |
908 |
908 |
19.59 |
|
|
|
| 19 |
A |
597 |
597 |
2.02 |
|
|
|
| 20 |
A |
394 |
394 |
2.62 |
|
|
|
| 21 |
A |
359 |
359 |
1.24 |
|
|
|
| 22 |
A |
257 |
257 |
10.40 |
|
|
|
| 23 |
A |
178 |
178 |
4.13 |
|
|
|
| 24 |
A |
119 |
119 |
8.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17554.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17554.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.561 |
-0.777 |
0.137 |
| O2 |
1.084 |
0.434 |
-0.437 |
| C3 |
-0.060 |
0.935 |
0.209 |
| C4 |
-1.244 |
0.063 |
0.036 |
| N5 |
-2.163 |
-0.645 |
-0.097 |
| H6 |
2.464 |
-1.041 |
-0.410 |
| H7 |
0.829 |
-1.585 |
0.039 |
| H8 |
1.805 |
-0.642 |
1.197 |
| H9 |
-0.275 |
1.909 |
-0.232 |
| H10 |
0.106 |
1.074 |
1.286 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4233 | 2.3594 | 2.9301 | 3.7337 | 1.0876 | 1.0944 | 1.0964 | 3.2746 | 2.6199 |
O2 | 1.4233 | | 1.4060 | 2.4042 | 3.4381 | 2.0202 | 2.0899 | 2.0861 | 2.0154 | 2.0818 | C3 | 2.3594 | 1.4060 | | 1.4809 | 2.6484 | 3.2646 | 2.6775 | 2.6353 | 1.0906 | 1.0984 | C4 | 2.9301 | 2.4042 | 1.4809 | | 1.1677 | 3.8942 | 2.6479 | 3.3382 | 2.1025 | 2.0992 | N5 | 3.7337 | 3.4381 | 2.6484 | 1.1677 | | 4.6538 | 3.1386 | 4.1737 | 3.1795 | 3.1648 | H6 | 1.0876 | 2.0202 | 3.2646 | 3.8942 | 4.6538 | | 1.7804 | 1.7820 | 4.0288 | 3.5924 | H7 | 1.0944 | 2.0899 | 2.6775 | 2.6479 | 3.1386 | 1.7804 | | 1.7839 | 3.6739 | 3.0239 | H8 | 1.0964 | 2.0861 | 2.6353 | 3.3382 | 4.1737 | 1.7820 | 1.7839 | | 3.5889 | 2.4166 | H9 | 3.2746 | 2.0154 | 1.0906 | 2.1025 | 3.1795 | 4.0288 | 3.6739 | 3.5889 | | 1.7745 | H10 | 2.6199 | 2.0818 | 1.0984 | 2.0992 | 3.1648 | 3.5924 | 3.0239 | 2.4166 | 1.7745 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
113.004 |
|
O2 |
C1 |
H6 |
106.370 |
| O2 |
C1 |
H7 |
111.552 |
|
O2 |
C1 |
H8 |
111.114 |
| O2 |
C3 |
C4 |
112.753 |
|
O2 |
C3 |
H9 |
106.977 |
| O2 |
C3 |
H10 |
111.876 |
|
C3 |
C4 |
N5 |
178.751 |
| C4 |
C3 |
H9 |
108.752 |
|
C4 |
C3 |
H10 |
108.040 |
| H6 |
C1 |
H7 |
109.357 |
|
H6 |
C1 |
H8 |
109.360 |
| H7 |
C1 |
H8 |
109.031 |
|
H9 |
C3 |
H10 |
108.319 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.080 |
|
|
|
| 2 |
O |
-0.478 |
|
|
|
| 3 |
C |
0.023 |
|
|
|
| 4 |
C |
0.290 |
|
|
|
| 5 |
N |
-0.421 |
|
|
|
| 6 |
H |
0.135 |
|
|
|
| 7 |
H |
0.124 |
|
|
|
| 8 |
H |
0.102 |
|
|
|
| 9 |
H |
0.167 |
|
|
|
| 10 |
H |
0.139 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.422 |
1.182 |
1.420 |
3.046 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.000 |
-5.598 |
0.605 |
| y |
-5.598 |
-27.870 |
0.559 |
| z |
0.605 |
0.559 |
-29.607 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.262 |
-5.598 |
0.605 |
| y |
-5.598 |
4.934 |
0.559 |
| z |
0.605 |
0.559 |
2.328 |
|
| Polar |
|---|
| 3z2-r2 | 4.656 |
|---|
| x2-y2 | -8.130 |
|---|
| xy | -5.598 |
|---|
| xz | 0.605 |
|---|
| yz | 0.559 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.660 |
0.446 |
0.172 |
| y |
0.446 |
5.411 |
0.196 |
| z |
0.172 |
0.196 |
4.316 |
<r2> (average value of r
2) Å
2
| <r2> |
123.049 |
| (<r2>)1/2 |
11.093 |
