Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP/daug-cc-pVDZ
Geometric Data calculated at B2PLYP/daug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP/daug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -218.191643 |
| Energy at 298.15K | |
| HF Energy | -217.976453 |
| Nuclear repulsion energy | 129.591064 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/daug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3143 |
3143 |
23.82 |
|
|
|
| 2 |
A |
3131 |
3131 |
54.83 |
|
|
|
| 3 |
A |
3125 |
3125 |
28.27 |
|
|
|
| 4 |
A |
3096 |
3096 |
15.50 |
|
|
|
| 5 |
A |
3067 |
3067 |
33.85 |
|
|
|
| 6 |
A |
3051 |
3051 |
25.23 |
|
|
|
| 7 |
A |
3049 |
3049 |
18.20 |
|
|
|
| 8 |
A |
1500 |
1500 |
2.05 |
|
|
|
| 9 |
A |
1494 |
1494 |
8.17 |
|
|
|
| 10 |
A |
1482 |
1482 |
5.31 |
|
|
|
| 11 |
A |
1461 |
1461 |
2.53 |
|
|
|
| 12 |
A |
1408 |
1408 |
7.46 |
|
|
|
| 13 |
A |
1400 |
1400 |
4.86 |
|
|
|
| 14 |
A |
1367 |
1367 |
0.76 |
|
|
|
| 15 |
A |
1295 |
1295 |
1.18 |
|
|
|
| 16 |
A |
1262 |
1262 |
1.06 |
|
|
|
| 17 |
A |
1169 |
1169 |
1.45 |
|
|
|
| 18 |
A |
1118 |
1118 |
3.63 |
|
|
|
| 19 |
A |
1080 |
1080 |
43.29 |
|
|
|
| 20 |
A |
962 |
962 |
63.35 |
|
|
|
| 21 |
A |
913 |
913 |
4.53 |
|
|
|
| 22 |
A |
879 |
879 |
6.63 |
|
|
|
| 23 |
A |
760 |
760 |
1.03 |
|
|
|
| 24 |
A |
475 |
475 |
4.43 |
|
|
|
| 25 |
A |
314 |
314 |
1.31 |
|
|
|
| 26 |
A |
212 |
212 |
1.65 |
|
|
|
| 27 |
A |
138 |
138 |
2.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21176.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 21176.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/daug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.791 |
0.521 |
0.303 |
| C2 |
-0.598 |
0.651 |
-0.290 |
| C3 |
-1.537 |
-0.486 |
0.120 |
| F4 |
1.412 |
-0.657 |
-0.171 |
| H5 |
1.440 |
1.358 |
0.013 |
| H6 |
0.762 |
0.441 |
1.398 |
| H7 |
-0.511 |
0.702 |
-1.385 |
| H8 |
-1.003 |
1.620 |
0.044 |
| H9 |
-2.531 |
-0.353 |
-0.326 |
| H10 |
-1.143 |
-1.455 |
-0.209 |
| H11 |
-1.659 |
-0.520 |
1.212 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5159 | 2.5437 | 1.4139 | 1.0977 | 1.0991 | 2.1387 | 2.1192 | 3.4926 | 2.8124 | 2.8128 |
C2 | 1.5159 | | 1.5306 | 2.4014 | 2.1787 | 2.1784 | 1.0993 | 1.1016 | 2.1786 | 2.1769 | 2.1797 | C3 | 2.5437 | 1.5306 | | 2.9689 | 3.5040 | 2.7899 | 2.1747 | 2.1735 | 1.0975 | 1.0967 | 1.0991 | F4 | 1.4139 | 2.4014 | 2.9689 | | 2.0232 | 2.0229 | 2.6490 | 3.3259 | 3.9585 | 2.6774 | 3.3705 | H5 | 1.0977 | 2.1787 | 3.5040 | 2.0232 | | 1.7943 | 2.4877 | 2.4575 | 4.3377 | 3.8256 | 3.8167 | H6 | 1.0991 | 2.1784 | 2.7899 | 2.0229 | 1.7943 | | 3.0714 | 2.5176 | 3.8016 | 3.1321 | 2.6114 | H7 | 2.1387 | 1.0993 | 2.1747 | 2.6490 | 2.4877 | 3.0714 | | 1.7678 | 2.5135 | 2.5370 | 3.0908 | H8 | 2.1192 | 1.1016 | 2.1735 | 3.3259 | 2.4575 | 2.5176 | 1.7678 | | 2.5225 | 3.0881 | 2.5242 | H9 | 3.4926 | 2.1786 | 1.0975 | 3.9585 | 4.3377 | 3.8016 | 2.5135 | 2.5225 | | 1.7766 | 1.7760 | H10 | 2.8124 | 2.1769 | 1.0967 | 2.6774 | 3.8256 | 3.1321 | 2.5370 | 3.0881 | 1.7766 | | 1.7769 | H11 | 2.8128 | 2.1797 | 1.0991 | 3.3705 | 3.8167 | 2.6114 | 3.0908 | 2.5242 | 1.7760 | 1.7769 | |
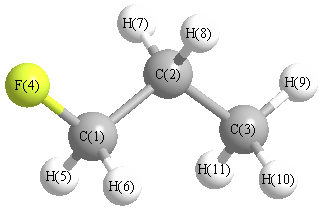 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.219 |
|
C1 |
C2 |
H7 |
108.680 |
| C1 |
C2 |
H8 |
107.051 |
|
C2 |
C1 |
F4 |
110.046 |
| C2 |
C1 |
H5 |
111.949 |
|
C2 |
C1 |
H6 |
111.838 |
| C2 |
C3 |
H9 |
110.912 |
|
C2 |
C3 |
H10 |
110.826 |
| C2 |
C3 |
H11 |
110.902 |
|
C3 |
C2 |
H7 |
110.493 |
| C3 |
C2 |
H8 |
110.267 |
|
F4 |
C1 |
H5 |
106.653 |
| F4 |
C1 |
H6 |
106.549 |
|
H5 |
C1 |
H6 |
109.535 |
| H7 |
C2 |
H8 |
106.876 |
|
H9 |
C3 |
H10 |
108.135 |
| H9 |
C3 |
H11 |
107.902 |
|
H10 |
C3 |
H11 |
108.042 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/daug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.910 |
|
|
|
| 2 |
C |
0.302 |
|
|
|
| 3 |
C |
1.193 |
|
|
|
| 4 |
F |
-0.574 |
|
|
|
| 5 |
H |
-0.464 |
|
|
|
| 6 |
H |
-0.395 |
|
|
|
| 7 |
H |
-0.545 |
|
|
|
| 8 |
H |
-0.608 |
|
|
|
| 9 |
H |
-0.359 |
|
|
|
| 10 |
H |
-0.209 |
|
|
|
| 11 |
H |
-0.251 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.113 |
1.566 |
0.498 |
1.984 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.779 |
1.878 |
0.749 |
| y |
1.878 |
-26.064 |
-0.013 |
| z |
0.749 |
-0.013 |
-25.313 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.091 |
1.878 |
0.749 |
| y |
1.878 |
0.483 |
-0.013 |
| z |
0.749 |
-0.013 |
1.608 |
|
| Polar |
|---|
| 3z2-r2 | 3.217 |
|---|
| x2-y2 | -1.716 |
|---|
| xy | 1.878 |
|---|
| xz | 0.749 |
|---|
| yz | -0.013 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.571 |
0.226 |
0.049 |
| y |
0.226 |
6.124 |
-0.026 |
| z |
0.049 |
-0.026 |
5.793 |
<r2> (average value of r
2) Å
2
| <r2> |
92.940 |
| (<r2>)1/2 |
9.641 |
