Jump to
S1C2
Energy calculated at HSEh1PBE/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -117.132295 |
| Energy at 298.15K | -117.136567 |
| HF Energy | -117.132295 |
| Nuclear repulsion energy | 64.764408 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3277 |
3153 |
13.72 |
|
|
|
| 2 |
A1 |
3173 |
3053 |
1.78 |
|
|
|
| 3 |
A1 |
3168 |
3049 |
12.98 |
|
|
|
| 4 |
A1 |
1506 |
1449 |
2.95 |
|
|
|
| 5 |
A1 |
1269 |
1221 |
1.63 |
|
|
|
| 6 |
A1 |
1037 |
998 |
0.01 |
|
|
|
| 7 |
A1 |
424 |
408 |
0.11 |
|
|
|
| 8 |
A2 |
803 |
773 |
0.00 |
|
|
|
| 9 |
A2 |
551 |
530 |
0.00 |
|
|
|
| 10 |
B1 |
1002 |
964 |
22.00 |
|
|
|
| 11 |
B1 |
827 |
796 |
75.99 |
|
|
|
| 12 |
B1 |
530 |
510 |
17.09 |
|
|
|
| 13 |
B2 |
3275 |
3151 |
3.98 |
|
|
|
| 14 |
B2 |
3165 |
3045 |
7.45 |
|
|
|
| 15 |
B2 |
1512 |
1455 |
0.62 |
|
|
|
| 16 |
B2 |
1408 |
1354 |
6.64 |
|
|
|
| 17 |
B2 |
1215 |
1169 |
0.26 |
|
|
|
| 18 |
B2 |
923 |
888 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14533.0 cm
-1
Scaled (by 0.9622) Zero Point Vibrational Energy (zpe) 13983.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.445 |
| H2 |
0.000 |
0.000 |
1.539 |
| C3 |
0.000 |
1.228 |
-0.197 |
| C4 |
0.000 |
-1.228 |
-0.197 |
| H5 |
0.000 |
2.162 |
0.364 |
| H6 |
0.000 |
-2.162 |
0.364 |
| H7 |
0.000 |
1.293 |
-1.286 |
| H8 |
0.000 |
-1.293 |
-1.286 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0943 | 1.3858 | 1.3858 | 2.1633 | 2.1633 | 2.1605 | 2.1605 |
H2 | 1.0943 | | 2.1266 | 2.1266 | 2.4608 | 2.4608 | 3.1070 | 3.1070 | C3 | 1.3858 | 2.1266 | | 2.4566 | 1.0889 | 3.4361 | 1.0910 | 2.7465 | C4 | 1.3858 | 2.1266 | 2.4566 | | 3.4361 | 1.0889 | 2.7465 | 1.0910 | H5 | 2.1633 | 2.4608 | 1.0889 | 3.4361 | | 4.3236 | 1.8644 | 3.8285 | H6 | 2.1633 | 2.4608 | 3.4361 | 1.0889 | 4.3236 | | 3.8285 | 1.8644 | H7 | 2.1605 | 3.1070 | 1.0910 | 2.7465 | 1.8644 | 3.8285 | | 2.5862 | H8 | 2.1605 | 3.1070 | 2.7465 | 1.0910 | 3.8285 | 1.8644 | 2.5862 | |
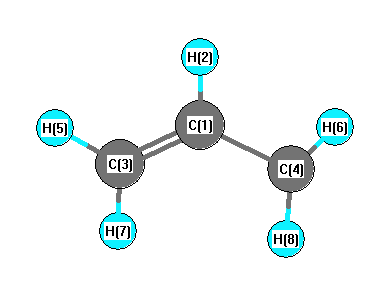 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.434 |
|
C1 |
C3 |
H7 |
120.992 |
| C1 |
C4 |
H6 |
121.434 |
|
C1 |
C4 |
H8 |
120.992 |
| H2 |
C1 |
C3 |
117.585 |
|
H2 |
C1 |
C4 |
117.585 |
| C3 |
C1 |
C4 |
124.829 |
|
H5 |
C3 |
H7 |
117.574 |
| H6 |
C4 |
H8 |
117.574 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.154 |
|
|
|
| 2 |
H |
-0.559 |
|
|
|
| 3 |
C |
0.510 |
|
|
|
| 4 |
C |
0.510 |
|
|
|
| 5 |
H |
-0.393 |
|
|
|
| 6 |
H |
-0.393 |
|
|
|
| 7 |
H |
-0.414 |
|
|
|
| 8 |
H |
-0.414 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.046 |
0.046 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.482 |
0.000 |
0.000 |
| y |
0.000 |
-18.083 |
0.000 |
| z |
0.000 |
0.000 |
-17.722 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.579 |
0.000 |
0.000 |
| y |
0.000 |
2.019 |
0.000 |
| z |
0.000 |
0.000 |
2.560 |
|
| Polar |
|---|
| 3z2-r2 | 5.120 |
|---|
| x2-y2 | -4.399 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.506 |
0.000 |
0.000 |
| y |
0.000 |
8.694 |
0.000 |
| z |
0.000 |
0.000 |
5.490 |
<r2> (average value of r
2) Å
2
| <r2> |
50.524 |
| (<r2>)1/2 |
7.108 |
Jump to
S1C1
Energy calculated at HSEh1PBE/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -117.132295 |
| Energy at 298.15K | -117.136568 |
| HF Energy | -117.132295 |
| Nuclear repulsion energy | 64.766380 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3277 |
3153 |
13.43 |
|
|
|
| 2 |
A' |
3275 |
3151 |
4.23 |
|
|
|
| 3 |
A' |
3174 |
3054 |
2.17 |
|
|
|
| 4 |
A' |
3168 |
3049 |
12.57 |
|
|
|
| 5 |
A' |
3164 |
3045 |
7.46 |
|
|
|
| 6 |
A' |
1512 |
1455 |
0.62 |
|
|
|
| 7 |
A' |
1506 |
1449 |
2.97 |
|
|
|
| 8 |
A' |
1407 |
1354 |
6.66 |
|
|
|
| 9 |
A' |
1269 |
1221 |
1.64 |
|
|
|
| 10 |
A' |
1215 |
1169 |
0.26 |
|
|
|
| 11 |
A' |
1037 |
998 |
0.01 |
|
|
|
| 12 |
A' |
923 |
888 |
0.01 |
|
|
|
| 13 |
A' |
425 |
409 |
0.11 |
|
|
|
| 14 |
A" |
1002 |
964 |
21.99 |
|
|
|
| 15 |
A" |
827 |
796 |
75.97 |
|
|
|
| 16 |
A" |
803 |
773 |
0.00 |
|
|
|
| 17 |
A" |
552 |
531 |
0.00 |
|
|
|
| 18 |
A" |
530 |
510 |
17.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14533.1 cm
-1
Scaled (by 0.9622) Zero Point Vibrational Energy (zpe) 13983.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.445 |
0.000 |
| H2 |
0.000 |
1.539 |
0.000 |
| C3 |
-1.228 |
-0.197 |
0.000 |
| C4 |
1.228 |
-0.197 |
0.000 |
| H5 |
-2.162 |
0.363 |
0.000 |
| H6 |
2.162 |
0.363 |
0.000 |
| H7 |
-1.292 |
-1.286 |
0.000 |
| H8 |
1.292 |
-1.286 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0943 | 1.3858 | 1.3858 | 2.1634 | 2.1633 | 2.1601 | 2.1602 |
H2 | 1.0943 | | 2.1269 | 2.1266 | 2.4613 | 2.4607 | 3.1069 | 3.1067 | C3 | 1.3858 | 2.1269 | | 2.4563 | 1.0890 | 3.4359 | 1.0910 | 2.7454 | C4 | 1.3858 | 2.1266 | 2.4563 | | 3.4360 | 1.0888 | 2.7452 | 1.0911 | H5 | 2.1634 | 2.4613 | 1.0890 | 3.4360 | | 4.3236 | 1.8649 | 3.8275 | H6 | 2.1633 | 2.4607 | 3.4359 | 1.0888 | 4.3236 | | 3.8272 | 1.8647 | H7 | 2.1601 | 3.1069 | 1.0910 | 2.7452 | 1.8649 | 3.8272 | | 2.5837 | H8 | 2.1602 | 3.1067 | 2.7454 | 1.0911 | 3.8275 | 1.8647 | 2.5837 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.434 |
|
C1 |
C3 |
H7 |
120.946 |
| C1 |
C4 |
H6 |
121.431 |
|
C1 |
C4 |
H8 |
120.951 |
| H2 |
C1 |
C3 |
117.613 |
|
H2 |
C1 |
C4 |
117.588 |
| C3 |
C1 |
C4 |
124.799 |
|
H5 |
C3 |
H7 |
117.620 |
| H6 |
C4 |
H8 |
117.618 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.153 |
|
|
|
| 2 |
H |
-0.559 |
|
|
|
| 3 |
C |
0.509 |
|
|
|
| 4 |
C |
0.510 |
|
|
|
| 5 |
H |
-0.392 |
|
|
|
| 6 |
H |
-0.393 |
|
|
|
| 7 |
H |
-0.414 |
|
|
|
| 8 |
H |
-0.414 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.046 |
0.000 |
0.046 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.085 |
0.000 |
0.000 |
| y |
0.000 |
-17.722 |
0.000 |
| z |
0.000 |
0.000 |
-22.482 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.017 |
0.000 |
0.000 |
| y |
0.000 |
2.562 |
0.000 |
| z |
0.000 |
0.000 |
-4.579 |
|
| Polar |
|---|
| 3z2-r2 | -9.157 |
|---|
| x2-y2 | -0.363 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.693 |
-0.000 |
0.000 |
| y |
-0.000 |
5.490 |
0.000 |
| z |
0.000 |
0.000 |
4.506 |
<r2> (average value of r
2) Å
2
| <r2> |
50.516 |
| (<r2>)1/2 |
7.107 |
