Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/aug-cc-pVTZ
Geometric Data calculated at HSEh1PBE/aug-cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1072.689919 |
| Energy at 298.15K | |
| HF Energy | -1072.689919 |
| Nuclear repulsion energy | 267.273384 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3113 |
2994 |
5.62 |
81.19 |
0.15 |
0.26 |
| 2 |
A |
2950 |
2837 |
46.79 |
136.64 |
0.28 |
0.43 |
| 3 |
A |
1862 |
1791 |
154.65 |
20.33 |
0.40 |
0.57 |
| 4 |
A |
1392 |
1339 |
12.32 |
2.72 |
0.34 |
0.51 |
| 5 |
A |
1264 |
1216 |
10.23 |
2.17 |
0.75 |
0.86 |
| 6 |
A |
1211 |
1164 |
9.61 |
4.59 |
0.68 |
0.81 |
| 7 |
A |
1037 |
998 |
14.93 |
1.60 |
0.37 |
0.54 |
| 8 |
A |
941 |
905 |
8.35 |
3.10 |
0.53 |
0.69 |
| 9 |
A |
817 |
786 |
75.12 |
5.30 |
0.69 |
0.81 |
| 10 |
A |
654 |
629 |
35.45 |
12.98 |
0.07 |
0.13 |
| 11 |
A |
621 |
597 |
32.22 |
5.74 |
0.39 |
0.56 |
| 12 |
A |
342 |
329 |
1.81 |
3.24 |
0.12 |
0.22 |
| 13 |
A |
274 |
264 |
3.65 |
3.14 |
0.61 |
0.76 |
| 14 |
A |
216 |
208 |
2.80 |
0.99 |
0.64 |
0.78 |
| 15 |
A |
88 |
85 |
9.81 |
1.26 |
0.72 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 8391.4 cm
-1
Scaled (by 0.9618) Zero Point Vibrational Energy (zpe) 8070.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/aug-cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.095 |
-0.016 |
0.519 |
| C2 |
-0.695 |
-1.280 |
0.203 |
| H3 |
0.227 |
0.072 |
1.596 |
| Cl4 |
1.724 |
-0.247 |
-0.169 |
| Cl5 |
-0.686 |
1.441 |
-0.058 |
| O6 |
-1.764 |
-1.299 |
-0.322 |
| H7 |
-0.163 |
-2.198 |
0.518 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5236 | 1.0893 | 1.7830 | 1.7512 | 2.4099 | 2.1971 |
C2 | 1.5236 | | 2.1493 | 2.6563 | 2.7341 | 1.1908 | 1.1068 | H3 | 1.0893 | 2.1493 | | 2.3364 | 2.3337 | 3.0859 | 2.5429 | Cl4 | 1.7830 | 2.6563 | 2.3364 | | 2.9441 | 3.6459 | 2.8000 | Cl5 | 1.7512 | 2.7341 | 2.3337 | 2.9441 | | 2.9569 | 3.7217 | O6 | 2.4099 | 1.1908 | 3.0859 | 3.6459 | 2.9569 | | 2.0188 | H7 | 2.1971 | 1.1068 | 2.5429 | 2.8000 | 3.7217 | 2.0188 | |
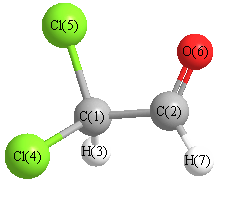 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.750 |
|
C1 |
C2 |
H7 |
112.324 |
| C2 |
C1 |
H3 |
109.562 |
|
C2 |
C1 |
Cl4 |
106.637 |
| C2 |
C1 |
Cl5 |
113.028 |
|
H3 |
C1 |
Cl4 |
106.342 |
| H3 |
C1 |
Cl5 |
108.222 |
|
Cl4 |
C1 |
Cl5 |
112.818 |
| O6 |
C2 |
H7 |
122.926 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.217 |
|
|
|
| 2 |
C |
0.103 |
|
|
|
| 3 |
H |
0.208 |
|
|
|
| 4 |
Cl |
-0.236 |
|
|
|
| 5 |
Cl |
-0.203 |
|
|
|
| 6 |
O |
-0.502 |
|
|
|
| 7 |
H |
0.414 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.345 |
-0.654 |
1.941 |
2.451 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.504 |
-2.243 |
-1.320 |
| y |
-2.243 |
-41.786 |
-1.656 |
| z |
-1.320 |
-1.656 |
-40.287 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.467 |
-2.243 |
-1.320 |
| y |
-2.243 |
2.109 |
-1.656 |
| z |
-1.320 |
-1.656 |
4.358 |
|
| Polar |
|---|
| 3z2-r2 | 8.716 |
|---|
| x2-y2 | -5.717 |
|---|
| xy | -2.243 |
|---|
| xz | -1.320 |
|---|
| yz | -1.656 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.746 |
-0.508 |
0.215 |
| y |
-0.508 |
8.796 |
-0.216 |
| z |
0.215 |
-0.216 |
6.625 |
<r2> (average value of r
2) Å
2
| <r2> |
183.978 |
| (<r2>)1/2 |
13.564 |
