Jump to
S1C2
S1C3
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -225.092723 |
| Energy at 298.15K | -225.098944 |
| HF Energy | -225.092723 |
| Nuclear repulsion energy | 124.071454 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3730 |
3579 |
22.57 |
|
|
|
| 2 |
A |
3617 |
3471 |
1.82 |
|
|
|
| 3 |
A |
1856 |
1781 |
421.07 |
|
|
|
| 4 |
A |
1679 |
1611 |
0.02 |
|
|
|
| 5 |
A |
1203 |
1154 |
3.59 |
|
|
|
| 6 |
A |
971 |
932 |
7.72 |
|
|
|
| 7 |
A |
601 |
577 |
89.70 |
|
|
|
| 8 |
A |
477 |
457 |
4.26 |
|
|
|
| 9 |
A |
389 |
374 |
78.55 |
|
|
|
| 10 |
B |
3730 |
3579 |
22.56 |
|
|
|
| 11 |
B |
3613 |
3467 |
33.95 |
|
|
|
| 12 |
B |
1682 |
1614 |
219.70 |
|
|
|
| 13 |
B |
1444 |
1386 |
208.09 |
|
|
|
| 14 |
B |
1069 |
1026 |
23.11 |
|
|
|
| 15 |
B |
803 |
771 |
97.77 |
|
|
|
| 16 |
B |
588 |
564 |
131.43 |
|
|
|
| 17 |
B |
555 |
532 |
175.38 |
|
|
|
| 18 |
B |
454 |
436 |
126.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14230.1 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 13655.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.145 |
| O2 |
0.000 |
0.000 |
1.356 |
| N3 |
0.000 |
1.156 |
-0.612 |
| N4 |
0.000 |
-1.156 |
-0.612 |
| H5 |
0.202 |
1.978 |
-0.068 |
| H6 |
0.461 |
1.130 |
-1.507 |
| H7 |
-0.202 |
-1.978 |
-0.068 |
| H8 |
-0.461 |
-1.130 |
-1.507 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2108 | 1.3817 | 1.3817 | 2.0000 | 2.0538 | 2.0000 | 2.0538 |
O2 | 1.2108 | | 2.2821 | 2.2821 | 2.4458 | 3.1119 | 2.4458 | 3.1119 | N3 | 1.3817 | 2.2821 | | 2.3119 | 1.0064 | 1.0072 | 3.1875 | 2.4978 | N4 | 1.3817 | 2.2821 | 2.3119 | | 3.1875 | 2.4978 | 1.0064 | 1.0072 | H5 | 2.0000 | 2.4458 | 1.0064 | 3.1875 | | 1.6901 | 3.9771 | 3.4887 | H6 | 2.0538 | 3.1119 | 1.0072 | 2.4978 | 1.6901 | | 3.4887 | 2.4410 | H7 | 2.0000 | 2.4458 | 3.1875 | 1.0064 | 3.9771 | 3.4887 | | 1.6901 | H8 | 2.0538 | 3.1119 | 2.4978 | 1.0072 | 3.4887 | 2.4410 | 1.6901 | |
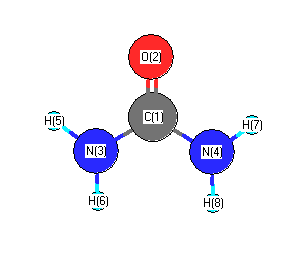 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
112.802 |
|
C1 |
N3 |
H6 |
117.719 |
| C1 |
N4 |
H7 |
112.802 |
|
C1 |
N4 |
H8 |
117.719 |
| O2 |
C1 |
N3 |
123.219 |
|
O2 |
C1 |
N4 |
123.219 |
| N3 |
C1 |
N4 |
113.562 |
|
H5 |
N3 |
H6 |
114.149 |
| H7 |
N4 |
H8 |
114.149 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.577 |
|
|
|
| 2 |
O |
-0.391 |
|
|
|
| 3 |
N |
-0.798 |
|
|
|
| 4 |
N |
-0.798 |
|
|
|
| 5 |
H |
0.361 |
|
|
|
| 6 |
H |
0.344 |
|
|
|
| 7 |
H |
0.361 |
|
|
|
| 8 |
H |
0.344 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.636 |
3.636 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.710 |
3.485 |
0.000 |
| y |
3.485 |
-17.547 |
0.000 |
| z |
0.000 |
0.000 |
-24.816 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.528 |
3.485 |
0.000 |
| y |
3.485 |
7.216 |
0.000 |
| z |
0.000 |
0.000 |
-3.688 |
|
| Polar |
|---|
| 3z2-r2 | -7.377 |
|---|
| x2-y2 | -7.163 |
|---|
| xy | 3.485 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.449 |
0.158 |
0.000 |
| y |
0.158 |
4.708 |
0.000 |
| z |
0.000 |
0.000 |
4.877 |
<r2> (average value of r
2) Å
2
| <r2> |
68.184 |
| (<r2>)1/2 |
8.257 |
Jump to
S1C1
S1C3
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -225.090717 |
| Energy at 298.15K | |
| HF Energy | -225.090717 |
| Nuclear repulsion energy | 124.119621 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3764 |
3612 |
25.31 |
|
|
|
| 2 |
A' |
3644 |
3497 |
7.33 |
|
|
|
| 3 |
A' |
1846 |
1771 |
452.54 |
|
|
|
| 4 |
A' |
1683 |
1615 |
13.56 |
|
|
|
| 5 |
A' |
1193 |
1145 |
2.79 |
|
|
|
| 6 |
A' |
979 |
940 |
8.94 |
|
|
|
| 7 |
A' |
787 |
755 |
28.82 |
|
|
|
| 8 |
A' |
560 |
537 |
36.04 |
|
|
|
| 9 |
A' |
483 |
464 |
13.93 |
|
|
|
| 10 |
A' |
369 |
354 |
440.53 |
|
|
|
| 11 |
A" |
3762 |
3610 |
29.33 |
|
|
|
| 12 |
A" |
3635 |
3488 |
34.38 |
|
|
|
| 13 |
A" |
1670 |
1603 |
242.27 |
|
|
|
| 14 |
A" |
1449 |
1390 |
215.29 |
|
|
|
| 15 |
A" |
1028 |
986 |
18.79 |
|
|
|
| 16 |
A" |
576 |
552 |
22.69 |
|
|
|
| 17 |
A" |
407 |
391 |
56.40 |
|
|
|
| 18 |
A" |
92i |
88i |
49.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13871.2 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 13310.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.008 |
0.141 |
0.000 |
| O2 |
0.034 |
1.354 |
0.000 |
| N3 |
0.034 |
-0.600 |
1.159 |
| N4 |
0.034 |
-0.600 |
-1.159 |
| H5 |
-0.146 |
-0.073 |
1.995 |
| H6 |
-0.250 |
-1.564 |
1.172 |
| H7 |
-0.146 |
-0.073 |
-1.995 |
| H8 |
-0.250 |
-1.564 |
-1.172 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2124 | 1.3761 | 1.3761 | 2.0124 | 2.0852 | 2.0124 | 2.0852 |
O2 | 1.2124 | | 2.2716 | 2.2716 | 2.4594 | 3.1567 | 2.4594 | 3.1567 | N3 | 1.3761 | 2.2716 | | 2.3177 | 1.0044 | 1.0046 | 3.2025 | 2.5380 | N4 | 1.3761 | 2.2716 | 2.3177 | | 3.2025 | 2.5380 | 1.0044 | 1.0046 | H5 | 2.0124 | 2.4594 | 1.0044 | 3.2025 | | 1.7057 | 3.9899 | 3.5016 | H6 | 2.0852 | 3.1567 | 1.0046 | 2.5380 | 1.7057 | | 3.5016 | 2.3438 | H7 | 2.0124 | 2.4594 | 3.2025 | 1.0044 | 3.9899 | 3.5016 | | 1.7057 | H8 | 2.0852 | 3.1567 | 2.5380 | 1.0046 | 3.5016 | 2.3438 | 1.7057 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
114.515 |
|
C1 |
N3 |
H6 |
121.506 |
| C1 |
N4 |
H7 |
114.515 |
|
C1 |
N4 |
H8 |
121.506 |
| O2 |
C1 |
N3 |
122.580 |
|
O2 |
C1 |
N4 |
122.580 |
| N3 |
C1 |
N4 |
114.727 |
|
H5 |
N3 |
H6 |
116.201 |
| H7 |
N4 |
H8 |
116.201 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.615 |
|
|
|
| 2 |
O |
-0.404 |
|
|
|
| 3 |
N |
-0.809 |
|
|
|
| 4 |
N |
-0.809 |
|
|
|
| 5 |
H |
0.365 |
|
|
|
| 6 |
H |
0.339 |
|
|
|
| 7 |
H |
0.365 |
|
|
|
| 8 |
H |
0.339 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.614 |
-4.065 |
0.000 |
4.374 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.414 |
1.684 |
0.000 |
| y |
1.684 |
-24.019 |
0.000 |
| z |
0.000 |
0.000 |
-16.852 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.979 |
1.684 |
0.000 |
| y |
1.684 |
-2.886 |
0.000 |
| z |
0.000 |
0.000 |
7.864 |
|
| Polar |
|---|
| 3z2-r2 | 15.729 |
|---|
| x2-y2 | -1.395 |
|---|
| xy | 1.684 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.360 |
0.115 |
0.000 |
| y |
0.115 |
4.856 |
0.000 |
| z |
0.000 |
0.000 |
4.731 |
<r2> (average value of r
2) Å
2
| <r2> |
68.224 |
| (<r2>)1/2 |
8.260 |
Jump to
S1C1
S1C2
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -225.090303 |
| Energy at 298.15K | |
| HF Energy | -225.090303 |
| Nuclear repulsion energy | 124.231417 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3799 |
3646 |
36.66 |
|
|
|
| 2 |
A1 |
3669 |
3521 |
4.39 |
|
|
|
| 3 |
A1 |
1839 |
1765 |
486.43 |
|
|
|
| 4 |
A1 |
1668 |
1600 |
1.00 |
|
|
|
| 5 |
A1 |
1167 |
1120 |
0.25 |
|
|
|
| 6 |
A1 |
988 |
948 |
7.66 |
|
|
|
| 7 |
A1 |
481 |
462 |
3.40 |
|
|
|
| 8 |
A2 |
374 |
359 |
0.00 |
|
|
|
| 9 |
A2 |
404i |
388i |
0.00 |
|
|
|
| 10 |
B1 |
790 |
758 |
11.38 |
|
|
|
| 11 |
B1 |
578 |
555 |
10.73 |
|
|
|
| 12 |
B1 |
256i |
245i |
561.43 |
|
|
|
| 13 |
B2 |
3797 |
3644 |
36.78 |
|
|
|
| 14 |
B2 |
3660 |
3512 |
50.72 |
|
|
|
| 15 |
B2 |
1662 |
1595 |
305.80 |
|
|
|
| 16 |
B2 |
1453 |
1395 |
221.36 |
|
|
|
| 17 |
B2 |
1002 |
961 |
11.50 |
|
|
|
| 18 |
B2 |
568 |
546 |
14.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13418.0 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 12875.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.142 |
| O2 |
0.000 |
0.000 |
1.356 |
| N3 |
0.000 |
1.154 |
-0.595 |
| N4 |
0.000 |
-1.154 |
-0.595 |
| H5 |
0.000 |
2.017 |
-0.085 |
| H6 |
0.000 |
1.186 |
-1.597 |
| H7 |
0.000 |
-2.017 |
-0.085 |
| H8 |
0.000 |
-1.186 |
-1.597 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2140 | 1.3698 | 1.3698 | 2.0300 | 2.1051 | 2.0300 | 2.1051 |
O2 | 1.2140 | | 2.2672 | 2.2672 | 2.4792 | 3.1824 | 2.4792 | 3.1824 | N3 | 1.3698 | 2.2672 | | 2.3089 | 1.0022 | 1.0023 | 3.2124 | 2.5461 | N4 | 1.3698 | 2.2672 | 2.3089 | | 3.2124 | 2.5461 | 1.0022 | 1.0023 | H5 | 2.0300 | 2.4792 | 1.0022 | 3.2124 | | 1.7251 | 4.0345 | 3.5424 | H6 | 2.1051 | 3.1824 | 1.0023 | 2.5461 | 1.7251 | | 3.5424 | 2.3726 | H7 | 2.0300 | 2.4792 | 3.2124 | 1.0022 | 4.0345 | 3.5424 | | 1.7251 | H8 | 2.1051 | 3.1824 | 2.5461 | 1.0023 | 3.5424 | 2.3726 | 1.7251 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
116.849 |
|
C1 |
N3 |
H6 |
124.383 |
| C1 |
N4 |
H7 |
116.849 |
|
C1 |
N4 |
H8 |
124.383 |
| O2 |
C1 |
N3 |
122.562 |
|
O2 |
C1 |
N4 |
122.562 |
| N3 |
C1 |
N4 |
114.876 |
|
H5 |
N3 |
H6 |
118.768 |
| H7 |
N4 |
H8 |
118.768 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.669 |
|
|
|
| 2 |
O |
-0.417 |
|
|
|
| 3 |
N |
-0.849 |
|
|
|
| 4 |
N |
-0.849 |
|
|
|
| 5 |
H |
0.374 |
|
|
|
| 6 |
H |
0.349 |
|
|
|
| 7 |
H |
0.374 |
|
|
|
| 8 |
H |
0.349 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.384 |
4.384 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.773 |
0.000 |
0.000 |
| y |
0.000 |
-16.121 |
0.000 |
| z |
0.000 |
0.000 |
-23.517 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.954 |
0.000 |
0.000 |
| y |
0.000 |
8.524 |
0.000 |
| z |
0.000 |
0.000 |
-2.570 |
|
| Polar |
|---|
| 3z2-r2 | -5.140 |
|---|
| x2-y2 | -9.652 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.280 |
0.000 |
0.000 |
| y |
0.000 |
4.710 |
0.000 |
| z |
0.000 |
0.000 |
4.798 |
<r2> (average value of r
2) Å
2
| <r2> |
68.137 |
| (<r2>)1/2 |
8.255 |
