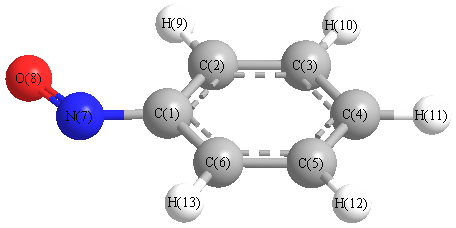
Vibrational Frequencies calculated at HSEh1PBE/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3258 |
3128 |
5.79 |
|
|
|
| 2 |
A' |
3251 |
3120 |
3.41 |
|
|
|
| 3 |
A' |
3240 |
3110 |
4.14 |
|
|
|
| 4 |
A' |
3232 |
3103 |
4.67 |
|
|
|
| 5 |
A' |
3220 |
3091 |
0.35 |
|
|
|
| 6 |
A' |
1650 |
1584 |
6.68 |
|
|
|
| 7 |
A' |
1641 |
1576 |
9.17 |
|
|
|
| 8 |
A' |
1536 |
1475 |
7.39 |
|
|
|
| 9 |
A' |
1520 |
1459 |
16.30 |
|
|
|
| 10 |
A' |
1387 |
1332 |
9.76 |
|
|
|
| 11 |
A' |
1375 |
1320 |
15.93 |
|
|
|
| 12 |
A' |
1354 |
1300 |
60.08 |
|
|
|
| 13 |
A' |
1232 |
1183 |
0.88 |
|
|
|
| 14 |
A' |
1231 |
1181 |
1.50 |
|
|
|
| 15 |
A' |
1134 |
1088 |
127.68 |
|
|
|
| 16 |
A' |
1116 |
1072 |
11.92 |
|
|
|
| 17 |
A' |
1057 |
1014 |
9.84 |
|
|
|
| 18 |
A' |
1042 |
1000 |
0.31 |
|
|
|
| 19 |
A' |
820 |
787 |
33.99 |
|
|
|
| 20 |
A' |
683 |
656 |
12.85 |
|
|
|
| 21 |
A' |
652 |
626 |
0.04 |
|
|
|
| 22 |
A' |
459 |
440 |
0.38 |
|
|
|
| 23 |
A' |
264 |
253 |
2.77 |
|
|
|
| 24 |
A" |
1082 |
1039 |
2.70 |
|
|
|
| 25 |
A" |
1057 |
1015 |
0.00 |
|
|
|
| 26 |
A" |
1012 |
972 |
7.54 |
|
|
|
| 27 |
A" |
903 |
867 |
0.00 |
|
|
|
| 28 |
A" |
818 |
785 |
48.85 |
|
|
|
| 29 |
A" |
736 |
706 |
60.10 |
|
|
|
| 30 |
A" |
500 |
480 |
4.54 |
|
|
|
| 31 |
A" |
437 |
420 |
0.03 |
|
|
|
| 32 |
A" |
261 |
251 |
0.06 |
|
|
|
| 33 |
A" |
138 |
132 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21648.2 cm
-1
Scaled (by 0.9599) Zero Point Vibrational Energy (zpe) 20780.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.151 |
|
|
|
| 2 |
C |
-0.176 |
|
|
|
| 3 |
C |
-0.225 |
|
|
|
| 4 |
C |
-0.200 |
|
|
|
| 5 |
C |
-0.225 |
|
|
|
| 6 |
C |
-0.169 |
|
|
|
| 7 |
N |
-0.095 |
|
|
|
| 8 |
O |
-0.272 |
|
|
|
| 9 |
H |
0.252 |
|
|
|
| 10 |
H |
0.236 |
|
|
|
| 11 |
H |
0.236 |
|
|
|
| 12 |
H |
0.236 |
|
|
|
| 13 |
H |
0.250 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.342 |
-3.524 |
0.000 |
3.771 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.119 |
2.680 |
0.000 |
| y |
2.680 |
-47.354 |
0.000 |
| z |
0.000 |
0.000 |
-48.298 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.707 |
2.680 |
0.000 |
| y |
2.680 |
-2.645 |
0.000 |
| z |
0.000 |
0.000 |
-4.062 |
|
| Polar |
|---|
| 3z2-r2 | -8.123 |
|---|
| x2-y2 | 6.235 |
|---|
| xy | 2.680 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.513 |
-1.765 |
0.000 |
| y |
-1.765 |
13.398 |
0.000 |
| z |
0.000 |
0.000 |
2.746 |
<r2> (average value of r
2) Å
2
| <r2> |
249.225 |
| (<r2>)1/2 |
15.787 |