Jump to
S1C2
Energy calculated at M06-2X/3-21G*
| | hartrees |
|---|
| Energy at 0K | -223.912864 |
| Energy at 298.15K | -223.918870 |
| HF Energy | -223.912864 |
| Nuclear repulsion energy | 123.015054 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3755 |
3557 |
53.93 |
|
|
|
| 2 |
A |
3632 |
3440 |
8.47 |
|
|
|
| 3 |
A |
1805 |
1710 |
364.02 |
|
|
|
| 4 |
A |
1695 |
1605 |
5.14 |
|
|
|
| 5 |
A |
1175 |
1113 |
0.75 |
|
|
|
| 6 |
A |
981 |
929 |
5.94 |
|
|
|
| 7 |
A |
504 |
477 |
0.00 |
|
|
|
| 8 |
A |
501 |
475 |
3.15 |
|
|
|
| 9 |
A |
214 |
203 |
0.00 |
|
|
|
| 10 |
B |
3752 |
3554 |
34.08 |
|
|
|
| 11 |
B |
3623 |
3431 |
75.12 |
|
|
|
| 12 |
B |
1666 |
1578 |
207.89 |
|
|
|
| 13 |
B |
1453 |
1376 |
300.22 |
|
|
|
| 14 |
B |
1026 |
972 |
10.67 |
|
|
|
| 15 |
B |
820 |
777 |
188.06 |
|
|
|
| 16 |
B |
584 |
553 |
78.35 |
|
|
|
| 17 |
B |
572 |
542 |
14.29 |
|
|
|
| 18 |
B |
507 |
481 |
604.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14132.1 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 13386.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/3-21G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.146 |
| O2 |
0.000 |
0.000 |
1.382 |
| N3 |
0.000 |
1.156 |
-0.606 |
| N4 |
0.000 |
-1.156 |
-0.606 |
| H5 |
-0.000 |
2.033 |
-0.104 |
| H6 |
-0.000 |
1.170 |
-1.616 |
| H7 |
0.000 |
-2.033 |
-0.104 |
| H8 |
0.000 |
-1.170 |
-1.616 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2358 | 1.3791 | 1.3791 | 2.0479 | 2.1154 | 2.0479 | 2.1154 |
O2 | 1.2358 | | 2.2997 | 2.2997 | 2.5175 | 3.2184 | 2.5175 | 3.2184 | N3 | 1.3791 | 2.2997 | | 2.3115 | 1.0109 | 1.0101 | 3.2278 | 2.5357 | N4 | 1.3791 | 2.2997 | 2.3115 | | 3.2278 | 2.5357 | 1.0109 | 1.0101 | H5 | 2.0479 | 2.5175 | 1.0109 | 3.2278 | | 1.7415 | 4.0653 | 3.5421 | H6 | 2.1154 | 3.2184 | 1.0101 | 2.5357 | 1.7415 | | 3.5421 | 2.3402 | H7 | 2.0479 | 2.5175 | 3.2278 | 1.0109 | 4.0653 | 3.5421 | | 1.7415 | H8 | 2.1154 | 3.2184 | 2.5357 | 1.0101 | 3.5421 | 2.3402 | 1.7415 | |
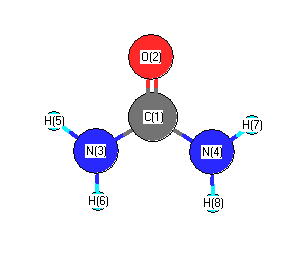 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.105 |
|
C1 |
N3 |
H6 |
123.878 |
| C1 |
N4 |
H7 |
117.105 |
|
C1 |
N4 |
H8 |
123.878 |
| O2 |
C1 |
N3 |
123.063 |
|
O2 |
C1 |
N4 |
123.063 |
| N3 |
C1 |
N4 |
113.875 |
|
H5 |
N3 |
H6 |
119.017 |
| H7 |
N4 |
H8 |
119.017 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.871 |
|
|
|
| 2 |
O |
-0.546 |
|
|
|
| 3 |
N |
-0.811 |
|
|
|
| 4 |
N |
-0.811 |
|
|
|
| 5 |
H |
0.337 |
|
|
|
| 6 |
H |
0.312 |
|
|
|
| 7 |
H |
0.337 |
|
|
|
| 8 |
H |
0.312 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.370 |
4.370 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.248 |
-0.002 |
0.000 |
| y |
-0.002 |
-16.098 |
0.000 |
| z |
0.000 |
0.000 |
-23.565 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.417 |
-0.002 |
0.000 |
| y |
-0.002 |
8.308 |
0.000 |
| z |
0.000 |
0.000 |
-2.892 |
|
| Polar |
|---|
| 3z2-r2 | -5.783 |
|---|
| x2-y2 | -9.150 |
|---|
| xy | -0.002 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.051 |
0.000 |
0.000 |
| y |
0.000 |
3.945 |
0.000 |
| z |
0.000 |
0.000 |
4.273 |
<r2> (average value of r
2) Å
2
| <r2> |
69.014 |
| (<r2>)1/2 |
8.307 |
Jump to
S1C1
Energy calculated at M06-2X/3-21G*
| | hartrees |
|---|
| Energy at 0K | -223.912864 |
| Energy at 298.15K | -223.918870 |
| HF Energy | -223.912864 |
| Nuclear repulsion energy | 123.016230 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3755 |
3557 |
53.92 |
|
|
|
| 2 |
A' |
3632 |
3440 |
8.51 |
|
|
|
| 3 |
A' |
1805 |
1710 |
364.05 |
|
|
|
| 4 |
A' |
1695 |
1605 |
5.21 |
|
|
|
| 5 |
A' |
1175 |
1113 |
0.75 |
|
|
|
| 6 |
A' |
981 |
929 |
5.95 |
|
|
|
| 7 |
A' |
820 |
777 |
188.07 |
|
|
|
| 8 |
A' |
584 |
553 |
77.77 |
|
|
|
| 9 |
A' |
507 |
481 |
604.65 |
|
|
|
| 10 |
A' |
502 |
475 |
3.15 |
|
|
|
| 11 |
A" |
3752 |
3554 |
34.12 |
|
|
|
| 12 |
A" |
3623 |
3431 |
75.01 |
|
|
|
| 13 |
A" |
1666 |
1578 |
207.83 |
|
|
|
| 14 |
A" |
1453 |
1376 |
300.23 |
|
|
|
| 15 |
A" |
1026 |
972 |
10.69 |
|
|
|
| 16 |
A" |
572 |
542 |
14.28 |
|
|
|
| 17 |
A" |
504 |
477 |
0.00 |
|
|
|
| 18 |
A" |
214 |
202 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14132.3 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 13386.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.000 |
0.146 |
0.000 |
| O2 |
0.001 |
1.382 |
0.000 |
| N3 |
-0.000 |
-0.606 |
1.156 |
| N4 |
-0.000 |
-0.606 |
-1.156 |
| H5 |
0.000 |
-0.104 |
2.033 |
| H6 |
-0.001 |
-1.616 |
1.170 |
| H7 |
0.000 |
-0.104 |
-2.033 |
| H8 |
-0.001 |
-1.616 |
-1.170 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2358 | 1.3791 | 1.3791 | 2.0479 | 2.1155 | 2.0479 | 2.1155 |
O2 | 1.2358 | | 2.2997 | 2.2997 | 2.5175 | 3.2185 | 2.5175 | 3.2185 | N3 | 1.3791 | 2.2997 | | 2.3114 | 1.0108 | 1.0101 | 3.2277 | 2.5355 | N4 | 1.3791 | 2.2997 | 2.3114 | | 3.2277 | 2.5355 | 1.0108 | 1.0101 | H5 | 2.0479 | 2.5175 | 1.0108 | 3.2277 | | 1.7415 | 4.0652 | 3.5419 | H6 | 2.1155 | 3.2185 | 1.0101 | 2.5355 | 1.7415 | | 3.5419 | 2.3399 | H7 | 2.0479 | 2.5175 | 3.2277 | 1.0108 | 4.0652 | 3.5419 | | 1.7415 | H8 | 2.1155 | 3.2185 | 2.5355 | 1.0101 | 3.5419 | 2.3399 | 1.7415 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.100 |
|
C1 |
N3 |
H6 |
123.878 |
| C1 |
N4 |
H7 |
117.100 |
|
C1 |
N4 |
H8 |
123.878 |
| O2 |
C1 |
N3 |
123.070 |
|
O2 |
C1 |
N4 |
123.070 |
| N3 |
C1 |
N4 |
113.861 |
|
H5 |
N3 |
H6 |
119.022 |
| H7 |
N4 |
H8 |
119.022 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.871 |
|
|
|
| 2 |
O |
-0.546 |
|
|
|
| 3 |
N |
-0.811 |
|
|
|
| 4 |
N |
-0.811 |
|
|
|
| 5 |
H |
0.337 |
|
|
|
| 6 |
H |
0.312 |
|
|
|
| 7 |
H |
0.337 |
|
|
|
| 8 |
H |
0.312 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.004 |
-4.369 |
0.000 |
4.369 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.248 |
0.000 |
0.000 |
| y |
0.000 |
-23.564 |
0.000 |
| z |
0.000 |
0.000 |
-16.100 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.416 |
0.000 |
0.000 |
| y |
0.000 |
-2.890 |
0.000 |
| z |
0.000 |
0.000 |
8.306 |
|
| Polar |
|---|
| 3z2-r2 | 16.612 |
|---|
| x2-y2 | -1.684 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.051 |
0.003 |
0.000 |
| y |
0.003 |
4.273 |
0.000 |
| z |
0.000 |
0.000 |
3.945 |
<r2> (average value of r
2) Å
2
| <r2> |
69.013 |
| (<r2>)1/2 |
8.307 |
