Jump to
S1C2
Energy calculated at B2PLYP=FULL/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -616.524590 |
| Energy at 298.15K | -616.531702 |
| HF Energy | -616.278900 |
| Nuclear repulsion energy | 204.631611 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3197 |
3075 |
5.75 |
|
|
|
| 2 |
A' |
3150 |
3030 |
14.40 |
|
|
|
| 3 |
A' |
3133 |
3013 |
17.44 |
|
|
|
| 4 |
A' |
3085 |
2968 |
18.21 |
|
|
|
| 5 |
A' |
3043 |
2926 |
29.92 |
|
|
|
| 6 |
A' |
1744 |
1678 |
0.69 |
|
|
|
| 7 |
A' |
1481 |
1424 |
12.11 |
|
|
|
| 8 |
A' |
1467 |
1411 |
2.27 |
|
|
|
| 9 |
A' |
1404 |
1350 |
3.00 |
|
|
|
| 10 |
A' |
1332 |
1281 |
1.68 |
|
|
|
| 11 |
A' |
1321 |
1271 |
25.83 |
|
|
|
| 12 |
A' |
1295 |
1245 |
10.21 |
|
|
|
| 13 |
A' |
1128 |
1085 |
0.39 |
|
|
|
| 14 |
A' |
1055 |
1015 |
11.63 |
|
|
|
| 15 |
A' |
915 |
880 |
9.41 |
|
|
|
| 16 |
A' |
712 |
685 |
19.57 |
|
|
|
| 17 |
A' |
583 |
560 |
5.34 |
|
|
|
| 18 |
A' |
337 |
324 |
1.24 |
|
|
|
| 19 |
A' |
175 |
168 |
0.79 |
|
|
|
| 20 |
A" |
3138 |
3019 |
2.89 |
|
|
|
| 21 |
A" |
3105 |
2986 |
16.55 |
|
|
|
| 22 |
A" |
1466 |
1410 |
6.33 |
|
|
|
| 23 |
A" |
1192 |
1146 |
2.69 |
|
|
|
| 24 |
A" |
1064 |
1023 |
1.49 |
|
|
|
| 25 |
A" |
984 |
946 |
35.81 |
|
|
|
| 26 |
A" |
950 |
914 |
0.10 |
|
|
|
| 27 |
A" |
731 |
703 |
0.53 |
|
|
|
| 28 |
A" |
245 |
235 |
4.16 |
|
|
|
| 29 |
A" |
210 |
202 |
2.20 |
|
|
|
| 30 |
A" |
119 |
115 |
0.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21879.1 cm
-1
Scaled (by 0.9618) Zero Point Vibrational Energy (zpe) 21043.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.321 |
0.311 |
0.000 |
| C2 |
0.000 |
1.022 |
0.000 |
| C3 |
1.223 |
0.475 |
0.000 |
| C4 |
2.502 |
1.261 |
0.000 |
| Cl5 |
-1.197 |
-1.496 |
0.000 |
| H6 |
-1.907 |
0.571 |
0.889 |
| H7 |
-1.907 |
0.571 |
-0.889 |
| H8 |
-0.101 |
2.114 |
0.000 |
| H9 |
1.309 |
-0.613 |
0.000 |
| H10 |
2.311 |
2.343 |
0.000 |
| H11 |
3.110 |
1.017 |
0.883 |
| H12 |
3.110 |
1.017 |
-0.883 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5002 | 2.5485 | 3.9389 | 1.8111 | 1.0968 | 1.0968 | 2.1765 | 2.7868 | 4.1615 | 4.5729 | 4.5729 |
C2 | 1.5002 | | 1.3395 | 2.5132 | 2.7887 | 2.1522 | 2.1522 | 1.0959 | 2.0949 | 2.6617 | 3.2333 | 3.2333 | C3 | 2.5485 | 1.3395 | | 1.5015 | 3.1211 | 3.2551 | 3.2551 | 2.1065 | 1.0920 | 2.1615 | 2.1535 | 2.1535 | C4 | 3.9389 | 2.5132 | 1.5015 | | 4.6137 | 4.5505 | 4.5505 | 2.7392 | 2.2223 | 1.0979 | 1.1001 | 1.1001 | Cl5 | 1.8111 | 2.7887 | 3.1211 | 4.6137 | | 2.3600 | 2.3600 | 3.7725 | 2.6567 | 5.2005 | 5.0647 | 5.0647 | H6 | 1.0968 | 2.1522 | 3.2551 | 4.5505 | 2.3600 | | 1.7789 | 2.5360 | 3.5406 | 4.6609 | 5.0372 | 5.3401 | H7 | 1.0968 | 2.1522 | 3.2551 | 4.5505 | 2.3600 | 1.7789 | | 2.5360 | 3.5406 | 4.6609 | 5.3401 | 5.0372 | H8 | 2.1765 | 1.0959 | 2.1065 | 2.7392 | 3.7725 | 2.5360 | 2.5360 | | 3.0700 | 2.4234 | 3.5067 | 3.5067 | H9 | 2.7868 | 2.0949 | 1.0920 | 2.2223 | 2.6567 | 3.5406 | 3.5406 | 3.0700 | | 3.1214 | 2.5856 | 2.5856 | H10 | 4.1615 | 2.6617 | 2.1615 | 1.0979 | 5.2005 | 4.6609 | 4.6609 | 2.4234 | 3.1214 | | 1.7820 | 1.7820 | H11 | 4.5729 | 3.2333 | 2.1535 | 1.1001 | 5.0647 | 5.0372 | 5.3401 | 3.5067 | 2.5856 | 1.7820 | | 1.7668 | H12 | 4.5729 | 3.2333 | 2.1535 | 1.1001 | 5.0647 | 5.3401 | 5.0372 | 3.5067 | 2.5856 | 1.7820 | 1.7668 | |
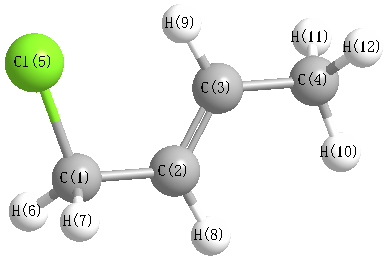 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.560 |
|
C1 |
C2 |
H8 |
113.019 |
| C2 |
C1 |
Cl5 |
114.417 |
|
C2 |
C1 |
H6 |
110.982 |
| C2 |
C1 |
H7 |
110.982 |
|
C2 |
C3 |
C4 |
124.314 |
| C2 |
C3 |
H9 |
118.633 |
|
C3 |
C2 |
H8 |
119.421 |
| C3 |
C4 |
H10 |
111.572 |
|
C3 |
C4 |
H11 |
110.794 |
| C3 |
C4 |
H12 |
110.794 |
|
C4 |
C3 |
H9 |
117.053 |
| Cl5 |
C1 |
H6 |
105.867 |
|
Cl5 |
C1 |
H7 |
105.867 |
| H6 |
C1 |
H7 |
108.369 |
|
H10 |
C4 |
H11 |
108.334 |
| H10 |
C4 |
H12 |
108.334 |
|
H11 |
C4 |
H12 |
106.845 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -616.526865 |
| Energy at 298.15K | -616.533921 |
| HF Energy | -616.281459 |
| Nuclear repulsion energy | 200.214553 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3187 |
3066 |
14.05 |
|
|
|
| 2 |
A |
3173 |
3051 |
1.47 |
|
|
|
| 3 |
A |
3161 |
3041 |
20.80 |
|
|
|
| 4 |
A |
3137 |
3017 |
7.53 |
|
|
|
| 5 |
A |
3107 |
2988 |
12.22 |
|
|
|
| 6 |
A |
3105 |
2987 |
16.44 |
|
|
|
| 7 |
A |
3044 |
2927 |
25.42 |
|
|
|
| 8 |
A |
1730 |
1663 |
13.02 |
|
|
|
| 9 |
A |
1479 |
1422 |
14.67 |
|
|
|
| 10 |
A |
1469 |
1413 |
2.72 |
|
|
|
| 11 |
A |
1465 |
1409 |
7.12 |
|
|
|
| 12 |
A |
1403 |
1349 |
2.34 |
|
|
|
| 13 |
A |
1341 |
1289 |
2.72 |
|
|
|
| 14 |
A |
1328 |
1278 |
0.68 |
|
|
|
| 15 |
A |
1263 |
1215 |
41.34 |
|
|
|
| 16 |
A |
1201 |
1155 |
2.79 |
|
|
|
| 17 |
A |
1118 |
1076 |
0.72 |
|
|
|
| 18 |
A |
1109 |
1066 |
2.03 |
|
|
|
| 19 |
A |
1061 |
1020 |
1.84 |
|
|
|
| 20 |
A |
989 |
952 |
37.91 |
|
|
|
| 21 |
A |
950 |
914 |
9.59 |
|
|
|
| 22 |
A |
891 |
857 |
2.97 |
|
|
|
| 23 |
A |
797 |
766 |
11.77 |
|
|
|
| 24 |
A |
677 |
651 |
62.92 |
|
|
|
| 25 |
A |
494 |
475 |
1.31 |
|
|
|
| 26 |
A |
354 |
341 |
4.36 |
|
|
|
| 27 |
A |
280 |
269 |
2.12 |
|
|
|
| 28 |
A |
214 |
206 |
1.01 |
|
|
|
| 29 |
A |
160 |
154 |
2.55 |
|
|
|
| 30 |
A |
90 |
87 |
0.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21887.1 cm
-1
Scaled (by 0.9618) Zero Point Vibrational Energy (zpe) 21051.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.772 |
0.841 |
0.105 |
| C2 |
0.519 |
0.187 |
0.455 |
| C3 |
1.606 |
0.234 |
-0.333 |
| C4 |
2.930 |
-0.390 |
-0.006 |
| Cl5 |
-2.105 |
-0.405 |
-0.101 |
| H6 |
-1.135 |
1.505 |
0.896 |
| H7 |
-0.714 |
1.384 |
-0.843 |
| H8 |
0.556 |
-0.346 |
1.409 |
| H9 |
1.534 |
0.763 |
-1.289 |
| H10 |
2.902 |
-0.900 |
0.965 |
| H11 |
3.215 |
-1.123 |
-0.776 |
| H12 |
3.726 |
0.369 |
0.023 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4896 | 2.4928 | 3.9032 | 1.8353 | 1.0950 | 1.0943 | 2.2076 | 2.6959 | 4.1561 | 4.5309 | 4.5237 |
C2 | 1.4896 | | 1.3429 | 2.5215 | 2.7468 | 2.1607 | 2.1543 | 1.0932 | 2.0986 | 2.6683 | 3.2399 | 3.2409 | C3 | 2.4928 | 1.3429 | | 1.5001 | 3.7722 | 3.2614 | 2.6389 | 2.1150 | 1.0951 | 2.1567 | 2.1510 | 2.1541 | C4 | 3.9032 | 2.5215 | 1.5001 | | 5.0359 | 4.5750 | 4.1385 | 2.7648 | 2.2188 | 1.0974 | 1.1001 | 1.1001 | Cl5 | 1.8353 | 2.7468 | 3.7722 | 5.0359 | | 2.3620 | 2.3839 | 3.0596 | 4.0023 | 5.1433 | 5.4104 | 5.8832 | H6 | 1.0950 | 2.1607 | 3.2614 | 4.5750 | 2.3620 | | 1.7939 | 2.5589 | 3.5283 | 4.6999 | 5.3500 | 5.0679 | H7 | 1.0943 | 2.1543 | 2.6389 | 4.1385 | 2.3839 | 1.7939 | | 3.1111 | 2.3745 | 4.6438 | 4.6609 | 4.6361 | H8 | 2.2076 | 1.0932 | 2.1150 | 2.7648 | 3.0596 | 2.5589 | 3.1111 | | 3.0769 | 2.4518 | 3.5284 | 3.5335 | H9 | 2.6959 | 2.0986 | 1.0951 | 2.2188 | 4.0023 | 3.5283 | 2.3745 | 3.0769 | | 3.1172 | 2.5778 | 2.5842 | H10 | 4.1561 | 2.6683 | 2.1567 | 1.0974 | 5.1433 | 4.6999 | 4.6438 | 2.4518 | 3.1172 | | 1.7825 | 1.7825 | H11 | 4.5309 | 3.2399 | 2.1510 | 1.1001 | 5.4104 | 5.3500 | 4.6609 | 3.5284 | 2.5778 | 1.7825 | | 1.7674 | H12 | 4.5237 | 3.2409 | 2.1541 | 1.1001 | 5.8832 | 5.0679 | 4.6361 | 3.5335 | 2.5842 | 1.7825 | 1.7674 | |
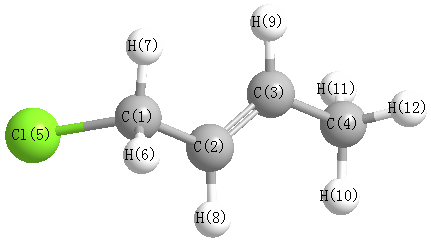 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.220 |
|
C1 |
C2 |
H8 |
116.625 |
| C2 |
C1 |
Cl5 |
110.981 |
|
C2 |
C1 |
H6 |
112.552 |
| C2 |
C1 |
H7 |
112.067 |
|
C2 |
C3 |
C4 |
124.887 |
| C2 |
C3 |
H9 |
118.463 |
|
C3 |
C2 |
H8 |
120.154 |
| C3 |
C4 |
H10 |
111.317 |
|
C3 |
C4 |
H11 |
110.698 |
| C3 |
C4 |
H12 |
110.948 |
|
C4 |
C3 |
H9 |
116.650 |
| Cl5 |
C1 |
H6 |
104.585 |
|
Cl5 |
C1 |
H7 |
106.155 |
| H6 |
C1 |
H7 |
110.057 |
|
H10 |
C4 |
H11 |
108.417 |
| H10 |
C4 |
H12 |
108.416 |
|
H11 |
C4 |
H12 |
106.894 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
