return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2CN (Butanenitrile)

using model chemistry: B2PLYP=FULL/3-21G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP=FULL/3-21G
 hartrees
Energy at 0K-209.918351
Energy at 298.15K-209.925817
HF Energy-209.751823
Nuclear repulsion energy153.198910
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/3-21G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3149 2997 23.67      
2 A' 3098 2949 16.71      
3 A' 3081 2933 1.83      
4 A' 3075 2927 15.29      
5 A' 2273 2163 0.26      
6 A' 1590 1514 7.26      
7 A' 1576 1500 3.20      
8 A' 1555 1480 4.65      
9 A' 1486 1415 4.85      
10 A' 1418 1350 2.65      
11 A' 1368 1302 0.60      
12 A' 1147 1092 6.55      
13 A' 1033 984 0.14      
14 A' 952 906 1.87      
15 A' 893 850 2.80      
16 A' 566 539 1.75      
17 A' 366 348 0.07      
18 A' 185 176 4.03      
19 A" 3154 3003 39.15      
20 A" 3130 2980 1.18      
21 A" 3115 2965 0.69      
22 A" 1589 1512 9.72      
23 A" 1386 1319 0.01      
24 A" 1328 1264 0.00      
25 A" 1192 1134 0.28      
26 A" 925 881 0.42      
27 A" 774 737 5.22      
28 A" 450 429 0.02      
29 A" 249 237 0.02      
30 A" 105 100 3.29      

Unscaled Zero Point Vibrational Energy (zpe) 23101.9 cm-1
Scaled (by 0.952) Zero Point Vibrational Energy (zpe) 21993.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULL/3-21G
ABC
0.78826 0.07454 0.07081

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULL/3-21G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -2.624 0.385 0.000
C2 -1.458 0.504 0.000
C3 0.000 0.637 0.000
C4 0.717 -0.741 0.000
C5 2.249 -0.559 0.000
H6 0.306 1.207 0.884
H7 0.306 1.207 -0.884
H8 0.405 -1.305 0.883
H9 0.405 -1.305 -0.883
H10 2.752 -1.530 0.000
H11 2.572 -0.006 0.887
H12 2.572 -0.006 -0.887

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.17212.63613.52594.96383.16943.16943.57893.57895.70645.28535.2853
C21.17211.46412.50643.85662.09502.09502.74262.74264.67514.15754.1575
C32.63611.46411.55382.54751.09561.09562.17162.17163.50242.79532.7953
C43.52592.50641.55381.54272.17882.17881.09361.09362.18182.18322.1832
C54.96383.85662.54751.54272.77072.77072.17702.17701.09301.09421.0942
H63.16942.09501.09562.17882.77071.76782.51433.07333.77532.56973.1210
H73.16942.09501.09562.17882.77071.76783.07332.51433.77533.12102.5697
H83.57892.74262.17161.09362.17702.51433.07331.76662.51782.52663.0853
H93.57892.74262.17161.09362.17703.07332.51431.76662.51783.08532.5266
H105.70644.67513.50242.18181.09303.77533.77532.51782.51781.77261.7726
H115.28534.15752.79532.18321.09422.56973.12102.52663.08531.77261.7747
H125.28534.15752.79532.18321.09423.12102.56973.08532.52661.77261.7747

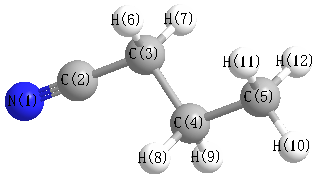
picture of Butanenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 179.390 C2 C3 C4 112.271
C2 C3 H6 109.014 C2 C3 H7 109.014
C3 C4 C5 110.713 C3 C4 H8 108.987
C3 C4 H9 108.987 C4 C3 H6 109.429
C4 C3 H7 109.429 C4 C5 H10 110.587
C4 C5 H11 110.624 C4 C5 H12 110.624
C5 C4 H8 110.169 C5 C4 H9 110.169
H6 C3 H7 107.568 H8 C4 H9 107.748
H10 C5 H11 108.276 H10 C5 H12 108.276
H11 C5 H12 108.370
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability