Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP=FULL/cc-pVDZ
Geometric Data calculated at B2PLYP=FULL/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -1072.600268 |
| Energy at 298.15K | |
| HF Energy | -1072.354657 |
| Nuclear repulsion energy | 264.702776 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3134 |
3004 |
4.40 |
87.96 |
0.20 |
0.34 |
| 2 |
A |
2970 |
2847 |
62.04 |
149.14 |
0.31 |
0.47 |
| 3 |
A |
1825 |
1749 |
123.15 |
12.02 |
0.54 |
0.70 |
| 4 |
A |
1390 |
1333 |
13.41 |
4.88 |
0.54 |
0.70 |
| 5 |
A |
1262 |
1210 |
12.16 |
3.62 |
0.70 |
0.82 |
| 6 |
A |
1211 |
1161 |
12.47 |
8.53 |
0.73 |
0.85 |
| 7 |
A |
1035 |
992 |
11.78 |
1.89 |
0.54 |
0.70 |
| 8 |
A |
938 |
899 |
8.10 |
6.04 |
0.66 |
0.80 |
| 9 |
A |
812 |
779 |
80.56 |
5.97 |
0.73 |
0.84 |
| 10 |
A |
646 |
619 |
36.87 |
9.77 |
0.09 |
0.16 |
| 11 |
A |
614 |
588 |
36.07 |
7.06 |
0.48 |
0.65 |
| 12 |
A |
345 |
330 |
1.57 |
3.42 |
0.24 |
0.38 |
| 13 |
A |
274 |
263 |
3.06 |
6.06 |
0.64 |
0.78 |
| 14 |
A |
220 |
211 |
2.36 |
1.56 |
0.70 |
0.83 |
| 15 |
A |
92 |
88 |
8.69 |
2.22 |
0.73 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 8384.6 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 8036.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.099 |
-0.026 |
0.522 |
| C2 |
-0.705 |
-1.292 |
0.196 |
| H3 |
0.231 |
0.062 |
1.609 |
| Cl4 |
1.748 |
-0.247 |
-0.169 |
| Cl5 |
-0.694 |
1.449 |
-0.058 |
| O6 |
-1.792 |
-1.297 |
-0.319 |
| H7 |
-0.165 |
-2.219 |
0.498 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5347 | 1.0982 | 1.8020 | 1.7716 | 2.4288 | 2.2096 |
C2 | 1.5347 | | 2.1695 | 2.6910 | 2.7520 | 1.2029 | 1.1154 | H3 | 1.0982 | 2.1695 | | 2.3578 | 2.3573 | 3.1078 | 2.5683 | Cl4 | 1.8020 | 2.6910 | 2.3578 | | 2.9748 | 3.6954 | 2.8274 | Cl5 | 1.7716 | 2.7520 | 2.3573 | 2.9748 | | 2.9684 | 3.7475 | O6 | 2.4288 | 1.2029 | 3.1078 | 3.6954 | 2.9684 | | 2.0416 | H7 | 2.2096 | 1.1154 | 2.5683 | 2.8274 | 3.7475 | 2.0416 | |
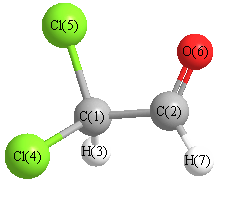 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.606 |
|
C1 |
C2 |
H7 |
112.007 |
| C2 |
C1 |
H3 |
109.859 |
|
C2 |
C1 |
Cl4 |
107.236 |
| C2 |
C1 |
Cl5 |
112.486 |
|
H3 |
C1 |
Cl4 |
106.222 |
| H3 |
C1 |
Cl5 |
108.149 |
|
Cl4 |
C1 |
Cl5 |
112.696 |
| O6 |
C2 |
H7 |
123.387 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
