Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Geometric Data calculated at B2PLYP=FULL/6-311G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -345.051493 |
| Energy at 298.15K | -345.068570 |
| HF Energy | -344.620387 |
| Nuclear repulsion energy | 424.697627 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3090 |
3090 |
0.00 |
|
|
|
| 2 |
A1 |
3068 |
3068 |
0.00 |
|
|
|
| 3 |
A1 |
1535 |
1535 |
0.00 |
|
|
|
| 4 |
A1 |
1377 |
1377 |
0.00 |
|
|
|
| 5 |
A1 |
1284 |
1284 |
0.00 |
|
|
|
| 6 |
A1 |
1032 |
1032 |
0.00 |
|
|
|
| 7 |
A1 |
980 |
980 |
0.00 |
|
|
|
| 8 |
A1 |
818 |
818 |
0.00 |
|
|
|
| 9 |
A1 |
609 |
609 |
0.00 |
|
|
|
| 10 |
A1 |
35 |
35 |
0.00 |
|
|
|
| 11 |
A2 |
3112 |
3112 |
0.00 |
|
|
|
| 12 |
A2 |
3055 |
3055 |
130.61 |
|
|
|
| 13 |
A2 |
1526 |
1526 |
6.76 |
|
|
|
| 14 |
A2 |
1398 |
1398 |
2.15 |
|
|
|
| 15 |
A2 |
1216 |
1216 |
0.00 |
|
|
|
| 16 |
A2 |
1003 |
1003 |
23.16 |
|
|
|
| 17 |
A2 |
815 |
815 |
0.02 |
|
|
|
| 18 |
A2 |
773 |
773 |
71.37 |
|
|
|
| 19 |
E |
3120 |
3120 |
108.43 |
|
|
|
| 19 |
E |
3120 |
3120 |
108.49 |
|
|
|
| 20 |
E |
3094 |
3094 |
0.01 |
|
|
|
| 20 |
E |
3094 |
3094 |
0.01 |
|
|
|
| 21 |
E |
3061 |
3061 |
123.68 |
|
|
|
| 21 |
E |
3061 |
3061 |
123.63 |
|
|
|
| 22 |
E |
3051 |
3051 |
0.00 |
|
|
|
| 22 |
E |
3051 |
3051 |
0.00 |
|
|
|
| 23 |
E |
1524 |
1524 |
13.54 |
|
|
|
| 23 |
E |
1524 |
1524 |
13.55 |
|
|
|
| 24 |
E |
1511 |
1511 |
0.00 |
|
|
|
| 24 |
E |
1511 |
1511 |
0.00 |
|
|
|
| 25 |
E |
1375 |
1375 |
8.44 |
|
|
|
| 25 |
E |
1375 |
1375 |
8.43 |
|
|
|
| 26 |
E |
1360 |
1360 |
0.00 |
|
|
|
| 26 |
E |
1360 |
1360 |
0.00 |
|
|
|
| 27 |
E |
1347 |
1347 |
0.33 |
|
|
|
| 27 |
E |
1347 |
1347 |
0.33 |
|
|
|
| 28 |
E |
1344 |
1344 |
0.01 |
|
|
|
| 28 |
E |
1344 |
1344 |
0.01 |
|
|
|
| 29 |
E |
1219 |
1219 |
0.00 |
|
|
|
| 29 |
E |
1219 |
1219 |
0.00 |
|
|
|
| 30 |
E |
1095 |
1095 |
42.75 |
|
|
|
| 30 |
E |
1095 |
1095 |
42.75 |
|
|
|
| 31 |
E |
1051 |
1051 |
0.00 |
|
|
|
| 31 |
E |
1051 |
1051 |
0.00 |
|
|
|
| 32 |
E |
908 |
908 |
3.88 |
|
|
|
| 32 |
E |
908 |
908 |
3.88 |
|
|
|
| 33 |
E |
837 |
837 |
4.69 |
|
|
|
| 33 |
E |
837 |
837 |
4.69 |
|
|
|
| 34 |
E |
589 |
589 |
0.00 |
|
|
|
| 34 |
E |
589 |
589 |
0.00 |
|
|
|
| 35 |
E |
429 |
429 |
0.00 |
|
|
|
| 35 |
E |
429 |
429 |
0.00 |
|
|
|
| 36 |
E |
332 |
332 |
0.00 |
|
|
|
| 36 |
E |
332 |
332 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40607.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 40607.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is D3
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.289 |
| N2 |
0.000 |
0.000 |
-1.289 |
| C3 |
-0.003 |
1.380 |
0.780 |
| C4 |
1.196 |
-0.688 |
0.780 |
| C5 |
-1.194 |
-0.692 |
0.780 |
| C6 |
0.003 |
1.380 |
-0.780 |
| C7 |
-1.196 |
-0.688 |
-0.780 |
| C8 |
1.194 |
-0.692 |
-0.780 |
| H9 |
0.875 |
1.891 |
1.185 |
| H10 |
-0.887 |
1.885 |
1.179 |
| H11 |
1.201 |
-1.703 |
1.185 |
| H12 |
2.076 |
-0.175 |
1.179 |
| H13 |
-2.075 |
-0.188 |
1.185 |
| H14 |
-1.189 |
-1.710 |
1.179 |
| H15 |
-0.875 |
1.891 |
-1.185 |
| H16 |
0.887 |
1.885 |
-1.179 |
| H17 |
-1.201 |
-1.703 |
-1.185 |
| H18 |
-2.076 |
-0.175 |
-1.179 |
| H19 |
2.075 |
-0.188 |
-1.185 |
| H20 |
1.189 |
-1.710 |
-1.179 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5773 | 1.4706 | 1.4706 | 1.4706 | 2.4868 | 2.4868 | 2.4868 | 2.0863 | 2.0858 | 2.0863 | 2.0858 | 2.0863 | 2.0858 | 3.2346 | 3.2292 | 3.2346 | 3.2292 | 3.2346 | 3.2292 |
N2 | 2.5773 | | 2.4868 | 2.4868 | 2.4868 | 1.4706 | 1.4706 | 1.4706 | 3.2346 | 3.2292 | 3.2346 | 3.2292 | 3.2346 | 3.2292 | 2.0863 | 2.0858 | 2.0863 | 2.0858 | 2.0863 | 2.0858 | C3 | 1.4706 | 2.4868 | | 2.3901 | 2.3901 | 1.5603 | 2.8521 | 2.8566 | 1.0933 | 1.0933 | 3.3343 | 2.6257 | 2.6304 | 3.3340 | 2.2102 | 2.2100 | 3.8475 | 3.2484 | 3.2619 | 3.8480 | C4 | 1.4706 | 2.4868 | 2.3901 | | 2.3901 | 2.8521 | 2.8566 | 1.5603 | 2.6304 | 3.3340 | 1.0933 | 1.0933 | 3.3343 | 2.6257 | 3.8475 | 3.2484 | 3.2619 | 3.8480 | 2.2102 | 2.2100 | C5 | 1.4706 | 2.4868 | 2.3901 | 2.3901 | | 2.8566 | 1.5603 | 2.8521 | 3.3343 | 2.6257 | 2.6304 | 3.3340 | 1.0933 | 1.0933 | 3.2619 | 3.8480 | 2.2102 | 2.2100 | 3.8475 | 3.2484 | C6 | 2.4868 | 1.4706 | 1.5603 | 2.8521 | 2.8566 | | 2.3901 | 2.3901 | 2.2102 | 2.2100 | 3.8475 | 3.2484 | 3.2619 | 3.8480 | 1.0933 | 1.0933 | 3.3343 | 2.6257 | 2.6304 | 3.3340 | C7 | 2.4868 | 1.4706 | 2.8521 | 2.8566 | 1.5603 | 2.3901 | | 2.3901 | 3.8475 | 3.2484 | 3.2619 | 3.8480 | 2.2102 | 2.2100 | 2.6304 | 3.3340 | 1.0933 | 1.0933 | 3.3343 | 2.6257 | C8 | 2.4868 | 1.4706 | 2.8566 | 1.5603 | 2.8521 | 2.3901 | 2.3901 | | 3.2619 | 3.8480 | 2.2102 | 2.2100 | 3.8475 | 3.2484 | 3.3343 | 2.6257 | 2.6304 | 3.3340 | 1.0933 | 1.0933 | H9 | 2.0863 | 3.2346 | 1.0933 | 2.6304 | 3.3343 | 2.2102 | 3.8475 | 3.2619 | | 1.7612 | 3.6092 | 2.3896 | 3.6092 | 4.1508 | 2.9462 | 2.3644 | 4.7799 | 4.3083 | 3.3744 | 4.3197 | H10 | 2.0858 | 3.2292 | 1.0933 | 3.3340 | 2.6257 | 2.2100 | 3.2484 | 3.8480 | 1.7612 | | 4.1508 | 3.6076 | 2.3896 | 3.6076 | 2.3644 | 2.9504 | 4.3083 | 3.3489 | 4.3197 | 4.7741 | H11 | 2.0863 | 3.2346 | 3.3343 | 1.0933 | 2.6304 | 3.8475 | 3.2619 | 2.2102 | 3.6092 | 4.1508 | | 1.7612 | 3.6092 | 2.3896 | 4.7799 | 4.3083 | 3.3744 | 4.3197 | 2.9462 | 2.3644 | H12 | 2.0858 | 3.2292 | 2.6257 | 1.0933 | 3.3340 | 3.2484 | 3.8480 | 2.2100 | 2.3896 | 3.6076 | 1.7612 | | 4.1508 | 3.6076 | 4.3083 | 3.3489 | 4.3197 | 4.7741 | 2.3644 | 2.9504 | H13 | 2.0863 | 3.2346 | 2.6304 | 3.3343 | 1.0933 | 3.2619 | 2.2102 | 3.8475 | 3.6092 | 2.3896 | 3.6092 | 4.1508 | | 1.7612 | 3.3744 | 4.3197 | 2.9462 | 2.3644 | 4.7799 | 4.3083 | H14 | 2.0858 | 3.2292 | 3.3340 | 2.6257 | 1.0933 | 3.8480 | 2.2100 | 3.2484 | 4.1508 | 3.6076 | 2.3896 | 3.6076 | 1.7612 | | 4.3197 | 4.7741 | 2.3644 | 2.9504 | 4.3083 | 3.3489 | H15 | 3.2346 | 2.0863 | 2.2102 | 3.8475 | 3.2619 | 1.0933 | 2.6304 | 3.3343 | 2.9462 | 2.3644 | 4.7799 | 4.3083 | 3.3744 | 4.3197 | | 1.7612 | 3.6092 | 2.3896 | 3.6092 | 4.1508 | H16 | 3.2292 | 2.0858 | 2.2100 | 3.2484 | 3.8480 | 1.0933 | 3.3340 | 2.6257 | 2.3644 | 2.9504 | 4.3083 | 3.3489 | 4.3197 | 4.7741 | 1.7612 | | 4.1508 | 3.6076 | 2.3896 | 3.6076 | H17 | 3.2346 | 2.0863 | 3.8475 | 3.2619 | 2.2102 | 3.3343 | 1.0933 | 2.6304 | 4.7799 | 4.3083 | 3.3744 | 4.3197 | 2.9462 | 2.3644 | 3.6092 | 4.1508 | | 1.7612 | 3.6092 | 2.3896 | H18 | 3.2292 | 2.0858 | 3.2484 | 3.8480 | 2.2100 | 2.6257 | 1.0933 | 3.3340 | 4.3083 | 3.3489 | 4.3197 | 4.7741 | 2.3644 | 2.9504 | 2.3896 | 3.6076 | 1.7612 | | 4.1508 | 3.6076 | H19 | 3.2346 | 2.0863 | 3.2619 | 2.2102 | 3.8475 | 2.6304 | 3.3343 | 1.0933 | 3.3744 | 4.3197 | 2.9462 | 2.3644 | 4.7799 | 4.3083 | 3.6092 | 2.3896 | 3.6092 | 4.1508 | | 1.7612 | H20 | 3.2292 | 2.0858 | 3.8480 | 2.2100 | 3.2484 | 3.3340 | 2.6257 | 1.0933 | 4.3197 | 4.7741 | 2.3644 | 2.9504 | 4.3083 | 3.3489 | 4.1508 | 3.6076 | 2.3896 | 3.6076 | 1.7612 | |
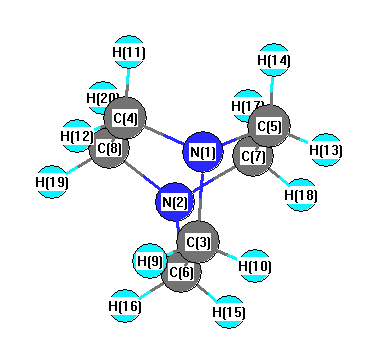 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
110.227 |
|
N1 |
C3 |
H9 |
108.020 |
| N1 |
C3 |
H10 |
107.973 |
|
N1 |
C4 |
C8 |
110.227 |
| N1 |
C4 |
H11 |
108.020 |
|
N1 |
C4 |
H12 |
107.973 |
| N1 |
C5 |
C7 |
110.227 |
|
N1 |
C5 |
H13 |
108.020 |
| N1 |
C5 |
H14 |
107.973 |
|
N2 |
C6 |
C3 |
110.227 |
| N2 |
C6 |
H15 |
108.020 |
|
N2 |
C6 |
H16 |
107.973 |
| N2 |
C7 |
C5 |
110.227 |
|
N2 |
C7 |
H17 |
108.020 |
| N2 |
C7 |
H18 |
107.973 |
|
N2 |
C8 |
C4 |
110.227 |
| N2 |
C8 |
H19 |
108.020 |
|
N2 |
C8 |
H20 |
107.973 |
| C3 |
N1 |
C4 |
108.704 |
|
C3 |
N1 |
C5 |
108.704 |
| C3 |
C6 |
H15 |
111.585 |
|
C3 |
C6 |
H16 |
111.566 |
| C4 |
N1 |
C5 |
108.704 |
|
C4 |
C8 |
H19 |
111.585 |
| C4 |
C8 |
H20 |
111.566 |
|
C5 |
C6 |
H15 |
101.870 |
| C5 |
C6 |
H16 |
150.793 |
|
C6 |
N2 |
C7 |
108.704 |
| C6 |
N2 |
C8 |
108.704 |
|
C6 |
C3 |
H9 |
111.585 |
| C6 |
C3 |
H10 |
111.566 |
|
C7 |
N2 |
C8 |
108.704 |
| C7 |
C5 |
H13 |
111.585 |
|
C7 |
C5 |
H14 |
111.566 |
| C8 |
C4 |
H11 |
111.585 |
|
C8 |
C4 |
H12 |
111.566 |
| H9 |
C3 |
H10 |
107.310 |
|
H11 |
C4 |
H12 |
107.310 |
| H13 |
C5 |
H14 |
107.310 |
|
H15 |
C6 |
H16 |
107.310 |
| H17 |
C7 |
H18 |
107.310 |
|
H19 |
C8 |
H20 |
107.310 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
