return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7NO (dimethylformamide)

using model chemistry: CCD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/6-31G**
 hartrees
Energy at 0K-247.806175
Energy at 298.15K-247.814527
HF Energy-247.000753
Nuclear repulsion energy181.648774
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3259 3056 1.15      
2 A' 3216 3016 19.58      
3 A' 3108 2914 49.80      
4 A' 3099 2906 46.96      
5 A' 3066 2875 84.22      
6 A' 1857 1741 456.30      
7 A' 1602 1502 20.58      
8 A' 1572 1474 12.65      
9 A' 1528 1433 6.87      
10 A' 1504 1410 70.49      
11 A' 1490 1397 3.04      
12 A' 1476 1383 77.81      
13 A' 1342 1258 41.56      
14 A' 1138 1067 118.42      
15 A' 1120 1050 2.54      
16 A' 907 851 3.42      
17 A' 677 635 7.43      
18 A' 401 376 1.31      
19 A' 333 312 12.27      
20 A" 3172 2974 21.81      
21 A" 3165 2967 50.16      
22 A" 1561 1464 11.61      
23 A" 1538 1442 2.99      
24 A" 1215 1139 5.35      
25 A" 1166 1093 0.01      
26 A" 1055 989 1.19      
27 A" 331 310 20.91      
28 A" 226 212 2.51      
29 A" 161 151 0.53      
30 A" 80 75 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 23181.3 cm-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 21734.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/6-31G**
ABC
0.29892 0.13958 0.09868

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.699 -0.822 0.000
O2 0.213 -1.939 0.000
N3 0.000 0.347 0.000
C4 -1.450 0.331 0.000
C5 0.660 1.634 0.000
H6 1.787 -0.653 0.000
H7 -1.777 -0.704 0.000
H8 -1.838 0.836 0.887
H9 -1.838 0.836 -0.887
H10 1.739 1.492 0.000
H11 0.387 2.212 -0.886
H12 0.387 2.212 0.886

Atom - Atom Distances (Å)
  C1 O2 N3 C4 C5 H6 H7 H8 H9 H10 H11 H12
C11.21881.36152.43882.45641.10062.47863.15763.15762.53693.17583.1758
O21.21882.29582.81403.60162.03262.34183.56303.56303.75574.24834.2483
N31.36152.29581.45051.44712.04732.06422.09862.09862.08252.10082.1008
C42.43882.81401.45052.48083.38321.08491.09191.09193.39452.77462.7746
C52.45643.60161.44712.48082.54973.37752.76842.76841.08831.09231.0923
H61.10062.03262.04733.38322.54973.56384.01764.01762.14573.30963.3096
H72.47862.34182.06421.08493.37753.56381.77831.77834.14553.73733.7373
H83.15763.56302.09861.09192.76844.01761.77831.77383.74333.15952.6154
H93.15763.56302.09861.09192.76844.01761.77831.77383.74332.61543.1595
H102.53693.75572.08253.39451.08832.14574.14553.74333.74331.76991.7699
H113.17584.24832.10082.77461.09233.30963.73733.15952.61541.76991.7716
H123.17584.24832.10082.77461.09233.30963.73732.61543.15951.76991.7716

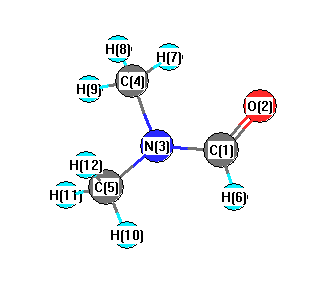
picture of dimethylformamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 C4 120.259 C1 N3 C5 121.965
O2 C1 N3 125.594 O2 C1 H6 122.328
N3 C1 H6 112.077 N3 C4 H7 108.143
N3 C4 H8 110.469 N3 C4 H9 110.469
N3 C5 H10 109.633 N3 C5 H11 110.866
N3 C5 H12 110.866 C4 N3 C5 117.776
H7 C4 H8 109.560 H7 C4 H9 109.560
H8 C4 H9 108.627 H10 C5 H11 108.517
H10 C5 H12 108.517 H11 C5 H12 108.373
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability