return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6 (Hexa-1,5-diene-3-yne)

using model chemistry: CCD/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at CCD/6-31+G**
 hartrees
Energy at 0K-231.444779
Energy at 298.15K-231.448356
HF Energy-230.615627
Nuclear repulsion energy174.440796
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3332 3140 0.00      
2 Ag 3243 3056 0.00      
3 Ag 3231 3045 0.00      
4 Ag 2346 2211 0.00      
5 Ag 1705 1607 0.00      
6 Ag 1481 1396 0.00      
7 Ag 1356 1278 0.00      
8 Ag 1114 1049 0.00      
9 Ag 729 687 0.00      
10 Ag 530 500 0.00      
11 Ag 226 213 0.00      
12 Au 1008 950 53.13      
13 Au 937 883 83.23      
14 Au 669 631 10.99      
15 Au 164 154 7.83      
16 Au 26 24 0.05      
17 Bg 1006 948 0.00      
18 Bg 937 883 0.00      
19 Bg 645 607 0.00      
20 Bg 221 208 0.00      
21 Bu 3332 3140 20.46      
22 Bu 3243 3056 20.80      
23 Bu 3231 3045 7.55      
24 Bu 1728 1629 36.08      
25 Bu 1499 1413 5.60      
26 Bu 1353 1275 4.84      
27 Bu 1232 1161 15.20      
28 Bu 1035 975 0.13      
29 Bu 534 503 8.13      
30 Bu 111 104 2.64      

Unscaled Zero Point Vibrational Energy (zpe) 21102.7 cm-1
Scaled (by 0.9423) Zero Point Vibrational Energy (zpe) 19885.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/6-31+G**
ABC
0.85834 0.04516 0.04290

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/6-31+G**

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.609 0.000
C2 0.000 -0.609 0.000
C3 0.000 2.045 0.000
C4 0.000 -2.045 0.000
C5 1.120 2.787 0.000
C6 -1.120 -2.787 0.000
H7 -0.970 2.530 0.000
H8 0.970 -2.530 0.000
H9 2.102 2.334 0.000
H10 1.066 3.867 0.000
H11 -2.102 -2.334 0.000
H12 -1.066 -3.867 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
C11.21751.43622.65372.44953.57592.15183.28482.71913.42823.61624.6009
C21.21752.65371.43623.57592.44953.28482.15183.61624.60092.71913.4282
C31.43622.65374.08981.34364.96031.08424.67622.12152.11104.85716.0072
C42.65371.43624.08984.96031.34364.67621.08424.85716.00722.12152.1110
C52.44953.57591.34364.96036.00772.10555.31901.08141.08106.05037.0041
C63.57592.44954.96031.34366.00775.31902.10556.05037.00411.08141.0810
H72.15183.28481.08424.67622.10555.31905.41823.07772.43594.99356.3972
H83.28482.15184.67621.08425.31902.10555.41824.99356.39723.07772.4359
H92.71913.61622.12154.85711.08146.05033.07774.99351.85006.28166.9632
H103.42824.60092.11106.00721.08107.00412.43596.39721.85006.96328.0224
H113.61622.71914.85712.12156.05031.08144.99353.07776.28166.96321.8500
H124.60093.42826.00722.11107.00411.08106.39722.43596.96328.02241.8500

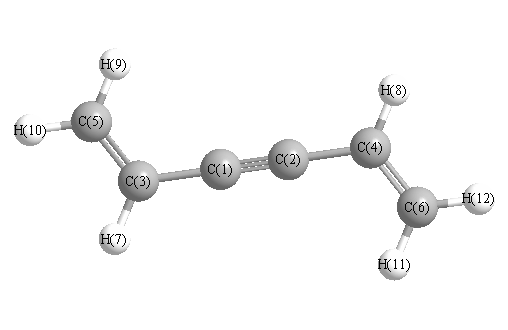
picture of Hexa-1,5-diene-3-yne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 180.000 C1 C3 C5 123.541
C1 C3 H7 116.553 C2 C1 C3 180.000
C2 C4 C6 123.541 C2 C4 H8 116.553
C3 C5 H9 121.676 C3 C5 H10 120.689
C4 C6 H11 121.676 C4 C6 H12 120.689
C5 C3 H7 119.906 C6 C4 H8 119.906
H9 C5 H10 117.635 H11 C6 H12 117.635
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability