All results from a given calculation for C6H12 (2,3-dimethylbut-1-ene)
using model chemistry: CCD/cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C2 |
1A' |
Energy calculated at CCD/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -235.354928 |
| Energy at 298.15K | |
| HF Energy | -234.260474 |
| Nuclear repulsion energy | 244.340897 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVTZ
Geometric Data calculated at CCD/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.947 |
-1.815 |
0.000 |
| C2 |
-0.064 |
-0.816 |
0.000 |
| C3 |
-0.523 |
0.628 |
0.000 |
| C4 |
1.418 |
-1.086 |
0.000 |
| C5 |
-0.064 |
1.372 |
1.258 |
| C6 |
-0.064 |
1.372 |
-1.258 |
| H7 |
-0.632 |
-2.849 |
0.000 |
| H8 |
-2.012 |
-1.624 |
0.000 |
| H9 |
-1.615 |
0.612 |
0.000 |
| H10 |
1.615 |
-2.156 |
0.000 |
| H11 |
1.900 |
-0.653 |
-0.878 |
| H12 |
1.900 |
-0.653 |
0.878 |
| H13 |
-0.487 |
2.377 |
1.280 |
| H14 |
-0.379 |
0.849 |
2.162 |
| H15 |
1.022 |
1.472 |
1.285 |
| H16 |
-0.487 |
2.377 |
-1.280 |
| H17 |
-0.379 |
0.849 |
-2.162 |
| H18 |
1.022 |
1.472 |
-1.285 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.3328 | 2.4789 | 2.4750 | 3.5379 | 3.5379 | 1.0818 | 1.0820 | 2.5168 | 2.5851 | 3.1982 | 3.1982 | 4.4067 | 3.4773 | 4.0413 | 4.4067 | 3.4773 | 4.0413 |
C2 | 1.3328 | | 1.5150 | 1.5067 | 2.5241 | 2.5241 | 2.1108 | 2.1089 | 2.1082 | 2.1486 | 2.1579 | 2.1579 | 3.4661 | 2.7470 | 2.8404 | 3.4661 | 2.7470 | 2.8404 | C3 | 2.4789 | 1.5150 | | 2.5892 | 1.5321 | 1.5321 | 3.4787 | 2.6997 | 1.0923 | 3.5103 | 2.8778 | 2.8778 | 2.1678 | 2.1776 | 2.1794 | 2.1678 | 2.1776 | 2.1794 | C4 | 2.4750 | 1.5067 | 2.5892 | | 3.1340 | 3.1340 | 2.7038 | 3.4723 | 3.4760 | 1.0884 | 1.0911 | 1.0911 | 4.1546 | 3.4131 | 2.8902 | 4.1546 | 3.4131 | 2.8902 | C5 | 3.5379 | 2.5241 | 1.5321 | 3.1340 | | 2.5164 | 4.4412 | 3.7886 | 2.1367 | 4.1052 | 3.5386 | 2.8468 | 1.0907 | 1.0901 | 1.0909 | 2.7622 | 3.4737 | 2.7669 | C6 | 3.5379 | 2.5241 | 1.5321 | 3.1340 | 2.5164 | | 4.4412 | 3.7886 | 2.1367 | 4.1052 | 2.8468 | 3.5386 | 2.7622 | 3.4737 | 2.7669 | 1.0907 | 1.0901 | 1.0909 | H7 | 1.0818 | 2.1108 | 3.4787 | 2.7038 | 4.4412 | 4.4412 | | 1.8461 | 3.5983 | 2.3512 | 3.4648 | 3.4648 | 5.3827 | 4.2914 | 4.8023 | 5.3827 | 4.2914 | 4.8023 | H8 | 1.0820 | 2.1089 | 2.6997 | 3.4723 | 3.7886 | 3.7886 | 1.8461 | | 2.2710 | 3.6664 | 4.1257 | 4.1257 | 4.4689 | 3.6682 | 4.5214 | 4.4689 | 3.6682 | 4.5214 | H9 | 2.5168 | 2.1082 | 1.0923 | 3.4760 | 2.1367 | 2.1367 | 3.5983 | 2.2710 | | 4.2542 | 3.8376 | 3.8376 | 2.4545 | 2.5010 | 3.0566 | 2.4545 | 2.5010 | 3.0566 | H10 | 2.5851 | 2.1486 | 3.5103 | 1.0884 | 4.1052 | 4.1052 | 2.3512 | 3.6664 | 4.2542 | | 1.7642 | 1.7642 | 5.1585 | 4.2054 | 3.8950 | 5.1585 | 4.2054 | 3.8950 | H11 | 3.1982 | 2.1579 | 2.8778 | 1.0911 | 3.5386 | 2.8468 | 3.4648 | 4.1257 | 3.8376 | 1.7642 | | 1.7556 | 4.4199 | 4.0855 | 3.1568 | 3.8785 | 3.0169 | 2.3355 | H12 | 3.1982 | 2.1579 | 2.8778 | 1.0911 | 2.8468 | 3.5386 | 3.4648 | 4.1257 | 3.8376 | 1.7642 | 1.7556 | | 3.8785 | 3.0169 | 2.3355 | 4.4199 | 4.0855 | 3.1568 | H13 | 4.4067 | 3.4661 | 2.1678 | 4.1546 | 1.0907 | 2.7622 | 5.3827 | 4.4689 | 2.4545 | 5.1585 | 4.4199 | 3.8785 | | 1.7672 | 1.7593 | 2.5594 | 3.7666 | 3.1099 | H14 | 3.4773 | 2.7470 | 2.1776 | 3.4131 | 1.0901 | 3.4737 | 4.2914 | 3.6682 | 2.5010 | 4.2054 | 4.0855 | 3.0169 | 1.7672 | | 1.7664 | 3.7666 | 4.3231 | 3.7720 | H15 | 4.0413 | 2.8404 | 2.1794 | 2.8902 | 1.0909 | 2.7669 | 4.8023 | 4.5214 | 3.0566 | 3.8950 | 3.1568 | 2.3355 | 1.7593 | 1.7664 | | 3.1099 | 3.7720 | 2.5695 | H16 | 4.4067 | 3.4661 | 2.1678 | 4.1546 | 2.7622 | 1.0907 | 5.3827 | 4.4689 | 2.4545 | 5.1585 | 3.8785 | 4.4199 | 2.5594 | 3.7666 | 3.1099 | | 1.7672 | 1.7593 | H17 | 3.4773 | 2.7470 | 2.1776 | 3.4131 | 3.4737 | 1.0901 | 4.2914 | 3.6682 | 2.5010 | 4.2054 | 3.0169 | 4.0855 | 3.7666 | 4.3231 | 3.7720 | 1.7672 | | 1.7664 | H18 | 4.0413 | 2.8404 | 2.1794 | 2.8902 | 2.7669 | 1.0909 | 4.8023 | 4.5214 | 3.0566 | 3.8950 | 2.3355 | 3.1568 | 3.1099 | 3.7720 | 2.5695 | 1.7593 | 1.7664 | |
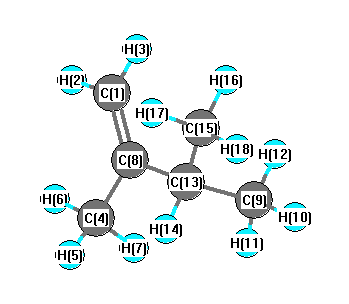 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.887 |
|
C1 |
C2 |
C4 |
121.181 |
| C2 |
C1 |
H7 |
121.540 |
|
C2 |
C1 |
H8 |
121.344 |
| C2 |
C3 |
C5 |
111.863 |
|
C2 |
C3 |
C6 |
111.863 |
| C2 |
C3 |
H9 |
106.798 |
|
C2 |
C4 |
H10 |
110.744 |
| C2 |
C4 |
H11 |
111.328 |
|
C2 |
C4 |
H12 |
111.328 |
| C3 |
C2 |
C4 |
117.932 |
|
C3 |
C5 |
H13 |
110.350 |
| C3 |
C5 |
H14 |
111.168 |
|
C3 |
C5 |
H15 |
111.267 |
| C3 |
C6 |
H16 |
110.350 |
|
C3 |
C6 |
H17 |
111.168 |
| C3 |
C6 |
H18 |
111.267 |
|
C5 |
C3 |
C6 |
110.415 |
| C5 |
C3 |
H9 |
107.834 |
|
C6 |
C3 |
H9 |
107.834 |
| H7 |
C1 |
H8 |
117.116 |
|
H10 |
C4 |
H11 |
108.079 |
| H10 |
C4 |
H12 |
108.079 |
|
H11 |
C4 |
H12 |
107.123 |
| H13 |
C5 |
H14 |
108.256 |
|
H13 |
C5 |
H15 |
107.499 |
| H14 |
C5 |
H15 |
108.168 |
|
H16 |
C6 |
H17 |
108.256 |
| H16 |
C6 |
H18 |
107.499 |
|
H17 |
C6 |
H18 |
108.168 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
