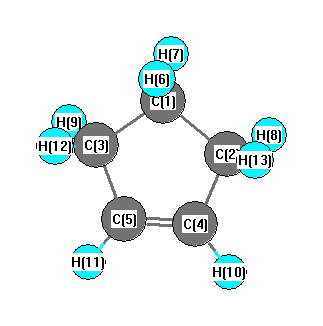
Vibrational Frequencies calculated at wB97X-D/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3242 |
3075 |
26.72 |
|
|
|
| 2 |
A' |
3152 |
2990 |
42.89 |
|
|
|
| 3 |
A' |
3113 |
2952 |
51.91 |
|
|
|
| 4 |
A' |
3091 |
2932 |
13.29 |
|
|
|
| 5 |
A' |
3054 |
2897 |
26.90 |
|
|
|
| 6 |
A' |
1735 |
1646 |
1.47 |
|
|
|
| 7 |
A' |
1550 |
1470 |
1.27 |
|
|
|
| 8 |
A' |
1521 |
1443 |
1.29 |
|
|
|
| 9 |
A' |
1360 |
1290 |
0.63 |
|
|
|
| 10 |
A' |
1253 |
1189 |
1.32 |
|
|
|
| 11 |
A' |
1139 |
1080 |
0.15 |
|
|
|
| 12 |
A' |
1091 |
1035 |
5.82 |
|
|
|
| 13 |
A' |
993 |
942 |
1.07 |
|
|
|
| 14 |
A' |
925 |
878 |
0.53 |
|
|
|
| 15 |
A' |
842 |
799 |
0.71 |
|
|
|
| 16 |
A' |
727 |
689 |
31.02 |
|
|
|
| 17 |
A' |
621 |
589 |
10.83 |
|
|
|
| 18 |
A' |
158 |
149 |
0.21 |
|
|
|
| 19 |
A" |
3216 |
3050 |
8.66 |
|
|
|
| 20 |
A" |
3107 |
2947 |
26.66 |
|
|
|
| 21 |
A" |
3055 |
2898 |
61.95 |
|
|
|
| 22 |
A" |
1528 |
1449 |
0.99 |
|
|
|
| 23 |
A" |
1402 |
1330 |
1.61 |
|
|
|
| 24 |
A" |
1345 |
1276 |
0.73 |
|
|
|
| 25 |
A" |
1325 |
1257 |
1.58 |
|
|
|
| 26 |
A" |
1250 |
1185 |
0.00 |
|
|
|
| 27 |
A" |
1173 |
1112 |
1.58 |
|
|
|
| 28 |
A" |
1062 |
1008 |
1.45 |
|
|
|
| 29 |
A" |
990 |
939 |
0.09 |
|
|
|
| 30 |
A" |
935 |
887 |
6.75 |
|
|
|
| 31 |
A" |
912 |
865 |
3.03 |
|
|
|
| 32 |
A" |
786 |
745 |
0.23 |
|
|
|
| 33 |
A" |
395 |
374 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26023.0 cm
-1
Scaled (by 0.9485) Zero Point Vibrational Energy (zpe) 24682.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.326 |
|
|
|
| 2 |
C |
-0.345 |
|
|
|
| 3 |
C |
-0.345 |
|
|
|
| 4 |
C |
-0.141 |
|
|
|
| 5 |
C |
-0.141 |
|
|
|
| 6 |
H |
0.166 |
|
|
|
| 7 |
H |
0.160 |
|
|
|
| 8 |
H |
0.166 |
|
|
|
| 9 |
H |
0.166 |
|
|
|
| 10 |
H |
0.157 |
|
|
|
| 11 |
H |
0.157 |
|
|
|
| 12 |
H |
0.163 |
|
|
|
| 13 |
H |
0.163 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.046 |
-0.144 |
0.000 |
0.151 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.000 |
-0.020 |
0.000 |
| y |
-0.020 |
-30.566 |
0.000 |
| z |
0.000 |
0.000 |
-30.027 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.703 |
-0.020 |
0.000 |
| y |
-0.020 |
0.447 |
0.000 |
| z |
0.000 |
0.000 |
1.256 |
|
| Polar |
|---|
| 3z2-r2 | 2.512 |
|---|
| x2-y2 | -1.434 |
|---|
| xy | -0.020 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.384 |
-0.063 |
0.000 |
| y |
-0.063 |
7.340 |
0.000 |
| z |
0.000 |
0.000 |
8.793 |
<r2> (average value of r
2) Å
2
| <r2> |
104.587 |
| (<r2>)1/2 |
10.227 |