Jump to
S1C2
Energy calculated at wB97X-D/6-31G*
| | hartrees |
|---|
| Energy at 0K | -491.574318 |
| Energy at 298.15K | -491.575036 |
| HF Energy | -491.574318 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3756 |
3563 |
272.21 |
|
|
|
| 2 |
A' |
2104 |
1996 |
741.31 |
|
|
|
| 3 |
A' |
887 |
841 |
5.77 |
|
|
|
| 4 |
A' |
620 |
588 |
367.50 |
|
|
|
| 5 |
A' |
447 |
424 |
112.17 |
|
|
|
| 6 |
A" |
485 |
460 |
0.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4149.6 cm
-1
Scaled (by 0.9485) Zero Point Vibrational Energy (zpe) 3935.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-0.143 |
1.691 |
0.000 |
| C2 |
0.000 |
0.497 |
0.000 |
| S3 |
0.031 |
-1.080 |
0.000 |
| H4 |
0.505 |
2.459 |
0.000 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
S3 |
H4 |
| N1 | | 1.2017 | 2.7758 | 1.0055 |
C2 | 1.2017 | | 1.5774 | 2.0260 | S3 | 2.7758 | 1.5774 | | 3.5707 | H4 | 1.0055 | 2.0260 | 3.5707 | |
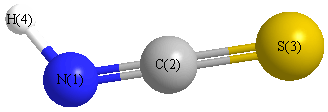 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
S3 |
174.293 |
|
C2 |
N1 |
H4 |
133.033 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.606 |
|
|
|
| 2 |
C |
0.296 |
|
|
|
| 3 |
S |
-0.086 |
|
|
|
| 4 |
H |
0.396 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.252 |
2.090 |
0.000 |
2.436 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.155 |
3.251 |
0.000 |
| y |
3.251 |
-19.019 |
0.000 |
| z |
0.000 |
0.000 |
-24.976 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.158 |
3.251 |
0.000 |
| y |
3.251 |
5.547 |
0.000 |
| z |
0.000 |
0.000 |
-3.389 |
|
| Polar |
|---|
| 3z2-r2 | -6.778 |
|---|
| x2-y2 | -5.137 |
|---|
| xy | 3.251 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.458 |
-0.203 |
0.000 |
| y |
-0.203 |
8.750 |
0.000 |
| z |
0.000 |
0.000 |
2.293 |
<r2> (average value of r
2) Å
2
| <r2> |
60.795 |
| (<r2>)1/2 |
7.797 |
Jump to
S1C1
Energy calculated at wB97X-D/6-31G*
| | hartrees |
|---|
| Energy at 0K | -491.571024 |
| Energy at 298.15K | |
| HF Energy | -491.571024 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3949 |
3745 |
713.16 |
|
|
|
| 2 |
Σ |
2218 |
2104 |
726.35 |
|
|
|
| 3 |
Σ |
860 |
816 |
39.26 |
|
|
|
| 4 |
Π |
481 |
456 |
3.08 |
|
|
|
| 4 |
Π |
481 |
456 |
3.08 |
|
|
|
| 5 |
Π |
438i |
415i |
189.46 |
|
|
|
| 5 |
Π |
438i |
415i |
189.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3556.4 cm
-1
Scaled (by 0.9485) Zero Point Vibrational Energy (zpe) 3373.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
-1.679 |
| C2 |
0.000 |
0.000 |
-0.501 |
| S3 |
0.000 |
0.000 |
1.089 |
| H4 |
0.000 |
0.000 |
-2.672 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
S3 |
H4 |
| N1 | | 1.1783 | 2.7681 | 0.9927 |
C2 | 1.1783 | | 1.5897 | 2.1710 | S3 | 2.7681 | 1.5897 | | 3.7608 | H4 | 0.9927 | 2.1710 | 3.7608 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
S3 |
180.000 |
|
C2 |
N1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.593 |
|
|
|
| 2 |
C |
0.325 |
|
|
|
| 3 |
S |
-0.131 |
|
|
|
| 4 |
H |
0.399 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.437 |
3.437 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.925 |
0.000 |
0.000 |
| y |
0.000 |
-24.925 |
0.000 |
| z |
0.000 |
0.000 |
-15.569 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.678 |
0.000 |
0.000 |
| y |
0.000 |
-4.678 |
0.000 |
| z |
0.000 |
0.000 |
9.357 |
|
| Polar |
|---|
| 3z2-r2 | 18.714 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.288 |
0.000 |
0.000 |
| y |
0.000 |
2.288 |
0.000 |
| z |
0.000 |
0.000 |
8.496 |
<r2> (average value of r
2) Å
2
| <r2> |
60.973 |
| (<r2>)1/2 |
7.809 |
