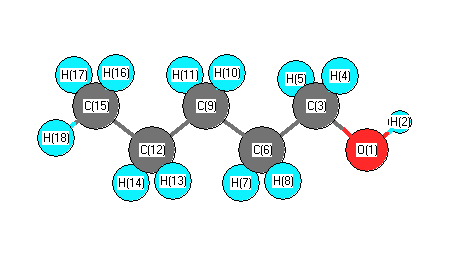
Vibrational Frequencies calculated at wB97X-D/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3859 |
3660 |
19.94 |
|
|
|
| 2 |
A' |
3150 |
2988 |
39.30 |
|
|
|
| 3 |
A' |
3074 |
2916 |
45.16 |
|
|
|
| 4 |
A' |
3068 |
2910 |
33.35 |
|
|
|
| 5 |
A' |
3060 |
2902 |
29.57 |
|
|
|
| 6 |
A' |
3042 |
2885 |
21.24 |
|
|
|
| 7 |
A' |
2996 |
2842 |
58.05 |
|
|
|
| 8 |
A' |
1567 |
1487 |
3.85 |
|
|
|
| 9 |
A' |
1547 |
1468 |
6.39 |
|
|
|
| 10 |
A' |
1539 |
1460 |
0.64 |
|
|
|
| 11 |
A' |
1529 |
1451 |
0.67 |
|
|
|
| 12 |
A' |
1526 |
1448 |
0.11 |
|
|
|
| 13 |
A' |
1492 |
1415 |
7.84 |
|
|
|
| 14 |
A' |
1450 |
1375 |
3.41 |
|
|
|
| 15 |
A' |
1437 |
1363 |
0.68 |
|
|
|
| 16 |
A' |
1382 |
1311 |
13.53 |
|
|
|
| 17 |
A' |
1320 |
1252 |
36.96 |
|
|
|
| 18 |
A' |
1259 |
1194 |
27.31 |
|
|
|
| 19 |
A' |
1151 |
1092 |
9.53 |
|
|
|
| 20 |
A' |
1114 |
1057 |
78.82 |
|
|
|
| 21 |
A' |
1094 |
1038 |
0.22 |
|
|
|
| 22 |
A' |
1064 |
1009 |
0.58 |
|
|
|
| 23 |
A' |
1025 |
973 |
15.59 |
|
|
|
| 24 |
A' |
918 |
871 |
3.48 |
|
|
|
| 25 |
A' |
506 |
480 |
12.35 |
|
|
|
| 26 |
A' |
371 |
352 |
0.09 |
|
|
|
| 27 |
A' |
320 |
304 |
4.83 |
|
|
|
| 28 |
A' |
136 |
129 |
1.60 |
|
|
|
| 29 |
A" |
3143 |
2981 |
67.62 |
|
|
|
| 30 |
A" |
3125 |
2964 |
56.74 |
|
|
|
| 31 |
A" |
3101 |
2942 |
13.83 |
|
|
|
| 32 |
A" |
3075 |
2916 |
5.14 |
|
|
|
| 33 |
A" |
3028 |
2872 |
63.59 |
|
|
|
| 34 |
A" |
1534 |
1455 |
7.75 |
|
|
|
| 35 |
A" |
1355 |
1285 |
0.90 |
|
|
|
| 36 |
A" |
1342 |
1273 |
0.94 |
|
|
|
| 37 |
A" |
1321 |
1253 |
0.04 |
|
|
|
| 38 |
A" |
1258 |
1193 |
0.58 |
|
|
|
| 39 |
A" |
1212 |
1149 |
1.35 |
|
|
|
| 40 |
A" |
1007 |
955 |
1.46 |
|
|
|
| 41 |
A" |
881 |
835 |
0.01 |
|
|
|
| 42 |
A" |
790 |
750 |
0.50 |
|
|
|
| 43 |
A" |
750 |
711 |
3.15 |
|
|
|
| 44 |
A" |
279 |
265 |
129.92 |
|
|
|
| 45 |
A" |
243 |
231 |
0.82 |
|
|
|
| 46 |
A" |
155 |
147 |
1.74 |
|
|
|
| 47 |
A" |
85 |
80 |
6.25 |
|
|
|
| 48 |
A" |
75 |
71 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36878.0 cm
-1
Scaled (by 0.9485) Zero Point Vibrational Energy (zpe) 34978.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.640 |
|
|
|
| 2 |
H |
0.407 |
|
|
|
| 3 |
C |
-0.055 |
|
|
|
| 4 |
H |
0.132 |
|
|
|
| 5 |
H |
0.132 |
|
|
|
| 6 |
C |
-0.308 |
|
|
|
| 7 |
H |
0.165 |
|
|
|
| 8 |
H |
0.165 |
|
|
|
| 9 |
C |
-0.299 |
|
|
|
| 10 |
H |
0.147 |
|
|
|
| 11 |
H |
0.147 |
|
|
|
| 12 |
C |
-0.292 |
|
|
|
| 13 |
H |
0.154 |
|
|
|
| 14 |
H |
0.154 |
|
|
|
| 15 |
C |
-0.495 |
|
|
|
| 16 |
H |
0.160 |
|
|
|
| 17 |
H |
0.160 |
|
|
|
| 18 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.312 |
0.743 |
0.000 |
1.507 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.079 |
-4.129 |
0.000 |
| y |
-4.129 |
-42.567 |
0.000 |
| z |
0.000 |
0.000 |
-39.167 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.788 |
-4.129 |
0.000 |
| y |
-4.129 |
-5.944 |
0.000 |
| z |
0.000 |
0.000 |
-0.845 |
|
| Polar |
|---|
| 3z2-r2 | -1.689 |
|---|
| x2-y2 | 8.488 |
|---|
| xy | -4.129 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.596 |
-1.030 |
0.000 |
| y |
-1.030 |
9.488 |
0.000 |
| z |
0.000 |
0.000 |
7.588 |
<r2> (average value of r
2) Å
2
| <r2> |
300.677 |
| (<r2>)1/2 |
17.340 |