Jump to
S1C2
Vibrational Frequencies calculated at wB97X-D/6-31G(2df,p)
Geometric Data calculated at wB97X-D/6-31G(2df,p)
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at wB97X-D/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -194.309886 |
| Energy at 298.15K | |
| HF Energy | -194.309886 |
| Nuclear repulsion energy | 132.967683 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3940 |
3940 |
29.72 |
95.16 |
0.29 |
0.45 |
| 2 |
A |
3155 |
3155 |
23.00 |
38.33 |
0.75 |
0.86 |
| 3 |
A |
3130 |
3130 |
43.70 |
50.02 |
0.59 |
0.74 |
| 4 |
A |
3102 |
3102 |
18.16 |
86.89 |
0.72 |
0.84 |
| 5 |
A |
3058 |
3058 |
37.37 |
110.74 |
0.07 |
0.13 |
| 6 |
A |
3053 |
3053 |
29.36 |
88.60 |
0.03 |
0.06 |
| 7 |
A |
3014 |
3014 |
56.98 |
79.98 |
0.74 |
0.85 |
| 8 |
A |
2977 |
2977 |
64.77 |
96.06 |
0.11 |
0.20 |
| 9 |
A |
1531 |
1531 |
3.44 |
5.32 |
0.67 |
0.80 |
| 10 |
A |
1515 |
1515 |
6.72 |
7.60 |
0.75 |
0.86 |
| 11 |
A |
1500 |
1500 |
6.44 |
17.97 |
0.75 |
0.86 |
| 12 |
A |
1485 |
1485 |
2.35 |
14.81 |
0.75 |
0.86 |
| 13 |
A |
1470 |
1470 |
4.79 |
1.54 |
0.45 |
0.62 |
| 14 |
A |
1417 |
1417 |
4.12 |
2.43 |
0.75 |
0.86 |
| 15 |
A |
1383 |
1383 |
1.17 |
1.18 |
0.75 |
0.86 |
| 16 |
A |
1335 |
1335 |
26.25 |
12.87 |
0.74 |
0.85 |
| 17 |
A |
1283 |
1283 |
2.63 |
8.39 |
0.74 |
0.85 |
| 18 |
A |
1264 |
1264 |
35.92 |
3.92 |
0.66 |
0.80 |
| 19 |
A |
1172 |
1172 |
7.26 |
1.15 |
0.24 |
0.39 |
| 20 |
A |
1152 |
1152 |
20.78 |
3.21 |
0.54 |
0.70 |
| 21 |
A |
1083 |
1083 |
36.68 |
2.72 |
0.69 |
0.81 |
| 22 |
A |
1008 |
1008 |
33.33 |
4.22 |
0.74 |
0.85 |
| 23 |
A |
934 |
934 |
2.25 |
0.36 |
0.68 |
0.81 |
| 24 |
A |
880 |
880 |
2.06 |
10.16 |
0.16 |
0.27 |
| 25 |
A |
780 |
780 |
0.53 |
0.50 |
0.48 |
0.65 |
| 26 |
A |
485 |
485 |
7.16 |
0.22 |
0.52 |
0.69 |
| 27 |
A |
331 |
331 |
7.26 |
0.26 |
0.43 |
0.60 |
| 28 |
A |
257 |
257 |
103.71 |
3.02 |
0.74 |
0.85 |
| 29 |
A |
225 |
225 |
2.39 |
0.08 |
0.69 |
0.82 |
| 30 |
A |
147 |
147 |
6.44 |
0.08 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 24031.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 24031.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.533 |
-0.516 |
0.128 |
| C2 |
-0.633 |
0.646 |
-0.286 |
| C3 |
0.770 |
0.539 |
0.290 |
| O4 |
1.372 |
-0.633 |
-0.221 |
| H5 |
-2.527 |
-0.424 |
-0.317 |
| H6 |
-1.652 |
-0.549 |
1.216 |
| H7 |
-1.099 |
-1.467 |
-0.191 |
| H8 |
-0.554 |
0.689 |
-1.378 |
| H9 |
-1.066 |
1.598 |
0.041 |
| H10 |
0.711 |
0.506 |
1.391 |
| H11 |
1.352 |
1.433 |
0.018 |
| H12 |
2.227 |
-0.738 |
0.194 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5272 | 2.5385 | 2.9281 | 1.0934 | 1.0956 | 1.0924 | 2.1628 | 2.1673 | 2.7703 | 3.4834 | 3.7673 |
C2 | 1.5272 | | 1.5208 | 2.3789 | 2.1761 | 2.1738 | 2.1660 | 1.0958 | 1.0960 | 2.1535 | 2.1566 | 3.2135 | C3 | 2.5385 | 1.5208 | | 1.4125 | 3.4887 | 2.8126 | 2.7838 | 2.1357 | 2.1347 | 1.1026 | 1.1011 | 1.9395 | O4 | 2.9281 | 2.3789 | 1.4125 | | 3.9064 | 3.3496 | 2.6085 | 2.6070 | 3.3151 | 2.0808 | 2.0798 | 0.9563 | H5 | 1.0934 | 2.1761 | 3.4887 | 3.9064 | | 1.7696 | 1.7731 | 2.5017 | 2.5205 | 3.7773 | 4.3139 | 4.7925 | H6 | 1.0956 | 2.1738 | 2.8126 | 3.3496 | 1.7696 | | 1.7688 | 3.0773 | 2.5173 | 2.5940 | 3.7935 | 4.0167 | H7 | 1.0924 | 2.1660 | 2.7838 | 2.6085 | 1.7731 | 1.7688 | | 2.5206 | 3.0742 | 3.1097 | 3.8030 | 3.4273 | H8 | 2.1628 | 1.0958 | 2.1357 | 2.6070 | 2.5017 | 3.0773 | 2.5206 | | 1.7615 | 3.0499 | 2.4773 | 3.4992 | H9 | 2.1673 | 1.0960 | 2.1347 | 3.3151 | 2.5205 | 2.5173 | 3.0742 | 1.7615 | | 2.4848 | 2.4235 | 4.0406 | H10 | 2.7703 | 2.1535 | 1.1026 | 2.0808 | 3.7773 | 2.5940 | 3.1097 | 3.0499 | 2.4848 | | 1.7763 | 2.2973 | H11 | 3.4834 | 2.1566 | 1.1011 | 2.0798 | 4.3139 | 3.7935 | 3.8030 | 2.4773 | 2.4235 | 1.7763 | | 2.3476 | H12 | 3.7673 | 3.2135 | 1.9395 | 0.9563 | 4.7925 | 4.0167 | 3.4273 | 3.4992 | 4.0406 | 2.2973 | 2.3476 | |
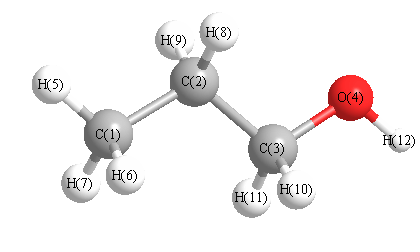 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.783 |
|
C1 |
C2 |
H8 |
109.996 |
| C1 |
C2 |
H9 |
110.338 |
|
C2 |
C1 |
H5 |
111.195 |
| C2 |
C1 |
H6 |
110.878 |
|
C2 |
C1 |
H7 |
110.450 |
| C2 |
C3 |
O4 |
108.330 |
|
C2 |
C3 |
H10 |
109.319 |
| C2 |
C3 |
H11 |
109.641 |
|
C3 |
C2 |
H8 |
108.322 |
| C3 |
C2 |
H9 |
108.237 |
|
C3 |
O4 |
H12 |
108.375 |
| O4 |
C3 |
H10 |
111.050 |
|
O4 |
C3 |
H11 |
111.056 |
| H5 |
C1 |
H6 |
107.882 |
|
H5 |
C1 |
H7 |
108.426 |
| H6 |
C1 |
H7 |
107.892 |
|
H8 |
C2 |
H9 |
106.968 |
| H10 |
C3 |
H11 |
107.428 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.416 |
|
|
|
| 2 |
C |
-0.216 |
|
|
|
| 3 |
C |
-0.079 |
|
|
|
| 4 |
O |
-0.448 |
|
|
|
| 5 |
H |
0.129 |
|
|
|
| 6 |
H |
0.125 |
|
|
|
| 7 |
H |
0.154 |
|
|
|
| 8 |
H |
0.129 |
|
|
|
| 9 |
H |
0.115 |
|
|
|
| 10 |
H |
0.098 |
|
|
|
| 11 |
H |
0.102 |
|
|
|
| 12 |
H |
0.307 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.726 |
0.770 |
0.932 |
1.410 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.492 |
-0.343 |
1.935 |
| y |
-0.343 |
-26.552 |
-0.562 |
| z |
1.935 |
-0.562 |
-26.240 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.905 |
-0.343 |
1.935 |
| y |
-0.343 |
-2.187 |
-0.562 |
| z |
1.935 |
-0.562 |
-1.718 |
|
| Polar |
|---|
| 3z2-r2 | -3.436 |
|---|
| x2-y2 | 4.061 |
|---|
| xy | -0.343 |
|---|
| xz | 1.935 |
|---|
| yz | -0.562 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.227 |
0.028 |
0.109 |
| y |
0.028 |
5.533 |
-0.001 |
| z |
0.109 |
-0.001 |
5.254 |
<r2> (average value of r
2) Å
2
| <r2> |
94.398 |
| (<r2>)1/2 |
9.716 |
