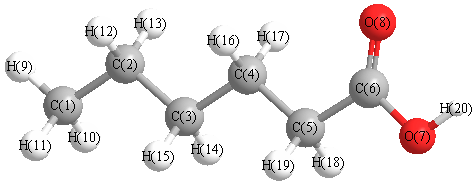
Vibrational Frequencies calculated at wB97X-D/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3847 |
3665 |
75.39 |
|
|
|
| 2 |
A |
3143 |
2994 |
39.47 |
|
|
|
| 3 |
A |
3132 |
2983 |
75.73 |
|
|
|
| 4 |
A |
3126 |
2978 |
24.58 |
|
|
|
| 5 |
A |
3105 |
2957 |
1.97 |
|
|
|
| 6 |
A |
3092 |
2945 |
21.07 |
|
|
|
| 7 |
A |
3074 |
2928 |
37.27 |
|
|
|
| 8 |
A |
3066 |
2921 |
2.30 |
|
|
|
| 9 |
A |
3065 |
2920 |
7.07 |
|
|
|
| 10 |
A |
3055 |
2910 |
37.39 |
|
|
|
| 11 |
A |
3049 |
2905 |
39.22 |
|
|
|
| 12 |
A |
3030 |
2887 |
21.11 |
|
|
|
| 13 |
A |
1862 |
1774 |
317.88 |
|
|
|
| 14 |
A |
1512 |
1440 |
9.81 |
|
|
|
| 15 |
A |
1511 |
1439 |
10.50 |
|
|
|
| 16 |
A |
1502 |
1431 |
4.81 |
|
|
|
| 17 |
A |
1494 |
1423 |
0.50 |
|
|
|
| 18 |
A |
1492 |
1421 |
0.36 |
|
|
|
| 19 |
A |
1465 |
1396 |
18.94 |
|
|
|
| 20 |
A |
1453 |
1384 |
64.52 |
|
|
|
| 21 |
A |
1429 |
1361 |
4.05 |
|
|
|
| 22 |
A |
1422 |
1355 |
16.92 |
|
|
|
| 23 |
A |
1374 |
1309 |
37.95 |
|
|
|
| 24 |
A |
1346 |
1283 |
1.04 |
|
|
|
| 25 |
A |
1339 |
1275 |
0.05 |
|
|
|
| 26 |
A |
1320 |
1257 |
5.25 |
|
|
|
| 27 |
A |
1304 |
1242 |
0.03 |
|
|
|
| 28 |
A |
1272 |
1211 |
14.33 |
|
|
|
| 29 |
A |
1232 |
1174 |
0.07 |
|
|
|
| 30 |
A |
1187 |
1130 |
237.34 |
|
|
|
| 31 |
A |
1145 |
1091 |
1.06 |
|
|
|
| 32 |
A |
1141 |
1087 |
33.43 |
|
|
|
| 33 |
A |
1089 |
1037 |
4.16 |
|
|
|
| 34 |
A |
1083 |
1031 |
6.19 |
|
|
|
| 35 |
A |
1038 |
989 |
9.14 |
|
|
|
| 36 |
A |
984 |
938 |
0.48 |
|
|
|
| 37 |
A |
932 |
888 |
5.33 |
|
|
|
| 38 |
A |
884 |
842 |
2.14 |
|
|
|
| 39 |
A |
863 |
822 |
6.59 |
|
|
|
| 40 |
A |
767 |
731 |
4.59 |
|
|
|
| 41 |
A |
726 |
692 |
9.85 |
|
|
|
| 42 |
A |
661 |
630 |
94.32 |
|
|
|
| 43 |
A |
642 |
612 |
28.62 |
|
|
|
| 44 |
A |
524 |
499 |
25.28 |
|
|
|
| 45 |
A |
516 |
492 |
23.03 |
|
|
|
| 46 |
A |
413 |
394 |
1.13 |
|
|
|
| 47 |
A |
310 |
296 |
1.79 |
|
|
|
| 48 |
A |
234 |
223 |
0.03 |
|
|
|
| 49 |
A |
227 |
216 |
2.17 |
|
|
|
| 50 |
A |
132 |
126 |
0.04 |
|
|
|
| 51 |
A |
98 |
93 |
1.05 |
|
|
|
| 52 |
A |
92 |
88 |
0.02 |
|
|
|
| 53 |
A |
53 |
50 |
0.09 |
|
|
|
| 54 |
A |
57i |
54i |
0.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38897.0 cm
-1
Scaled (by 0.9526) Zero Point Vibrational Energy (zpe) 37053.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.611 |
|
|
|
| 2 |
C |
-0.144 |
|
|
|
| 3 |
C |
-0.350 |
|
|
|
| 4 |
C |
-0.218 |
|
|
|
| 5 |
C |
-0.503 |
|
|
|
| 6 |
C |
0.616 |
|
|
|
| 7 |
O |
-0.462 |
|
|
|
| 8 |
O |
-0.477 |
|
|
|
| 9 |
H |
0.158 |
|
|
|
| 10 |
H |
0.154 |
|
|
|
| 11 |
H |
0.154 |
|
|
|
| 12 |
H |
0.151 |
|
|
|
| 13 |
H |
0.151 |
|
|
|
| 14 |
H |
0.145 |
|
|
|
| 15 |
H |
0.145 |
|
|
|
| 16 |
H |
0.167 |
|
|
|
| 17 |
H |
0.167 |
|
|
|
| 18 |
H |
0.189 |
|
|
|
| 19 |
H |
0.189 |
|
|
|
| 20 |
H |
0.380 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.451 |
-1.536 |
0.000 |
1.600 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.570 |
3.296 |
0.000 |
| y |
3.296 |
-55.971 |
0.001 |
| z |
0.000 |
0.001 |
-49.248 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.040 |
3.296 |
0.000 |
| y |
3.296 |
-8.062 |
0.001 |
| z |
0.000 |
0.001 |
2.022 |
|
| Polar |
|---|
| 3z2-r2 | 4.045 |
|---|
| x2-y2 | 9.402 |
|---|
| xy | 3.296 |
|---|
| xz | 0.000 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.655 |
0.005 |
0.000 |
| y |
0.005 |
11.281 |
-0.000 |
| z |
0.000 |
-0.000 |
9.183 |
<r2> (average value of r
2) Å
2
| <r2> |
510.538 |
| (<r2>)1/2 |
22.595 |