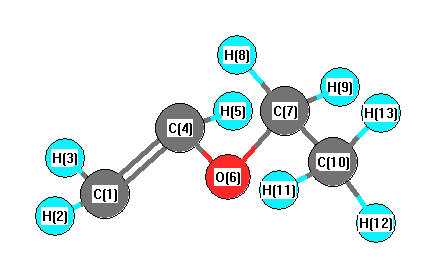
Vibrational Frequencies calculated at wB97X-D/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3282 |
3138 |
5.36 |
|
|
|
| 2 |
A |
3187 |
3047 |
0.06 |
|
|
|
| 3 |
A |
3152 |
3013 |
24.67 |
|
|
|
| 4 |
A |
3147 |
3009 |
31.35 |
|
|
|
| 5 |
A |
3146 |
3007 |
11.15 |
|
|
|
| 6 |
A |
3069 |
2934 |
16.01 |
|
|
|
| 7 |
A |
3057 |
2923 |
37.35 |
|
|
|
| 8 |
A |
3019 |
2886 |
53.02 |
|
|
|
| 9 |
A |
1740 |
1663 |
187.50 |
|
|
|
| 10 |
A |
1541 |
1473 |
7.01 |
|
|
|
| 11 |
A |
1518 |
1451 |
2.42 |
|
|
|
| 12 |
A |
1490 |
1425 |
6.51 |
|
|
|
| 13 |
A |
1453 |
1389 |
5.86 |
|
|
|
| 14 |
A |
1425 |
1363 |
63.08 |
|
|
|
| 15 |
A |
1402 |
1340 |
23.49 |
|
|
|
| 16 |
A |
1351 |
1291 |
5.35 |
|
|
|
| 17 |
A |
1315 |
1257 |
0.78 |
|
|
|
| 18 |
A |
1249 |
1194 |
261.10 |
|
|
|
| 19 |
A |
1195 |
1142 |
12.13 |
|
|
|
| 20 |
A |
1172 |
1121 |
121.88 |
|
|
|
| 21 |
A |
1107 |
1059 |
13.51 |
|
|
|
| 22 |
A |
1002 |
958 |
18.55 |
|
|
|
| 23 |
A |
998 |
954 |
6.14 |
|
|
|
| 24 |
A |
880 |
841 |
55.86 |
|
|
|
| 25 |
A |
868 |
829 |
6.03 |
|
|
|
| 26 |
A |
842 |
804 |
1.91 |
|
|
|
| 27 |
A |
720 |
688 |
6.25 |
|
|
|
| 28 |
A |
507 |
485 |
2.72 |
|
|
|
| 29 |
A |
443 |
423 |
1.39 |
|
|
|
| 30 |
A |
259 |
248 |
0.58 |
|
|
|
| 31 |
A |
213 |
203 |
0.28 |
|
|
|
| 32 |
A |
90 |
86 |
2.31 |
|
|
|
| 33 |
A |
77 |
74 |
0.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24957.0 cm
-1
Scaled (by 0.956) Zero Point Vibrational Energy (zpe) 23858.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.347 |
|
|
|
| 2 |
H |
0.115 |
|
|
|
| 3 |
H |
0.116 |
|
|
|
| 4 |
C |
0.071 |
|
|
|
| 5 |
H |
0.088 |
|
|
|
| 6 |
O |
-0.223 |
|
|
|
| 7 |
C |
0.036 |
|
|
|
| 8 |
H |
0.065 |
|
|
|
| 9 |
H |
0.060 |
|
|
|
| 10 |
C |
-0.246 |
|
|
|
| 11 |
H |
0.092 |
|
|
|
| 12 |
H |
0.092 |
|
|
|
| 13 |
H |
0.080 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.183 |
1.280 |
0.059 |
1.745 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.728 |
-0.009 |
0.101 |
| y |
-0.009 |
-30.498 |
-0.363 |
| z |
0.101 |
-0.363 |
-33.184 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.113 |
-0.009 |
0.101 |
| y |
-0.009 |
0.958 |
-0.363 |
| z |
0.101 |
-0.363 |
-3.071 |
|
| Polar |
|---|
| 3z2-r2 | -6.143 |
|---|
| x2-y2 | 0.770 |
|---|
| xy | -0.009 |
|---|
| xz | 0.101 |
|---|
| yz | -0.363 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.966 |
0.759 |
-0.062 |
| y |
0.759 |
7.321 |
-0.096 |
| z |
-0.062 |
-0.096 |
5.880 |
<r2> (average value of r
2) Å
2
| <r2> |
161.016 |
| (<r2>)1/2 |
12.689 |