Jump to
S1C2
S1C3
Energy calculated at PBEPBE/STO-3G
| | hartrees |
|---|
| Energy at 0K | -206.215795 |
| Energy at 298.15K | |
| HF Energy | -206.215795 |
| Nuclear repulsion energy | 115.110788 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3659 |
3343 |
6.95 |
|
|
|
| 2 |
A' |
3389 |
3097 |
0.80 |
|
|
|
| 3 |
A' |
3199 |
2923 |
18.05 |
|
|
|
| 4 |
A' |
3074 |
2809 |
37.03 |
|
|
|
| 5 |
A' |
1844 |
1685 |
132.05 |
|
|
|
| 6 |
A' |
1639 |
1498 |
7.31 |
|
|
|
| 7 |
A' |
1534 |
1401 |
11.17 |
|
|
|
| 8 |
A' |
1506 |
1376 |
3.79 |
|
|
|
| 9 |
A' |
1409 |
1288 |
3.00 |
|
|
|
| 10 |
A' |
1249 |
1141 |
55.93 |
|
|
|
| 11 |
A' |
1179 |
1077 |
21.70 |
|
|
|
| 12 |
A' |
1012 |
925 |
40.61 |
|
|
|
| 13 |
A' |
570 |
521 |
5.46 |
|
|
|
| 14 |
A' |
311 |
284 |
2.80 |
|
|
|
| 15 |
A" |
3336 |
3048 |
5.00 |
|
|
|
| 16 |
A" |
1604 |
1465 |
6.67 |
|
|
|
| 17 |
A" |
1164 |
1063 |
0.77 |
|
|
|
| 18 |
A" |
942 |
861 |
5.30 |
|
|
|
| 19 |
A" |
488 |
446 |
100.32 |
|
|
|
| 20 |
A" |
198 |
181 |
2.96 |
|
|
|
| 21 |
A" |
90i |
83i |
12.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16608.6 cm
-1
Scaled (by 0.9136) Zero Point Vibrational Energy (zpe) 15173.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.285 |
-0.801 |
0.000 |
| O2 |
1.443 |
-1.299 |
0.000 |
| N3 |
0.000 |
0.603 |
0.000 |
| C4 |
-1.364 |
1.185 |
0.000 |
| H5 |
-0.699 |
-1.366 |
0.000 |
| H6 |
0.811 |
1.270 |
0.000 |
| H7 |
-2.089 |
0.346 |
0.000 |
| H8 |
-1.546 |
1.811 |
0.903 |
| H9 |
-1.546 |
1.811 |
-0.903 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2605 | 1.4331 | 2.5812 | 1.1341 | 2.1366 | 2.6368 | 3.3158 | 3.3158 |
O2 | 1.2605 | | 2.3877 | 3.7481 | 2.1425 | 2.6456 | 3.8965 | 4.4075 | 4.4075 | N3 | 1.4331 | 2.3877 | | 1.4826 | 2.0895 | 1.0494 | 2.1052 | 2.1600 | 2.1600 | C4 | 2.5812 | 3.7481 | 1.4826 | | 2.6362 | 2.1760 | 1.1094 | 1.1143 | 1.1143 | H5 | 1.1341 | 2.1425 | 2.0895 | 2.6362 | | 3.0372 | 2.2055 | 3.4103 | 3.4103 | H6 | 2.1366 | 2.6456 | 1.0494 | 2.1760 | 3.0372 | | 3.0436 | 2.5812 | 2.5812 | H7 | 2.6368 | 3.8965 | 2.1052 | 1.1094 | 2.2055 | 3.0436 | | 1.8055 | 1.8055 | H8 | 3.3158 | 4.4075 | 2.1600 | 1.1143 | 3.4103 | 2.5812 | 1.8055 | | 1.8065 | H9 | 3.3158 | 4.4075 | 2.1600 | 1.1143 | 3.4103 | 2.5812 | 1.8055 | 1.8065 | |
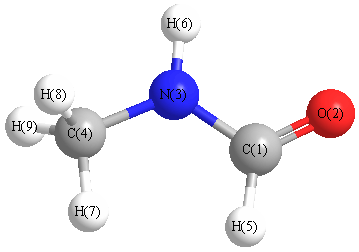 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.566 |
|
C1 |
N3 |
H6 |
117.960 |
| O2 |
C1 |
N3 |
124.738 |
|
O2 |
C1 |
H5 |
126.863 |
| N3 |
C1 |
H5 |
108.400 |
|
N3 |
C4 |
H7 |
107.749 |
| N3 |
C4 |
H8 |
111.777 |
|
N3 |
C4 |
H9 |
111.777 |
| C4 |
N3 |
H6 |
117.474 |
|
H7 |
C4 |
H8 |
108.569 |
| H7 |
C4 |
H9 |
108.569 |
|
H8 |
C4 |
H9 |
108.306 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/STO-3G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.106 |
|
|
|
| 2 |
O |
-0.202 |
|
|
|
| 3 |
N |
-0.282 |
|
|
|
| 4 |
C |
-0.153 |
|
|
|
| 5 |
H |
0.054 |
|
|
|
| 6 |
H |
0.201 |
|
|
|
| 7 |
H |
0.098 |
|
|
|
| 8 |
H |
0.089 |
|
|
|
| 9 |
H |
0.089 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.801 |
2.090 |
0.000 |
2.759 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.016 |
2.633 |
0.000 |
| y |
2.633 |
-21.961 |
0.000 |
| z |
0.000 |
0.000 |
-22.598 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.736 |
2.633 |
0.000 |
| y |
2.633 |
0.846 |
0.000 |
| z |
0.000 |
0.000 |
-0.110 |
|
| Polar |
|---|
| 3z2-r2 | -0.219 |
|---|
| x2-y2 | -1.055 |
|---|
| xy | 2.633 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.416 |
-0.970 |
0.000 |
| y |
-0.970 |
3.640 |
0.000 |
| z |
0.000 |
0.000 |
1.233 |
<r2> (average value of r
2) Å
2
| <r2> |
92.780 |
| (<r2>)1/2 |
9.632 |
Jump to
S1C1
S1C3
Energy calculated at PBEPBE/STO-3G
| | hartrees |
|---|
| Energy at 0K | -206.215795 |
| Energy at 298.15K | |
| HF Energy | -206.215795 |
| Nuclear repulsion energy | 115.110788 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/STO-3G
Geometric Data calculated at PBEPBE/STO-3G
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at PBEPBE/STO-3G
| | hartrees |
|---|
| Energy at 0K | -206.216527 |
| Energy at 298.15K | -206.222179 |
| HF Energy | -206.216527 |
| Nuclear repulsion energy | 117.614832 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3623 |
3310 |
4.12 |
|
|
|
| 2 |
A |
3380 |
3088 |
0.24 |
|
|
|
| 3 |
A |
3354 |
3064 |
4.50 |
|
|
|
| 4 |
A |
3204 |
2927 |
14.74 |
|
|
|
| 5 |
A |
3094 |
2826 |
38.87 |
|
|
|
| 6 |
A |
1825 |
1668 |
89.64 |
|
|
|
| 7 |
A |
1631 |
1490 |
3.83 |
|
|
|
| 8 |
A |
1609 |
1470 |
8.34 |
|
|
|
| 9 |
A |
1512 |
1382 |
39.05 |
|
|
|
| 10 |
A |
1490 |
1362 |
11.37 |
|
|
|
| 11 |
A |
1421 |
1298 |
0.32 |
|
|
|
| 12 |
A |
1205 |
1101 |
67.48 |
|
|
|
| 13 |
A |
1169 |
1068 |
3.43 |
|
|
|
| 14 |
A |
1133 |
1035 |
9.61 |
|
|
|
| 15 |
A |
978 |
894 |
19.29 |
|
|
|
| 16 |
A |
905 |
827 |
0.94 |
|
|
|
| 17 |
A |
735 |
672 |
4.36 |
|
|
|
| 18 |
A |
476 |
435 |
54.82 |
|
|
|
| 19 |
A |
255 |
233 |
28.90 |
|
|
|
| 20 |
A |
192 |
175 |
34.48 |
|
|
|
| 21 |
A |
149 |
136 |
7.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16670.2 cm
-1
Scaled (by 0.9136) Zero Point Vibrational Energy (zpe) 15229.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.895 |
0.454 |
0.007 |
| O2 |
1.470 |
-0.675 |
-0.001 |
| N3 |
-0.516 |
0.669 |
-0.022 |
| C4 |
-1.477 |
-0.468 |
0.005 |
| H5 |
1.423 |
1.463 |
0.019 |
| H6 |
-0.898 |
1.645 |
0.059 |
| H7 |
-2.499 |
-0.097 |
-0.219 |
| H8 |
-1.187 |
-1.221 |
-0.763 |
| H9 |
-1.493 |
-0.984 |
0.996 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2671 | 1.4283 | 2.5457 | 1.1385 | 2.1534 | 3.4463 | 2.7815 | 2.9581 |
O2 | 1.2671 | | 2.3984 | 2.9543 | 2.1386 | 3.3158 | 4.0168 | 2.8179 | 3.1411 | N3 | 1.4283 | 2.3984 | | 1.4891 | 2.0962 | 1.0515 | 2.1346 | 2.1379 | 2.1731 | C4 | 2.5457 | 2.9543 | 1.4891 | | 3.4846 | 2.1925 | 1.1102 | 1.1141 | 1.1170 | H5 | 1.1385 | 2.1386 | 2.0962 | 3.4846 | | 2.3285 | 4.2279 | 3.8250 | 3.9297 | H6 | 2.1534 | 3.3158 | 1.0515 | 2.1925 | 2.3285 | | 2.3828 | 2.9959 | 2.8541 | H7 | 3.4463 | 4.0168 | 2.1346 | 1.1102 | 4.2279 | 2.3828 | | 1.8109 | 1.8100 | H8 | 2.7815 | 2.8179 | 2.1379 | 1.1141 | 3.8250 | 2.9959 | 1.8109 | | 1.8017 | H9 | 2.9581 | 3.1411 | 2.1731 | 1.1170 | 3.9297 | 2.8541 | 1.8100 | 1.8017 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.504 |
|
C1 |
N3 |
H6 |
119.774 |
| O2 |
C1 |
N3 |
125.592 |
|
O2 |
C1 |
H5 |
125.415 |
| N3 |
C1 |
H5 |
108.985 |
|
N3 |
C4 |
H7 |
109.554 |
| N3 |
C4 |
H8 |
109.582 |
|
N3 |
C4 |
H9 |
112.198 |
| C4 |
N3 |
H6 |
118.285 |
|
H7 |
C4 |
H8 |
109.012 |
| H7 |
C4 |
H9 |
108.718 |
|
H8 |
C4 |
H9 |
107.711 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/STO-3G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.103 |
|
|
|
| 2 |
O |
-0.199 |
|
|
|
| 3 |
N |
-0.279 |
|
|
|
| 4 |
C |
-0.159 |
|
|
|
| 5 |
H |
0.058 |
|
|
|
| 6 |
H |
0.193 |
|
|
|
| 7 |
H |
0.089 |
|
|
|
| 8 |
H |
0.109 |
|
|
|
| 9 |
H |
0.084 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.959 |
1.533 |
0.614 |
2.562 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.766 |
1.326 |
-0.413 |
| y |
1.326 |
-20.590 |
0.778 |
| z |
-0.413 |
0.778 |
-22.584 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.179 |
1.326 |
-0.413 |
| y |
1.326 |
2.085 |
0.778 |
| z |
-0.413 |
0.778 |
-0.906 |
|
| Polar |
|---|
| 3z2-r2 | -1.812 |
|---|
| x2-y2 | -2.176 |
|---|
| xy | 1.326 |
|---|
| xz | -0.413 |
|---|
| yz | 0.778 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.838 |
-0.246 |
0.059 |
| y |
-0.246 |
2.963 |
0.125 |
| z |
0.059 |
0.125 |
1.270 |
<r2> (average value of r
2) Å
2
| <r2> |
80.328 |
| (<r2>)1/2 |
8.963 |
