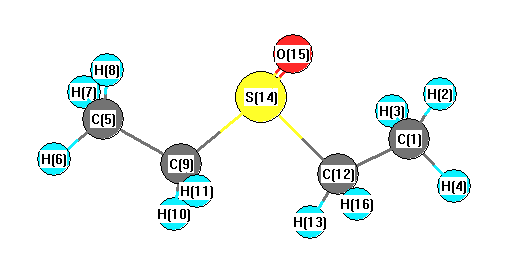
Vibrational Frequencies calculated at PBEPBE/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3060 |
3032 |
18.50 |
|
|
|
| 2 |
A |
3057 |
3029 |
26.67 |
|
|
|
| 3 |
A |
3048 |
3020 |
7.96 |
|
|
|
| 4 |
A |
3043 |
3015 |
17.15 |
|
|
|
| 5 |
A |
3039 |
3011 |
1.97 |
|
|
|
| 6 |
A |
3037 |
3009 |
3.72 |
|
|
|
| 7 |
A |
2998 |
2971 |
7.88 |
|
|
|
| 8 |
A |
2994 |
2967 |
15.21 |
|
|
|
| 9 |
A |
2974 |
2947 |
20.71 |
|
|
|
| 10 |
A |
2973 |
2946 |
19.78 |
|
|
|
| 11 |
A |
1456 |
1443 |
6.53 |
|
|
|
| 12 |
A |
1451 |
1438 |
9.24 |
|
|
|
| 13 |
A |
1448 |
1435 |
5.19 |
|
|
|
| 14 |
A |
1447 |
1434 |
14.33 |
|
|
|
| 15 |
A |
1414 |
1401 |
7.34 |
|
|
|
| 16 |
A |
1410 |
1397 |
5.56 |
|
|
|
| 17 |
A |
1360 |
1348 |
11.30 |
|
|
|
| 18 |
A |
1355 |
1343 |
6.51 |
|
|
|
| 19 |
A |
1252 |
1241 |
0.71 |
|
|
|
| 20 |
A |
1228 |
1216 |
3.11 |
|
|
|
| 21 |
A |
1208 |
1197 |
1.21 |
|
|
|
| 22 |
A |
1205 |
1194 |
0.39 |
|
|
|
| 23 |
A |
1048 |
1038 |
9.35 |
|
|
|
| 24 |
A |
1040 |
1030 |
23.15 |
|
|
|
| 25 |
A |
1028 |
1019 |
81.44 |
|
|
|
| 26 |
A |
1021 |
1012 |
10.42 |
|
|
|
| 27 |
A |
978 |
969 |
2.76 |
|
|
|
| 28 |
A |
956 |
947 |
0.81 |
|
|
|
| 29 |
A |
943 |
935 |
11.81 |
|
|
|
| 30 |
A |
766 |
759 |
9.90 |
|
|
|
| 31 |
A |
736 |
729 |
9.02 |
|
|
|
| 32 |
A |
624 |
618 |
23.65 |
|
|
|
| 33 |
A |
543 |
538 |
9.44 |
|
|
|
| 34 |
A |
454 |
449 |
3.51 |
|
|
|
| 35 |
A |
344 |
341 |
0.85 |
|
|
|
| 36 |
A |
290 |
288 |
2.84 |
|
|
|
| 37 |
A |
260 |
258 |
1.51 |
|
|
|
| 38 |
A |
210 |
208 |
6.27 |
|
|
|
| 39 |
A |
202 |
200 |
1.40 |
|
|
|
| 40 |
A |
190 |
188 |
3.67 |
|
|
|
| 41 |
A |
118 |
117 |
1.67 |
|
|
|
| 42 |
A |
45 |
44 |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29125.4 cm
-1
Scaled (by 0.9909) Zero Point Vibrational Energy (zpe) 28860.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.344 |
|
|
|
| 2 |
H |
0.136 |
|
|
|
| 3 |
H |
0.125 |
|
|
|
| 4 |
H |
0.160 |
|
|
|
| 5 |
C |
-0.349 |
|
|
|
| 6 |
H |
0.141 |
|
|
|
| 7 |
H |
0.142 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
| 9 |
C |
-0.442 |
|
|
|
| 10 |
H |
0.179 |
|
|
|
| 11 |
H |
0.172 |
|
|
|
| 12 |
C |
-0.457 |
|
|
|
| 13 |
H |
0.160 |
|
|
|
| 14 |
S |
0.595 |
|
|
|
| 15 |
O |
-0.526 |
|
|
|
| 16 |
H |
0.178 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.075 |
3.318 |
-0.738 |
3.983 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.154 |
4.254 |
-2.403 |
| y |
4.254 |
-50.487 |
0.842 |
| z |
-2.403 |
0.842 |
-45.777 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.978 |
4.254 |
-2.403 |
| y |
4.254 |
-5.522 |
0.842 |
| z |
-2.403 |
0.842 |
1.543 |
|
| Polar |
|---|
| 3z2-r2 | 3.087 |
|---|
| x2-y2 | 6.334 |
|---|
| xy | 4.254 |
|---|
| xz | -2.403 |
|---|
| yz | 0.842 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.109 |
0.078 |
-0.273 |
| y |
0.078 |
10.686 |
-0.181 |
| z |
-0.273 |
-0.181 |
9.275 |
<r2> (average value of r
2) Å
2
| <r2> |
228.211 |
| (<r2>)1/2 |
15.107 |