Jump to
S1C2
Energy calculated at PBEPBE/TZVP
| | hartrees |
|---|
| Energy at 0K | -168.466691 |
| Energy at 298.15K | |
| HF Energy | -168.466691 |
| Nuclear repulsion energy | 60.146228 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/TZVP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3450 |
3411 |
247.90 |
32.12 |
0.20 |
0.33 |
| 2 |
Σ |
2285 |
2259 |
354.41 |
25.78 |
0.08 |
0.14 |
| 3 |
Σ |
1278 |
1264 |
96.08 |
23.53 |
0.26 |
0.42 |
| 4 |
Π |
538 |
532 |
3.05 |
0.42 |
0.75 |
0.86 |
| 4 |
Π |
538 |
532 |
3.05 |
0.42 |
0.75 |
0.86 |
| 5 |
Π |
156i |
154i |
94.65 |
8.44 |
0.75 |
0.86 |
| 5 |
Π |
156i |
154i |
94.65 |
8.44 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 3888.4 cm
-1
Scaled (by 0.9888) Zero Point Vibrational Energy (zpe) 3844.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/TZVP
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.187 |
| N2 |
0.000 |
0.000 |
-0.017 |
| C3 |
0.000 |
0.000 |
-1.188 |
| H4 |
0.000 |
0.000 |
-2.255 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2043 | 2.3750 | 3.4425 |
N2 | 1.2043 | | 1.1707 | 2.2382 | C3 | 2.3750 | 1.1707 | | 1.0675 | H4 | 3.4425 | 2.2382 | 1.0675 | |
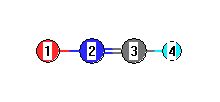 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
180.000 |
|
N2 |
C3 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/TZVP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.201 |
|
|
|
| 2 |
N |
0.217 |
|
|
|
| 3 |
C |
-0.203 |
|
|
|
| 4 |
H |
0.187 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.867 |
2.867 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.893 |
0.000 |
0.000 |
| y |
0.000 |
-16.893 |
0.000 |
| z |
0.000 |
0.000 |
-14.236 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.329 |
0.000 |
0.000 |
| y |
0.000 |
-1.329 |
0.000 |
| z |
0.000 |
0.000 |
2.657 |
|
| Polar |
|---|
| 3z2-r2 | 5.315 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.668 |
0.000 |
0.000 |
| y |
0.000 |
1.668 |
0.000 |
| z |
0.000 |
0.000 |
6.263 |
<r2> (average value of r
2) Å
2
| <r2> |
34.827 |
| (<r2>)1/2 |
5.901 |
Jump to
S1C1
Energy calculated at PBEPBE/TZVP
| | hartrees |
|---|
| Energy at 0K | -168.466753 |
| Energy at 298.15K | -168.467232 |
| HF Energy | -168.466753 |
| Nuclear repulsion energy | 60.146569 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/TZVP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3429 |
3390 |
217.78 |
|
|
|
| 2 |
A' |
2272 |
2247 |
364.96 |
|
|
|
| 3 |
A' |
1283 |
1268 |
86.82 |
|
|
|
| 4 |
A' |
538 |
532 |
4.14 |
|
|
|
| 5 |
A' |
209 |
207 |
120.90 |
|
|
|
| 6 |
A" |
538 |
532 |
3.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4134.4 cm
-1
Scaled (by 0.9888) Zero Point Vibrational Energy (zpe) 4088.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/TZVP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.756 |
-0.913 |
0.000 |
| N2 |
0.000 |
0.022 |
0.000 |
| C3 |
0.797 |
0.884 |
0.000 |
| H4 |
1.270 |
1.843 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2028 | 2.3751 | 3.4203 |
N2 | 1.2028 | | 1.1736 | 2.2195 | C3 | 2.3751 | 1.1736 | | 1.0690 | H4 | 3.4203 | 2.2195 | 1.0690 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
176.200 |
|
N2 |
C3 |
H4 |
163.505 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/TZVP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.193 |
|
|
|
| 2 |
N |
0.205 |
|
|
|
| 3 |
C |
-0.201 |
|
|
|
| 4 |
H |
0.189 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.461 |
2.348 |
0.000 |
2.766 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.671 |
1.016 |
0.000 |
| y |
1.016 |
-14.642 |
0.000 |
| z |
0.000 |
0.000 |
-16.904 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.898 |
1.016 |
0.000 |
| y |
1.016 |
2.145 |
0.000 |
| z |
0.000 |
0.000 |
-1.247 |
|
| Polar |
|---|
| 3z2-r2 | -2.494 |
|---|
| x2-y2 | -2.029 |
|---|
| xy | 1.016 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.748 |
2.285 |
0.000 |
| y |
2.285 |
4.241 |
0.000 |
| z |
0.000 |
0.000 |
1.679 |
<r2> (average value of r
2) Å
2
| <r2> |
34.791 |
| (<r2>)1/2 |
5.898 |
