Jump to
S1C2
Energy calculated at PBEPBE/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -114.334532 |
| Energy at 298.15K | -114.335936 |
| HF Energy | -114.334532 |
| Nuclear repulsion energy | 30.522222 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3568 |
3525 |
50.29 |
|
|
|
| 2 |
A' |
2744 |
2711 |
150.64 |
|
|
|
| 3 |
A' |
1468 |
1450 |
20.54 |
|
|
|
| 4 |
A' |
1285 |
1269 |
46.93 |
|
|
|
| 5 |
A' |
1172 |
1158 |
131.29 |
|
|
|
| 6 |
A" |
1081 |
1068 |
107.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5658.8 cm
-1
Scaled (by 0.9879) Zero Point Vibrational Energy (zpe) 5590.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.004 |
0.729 |
0.000 |
| O2 |
-0.004 |
-0.580 |
0.000 |
| H3 |
0.972 |
1.133 |
0.000 |
| H4 |
-0.921 |
-0.868 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3085 | 1.0559 | 1.8414 |
O2 | 1.3085 | | 1.9710 | 0.9619 | H3 | 1.0559 | 1.9710 | | 2.7544 | H4 | 1.8414 | 0.9619 | 2.7544 | |
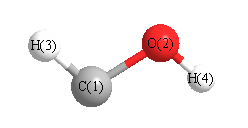 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
107.409 |
|
O2 |
C1 |
H3 |
112.500 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.216 |
|
|
|
| 2 |
O |
-0.128 |
|
|
|
| 3 |
H |
0.085 |
|
|
|
| 4 |
H |
0.259 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.207 |
-1.784 |
0.000 |
1.796 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-11.402 |
3.666 |
0.000 |
| y |
3.666 |
-13.984 |
0.000 |
| z |
0.000 |
0.000 |
-12.293 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.737 |
3.666 |
0.000 |
| y |
3.666 |
-2.136 |
0.000 |
| z |
0.000 |
0.000 |
0.399 |
|
| Polar |
|---|
| 3z2-r2 | 0.798 |
|---|
| x2-y2 | 2.582 |
|---|
| xy | 3.666 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.144 |
0.452 |
0.000 |
| y |
0.452 |
3.132 |
0.000 |
| z |
0.000 |
0.000 |
2.107 |
<r2> (average value of r
2) Å
2
| <r2> |
17.604 |
| (<r2>)1/2 |
4.196 |
Jump to
S1C1
Energy calculated at PBEPBE/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -114.327521 |
| Energy at 298.15K | -114.328911 |
| HF Energy | -114.327521 |
| Nuclear repulsion energy | 30.466302 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3328 |
3288 |
57.80 |
|
|
|
| 2 |
A' |
2656 |
2624 |
217.23 |
|
|
|
| 3 |
A' |
1428 |
1410 |
48.17 |
|
|
|
| 4 |
A' |
1303 |
1288 |
35.71 |
|
|
|
| 5 |
A' |
1154 |
1140 |
20.48 |
|
|
|
| 6 |
A" |
1009 |
996 |
26.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5438.5 cm
-1
Scaled (by 0.9879) Zero Point Vibrational Energy (zpe) 5372.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.117 |
0.736 |
0.000 |
| O2 |
0.117 |
-0.574 |
0.000 |
| H3 |
-0.861 |
1.156 |
0.000 |
| H4 |
-0.773 |
-0.974 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3100 | 1.0646 | 1.9272 |
O2 | 1.3100 | | 1.9879 | 0.9757 | H3 | 1.0646 | 1.9879 | | 2.1317 | H4 | 1.9272 | 0.9757 | 2.1317 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
114.154 |
|
O2 |
C1 |
H3 |
113.274 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.141 |
|
|
|
| 2 |
O |
-0.121 |
|
|
|
| 3 |
H |
0.061 |
|
|
|
| 4 |
H |
0.201 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.860 |
-1.762 |
0.000 |
3.360 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-12.667 |
-0.212 |
0.000 |
| y |
-0.212 |
-13.395 |
0.000 |
| z |
0.000 |
0.000 |
-12.218 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.140 |
-0.212 |
0.000 |
| y |
-0.212 |
-0.952 |
0.000 |
| z |
0.000 |
0.000 |
0.813 |
|
| Polar |
|---|
| 3z2-r2 | 1.625 |
|---|
| x2-y2 | 0.728 |
|---|
| xy | -0.212 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.122 |
0.106 |
0.000 |
| y |
0.106 |
3.753 |
0.000 |
| z |
0.000 |
0.000 |
2.079 |
<r2> (average value of r
2) Å
2
| <r2> |
17.757 |
| (<r2>)1/2 |
4.214 |
