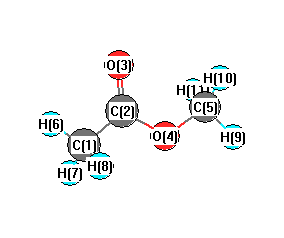
Vibrational Frequencies calculated at PBEPBE/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3106 |
3068 |
4.69 |
|
|
|
| 2 |
A' |
3093 |
3056 |
12.32 |
|
|
|
| 3 |
A' |
2992 |
2956 |
2.75 |
|
|
|
| 4 |
A' |
2983 |
2947 |
27.28 |
|
|
|
| 5 |
A' |
1747 |
1726 |
224.80 |
|
|
|
| 6 |
A' |
1446 |
1428 |
8.50 |
|
|
|
| 7 |
A' |
1420 |
1403 |
19.04 |
|
|
|
| 8 |
A' |
1414 |
1396 |
14.41 |
|
|
|
| 9 |
A' |
1339 |
1323 |
44.97 |
|
|
|
| 10 |
A' |
1214 |
1200 |
282.95 |
|
|
|
| 11 |
A' |
1163 |
1149 |
1.67 |
|
|
|
| 12 |
A' |
1032 |
1019 |
71.84 |
|
|
|
| 13 |
A' |
954 |
943 |
0.99 |
|
|
|
| 14 |
A' |
828 |
818 |
19.98 |
|
|
|
| 15 |
A' |
627 |
619 |
3.90 |
|
|
|
| 16 |
A' |
411 |
406 |
5.18 |
|
|
|
| 17 |
A' |
274 |
271 |
11.77 |
|
|
|
| 18 |
A" |
3057 |
3020 |
3.66 |
|
|
|
| 19 |
A" |
3055 |
3018 |
18.22 |
|
|
|
| 20 |
A" |
1429 |
1412 |
11.31 |
|
|
|
| 21 |
A" |
1423 |
1406 |
6.17 |
|
|
|
| 22 |
A" |
1130 |
1116 |
0.56 |
|
|
|
| 23 |
A" |
1021 |
1009 |
5.60 |
|
|
|
| 24 |
A" |
587 |
580 |
3.15 |
|
|
|
| 25 |
A" |
177 |
175 |
6.02 |
|
|
|
| 26 |
A" |
118 |
117 |
0.30 |
|
|
|
| 27 |
A" |
28i |
27i |
0.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19005.1 cm
-1
Scaled (by 0.9879) Zero Point Vibrational Energy (zpe) 18775.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.252 |
|
|
|
| 2 |
C |
0.247 |
|
|
|
| 3 |
O |
-0.292 |
|
|
|
| 4 |
O |
-0.203 |
|
|
|
| 5 |
C |
-0.159 |
|
|
|
| 6 |
H |
0.109 |
|
|
|
| 7 |
H |
0.103 |
|
|
|
| 8 |
H |
0.103 |
|
|
|
| 9 |
H |
0.117 |
|
|
|
| 10 |
H |
0.114 |
|
|
|
| 11 |
H |
0.114 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.575 |
-0.921 |
0.000 |
1.825 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.820 |
4.467 |
0.000 |
| y |
4.467 |
-26.919 |
0.000 |
| z |
0.000 |
0.000 |
-29.508 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.607 |
4.467 |
0.000 |
| y |
4.467 |
4.245 |
0.000 |
| z |
0.000 |
0.000 |
0.362 |
|
| Polar |
|---|
| 3z2-r2 | 0.723 |
|---|
| x2-y2 | -5.901 |
|---|
| xy | 4.467 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.069 |
0.537 |
0.000 |
| y |
0.537 |
8.097 |
0.000 |
| z |
0.000 |
0.000 |
5.341 |
<r2> (average value of r
2) Å
2
| <r2> |
121.016 |
| (<r2>)1/2 |
11.001 |