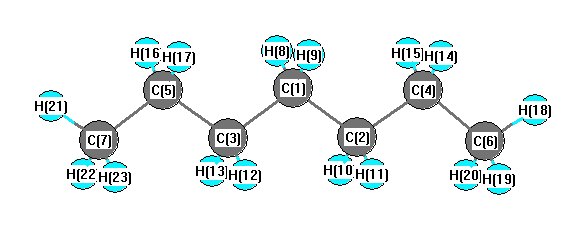
Vibrational Frequencies calculated at PBEPBE/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3062 |
3020 |
50.61 |
|
|
|
| 2 |
A1 |
2975 |
2934 |
29.99 |
|
|
|
| 3 |
A1 |
2963 |
2922 |
154.07 |
|
|
|
| 4 |
A1 |
2951 |
2910 |
10.61 |
|
|
|
| 5 |
A1 |
2943 |
2902 |
0.18 |
|
|
|
| 6 |
A1 |
1516 |
1495 |
11.07 |
|
|
|
| 7 |
A1 |
1503 |
1482 |
1.01 |
|
|
|
| 8 |
A1 |
1492 |
1471 |
0.00 |
|
|
|
| 9 |
A1 |
1486 |
1465 |
0.01 |
|
|
|
| 10 |
A1 |
1411 |
1392 |
3.72 |
|
|
|
| 11 |
A1 |
1372 |
1353 |
0.84 |
|
|
|
| 12 |
A1 |
1313 |
1295 |
0.01 |
|
|
|
| 13 |
A1 |
1147 |
1131 |
1.51 |
|
|
|
| 14 |
A1 |
1073 |
1058 |
0.57 |
|
|
|
| 15 |
A1 |
993 |
980 |
0.02 |
|
|
|
| 16 |
A1 |
916 |
904 |
2.15 |
|
|
|
| 17 |
A1 |
412 |
406 |
0.12 |
|
|
|
| 18 |
A1 |
300 |
296 |
0.01 |
|
|
|
| 19 |
A1 |
97 |
95 |
0.02 |
|
|
|
| 20 |
A2 |
3056 |
3013 |
0.00 |
|
|
|
| 21 |
A2 |
3006 |
2965 |
0.00 |
|
|
|
| 22 |
A2 |
2978 |
2936 |
0.00 |
|
|
|
| 23 |
A2 |
1499 |
1478 |
0.00 |
|
|
|
| 24 |
A2 |
1317 |
1299 |
0.00 |
|
|
|
| 25 |
A2 |
1308 |
1289 |
0.00 |
|
|
|
| 26 |
A2 |
1234 |
1217 |
0.00 |
|
|
|
| 27 |
A2 |
1028 |
1013 |
0.00 |
|
|
|
| 28 |
A2 |
853 |
841 |
0.00 |
|
|
|
| 29 |
A2 |
744 |
734 |
0.00 |
|
|
|
| 30 |
A2 |
236 |
233 |
0.00 |
|
|
|
| 31 |
A2 |
142 |
140 |
0.00 |
|
|
|
| 32 |
A2 |
84 |
83 |
0.00 |
|
|
|
| 33 |
B1 |
3056 |
3014 |
130.62 |
|
|
|
| 34 |
B1 |
3019 |
2977 |
115.07 |
|
|
|
| 35 |
B1 |
2991 |
2950 |
0.30 |
|
|
|
| 36 |
B1 |
2970 |
2929 |
0.02 |
|
|
|
| 37 |
B1 |
1499 |
1478 |
17.36 |
|
|
|
| 38 |
B1 |
1321 |
1303 |
0.76 |
|
|
|
| 39 |
B1 |
1282 |
1264 |
0.03 |
|
|
|
| 40 |
B1 |
1201 |
1184 |
0.25 |
|
|
|
| 41 |
B1 |
943 |
930 |
1.55 |
|
|
|
| 42 |
B1 |
780 |
769 |
2.41 |
|
|
|
| 43 |
B1 |
737 |
727 |
8.84 |
|
|
|
| 44 |
B1 |
238 |
234 |
0.00 |
|
|
|
| 45 |
B1 |
149 |
147 |
0.00 |
|
|
|
| 46 |
B1 |
66 |
65 |
0.00 |
|
|
|
| 47 |
B2 |
3062 |
3019 |
35.79 |
|
|
|
| 48 |
B2 |
2975 |
2934 |
52.29 |
|
|
|
| 49 |
B2 |
2960 |
2919 |
0.05 |
|
|
|
| 50 |
B2 |
2944 |
2903 |
0.00 |
|
|
|
| 51 |
B2 |
1510 |
1489 |
1.45 |
|
|
|
| 52 |
B2 |
1496 |
1475 |
2.63 |
|
|
|
| 53 |
B2 |
1486 |
1465 |
0.18 |
|
|
|
| 54 |
B2 |
1411 |
1392 |
3.93 |
|
|
|
| 55 |
B2 |
1366 |
1348 |
0.65 |
|
|
|
| 56 |
B2 |
1355 |
1336 |
2.10 |
|
|
|
| 57 |
B2 |
1259 |
1241 |
0.19 |
|
|
|
| 58 |
B2 |
1089 |
1074 |
8.40 |
|
|
|
| 59 |
B2 |
1069 |
1054 |
2.90 |
|
|
|
| 60 |
B2 |
1044 |
1029 |
0.00 |
|
|
|
| 61 |
B2 |
878 |
866 |
3.11 |
|
|
|
| 62 |
B2 |
479 |
472 |
0.07 |
|
|
|
| 63 |
B2 |
240 |
237 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 47139.6 cm
-1
Scaled (by 0.9862) Zero Point Vibrational Energy (zpe) 46489.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.273 |
|
|
|
| 2 |
C |
-0.266 |
|
|
|
| 3 |
C |
-0.266 |
|
|
|
| 4 |
C |
-0.275 |
|
|
|
| 5 |
C |
-0.275 |
|
|
|
| 6 |
C |
-0.454 |
|
|
|
| 7 |
C |
-0.454 |
|
|
|
| 8 |
H |
0.137 |
|
|
|
| 9 |
H |
0.137 |
|
|
|
| 10 |
H |
0.136 |
|
|
|
| 11 |
H |
0.136 |
|
|
|
| 12 |
H |
0.136 |
|
|
|
| 13 |
H |
0.136 |
|
|
|
| 14 |
H |
0.141 |
|
|
|
| 15 |
H |
0.141 |
|
|
|
| 16 |
H |
0.141 |
|
|
|
| 17 |
H |
0.141 |
|
|
|
| 18 |
H |
0.146 |
|
|
|
| 19 |
H |
0.148 |
|
|
|
| 20 |
H |
0.148 |
|
|
|
| 21 |
H |
0.146 |
|
|
|
| 22 |
H |
0.148 |
|
|
|
| 23 |
H |
0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.039 |
0.039 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.527 |
0.000 |
0.000 |
| y |
0.000 |
-48.677 |
0.000 |
| z |
0.000 |
0.000 |
-48.453 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.039 |
0.000 |
0.000 |
| y |
0.000 |
-0.687 |
0.000 |
| z |
0.000 |
0.000 |
-0.351 |
|
| Polar |
|---|
| 3z2-r2 | -0.703 |
|---|
| x2-y2 | 1.150 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.186 |
0.000 |
0.000 |
| y |
0.000 |
14.061 |
0.000 |
| z |
0.000 |
0.000 |
10.446 |
<r2> (average value of r
2) Å
2
| <r2> |
482.855 |
| (<r2>)1/2 |
21.974 |