Jump to
S1C2
Energy calculated at PBEPBE/6-31G
| | hartrees |
|---|
| Energy at 0K | -169.649885 |
| Energy at 298.15K | -169.653874 |
| HF Energy | -169.649885 |
| Nuclear repulsion energy | 69.917934 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3651 |
3601 |
20.38 |
|
|
|
| 2 |
A' |
3504 |
3455 |
16.37 |
|
|
|
| 3 |
A' |
2924 |
2884 |
115.16 |
|
|
|
| 4 |
A' |
1705 |
1682 |
292.03 |
|
|
|
| 5 |
A' |
1617 |
1594 |
38.41 |
|
|
|
| 6 |
A' |
1388 |
1369 |
13.66 |
|
|
|
| 7 |
A' |
1265 |
1248 |
77.03 |
|
|
|
| 8 |
A' |
1041 |
1027 |
1.64 |
|
|
|
| 9 |
A' |
548 |
540 |
10.16 |
|
|
|
| 10 |
A" |
1007 |
993 |
0.90 |
|
|
|
| 11 |
A" |
670 |
660 |
146.13 |
|
|
|
| 12 |
A" |
521 |
513 |
232.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9920.0 cm
-1
Scaled (by 0.9862) Zero Point Vibrational Energy (zpe) 9783.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.426 |
0.000 |
| O2 |
1.232 |
0.206 |
0.000 |
| N3 |
-0.969 |
-0.545 |
0.000 |
| H4 |
-0.428 |
1.452 |
0.000 |
| H5 |
-0.692 |
-1.524 |
0.000 |
| H6 |
-1.957 |
-0.313 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2516 | 1.3716 | 1.1119 | 2.0696 | 2.0922 |
O2 | 1.2516 | | 2.3252 | 2.0759 | 2.5874 | 3.2312 | N3 | 1.3716 | 2.3252 | | 2.0696 | 1.0176 | 1.0157 | H4 | 1.1119 | 2.0759 | 2.0696 | | 2.9887 | 2.3359 | H5 | 2.0696 | 2.5874 | 1.0176 | 2.9887 | | 1.7519 | H6 | 2.0922 | 3.2312 | 1.0157 | 2.3359 | 1.7519 | |
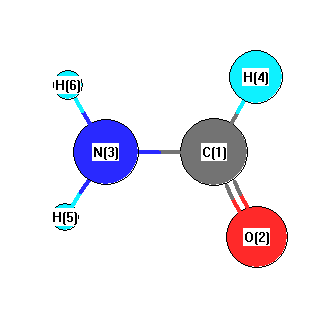 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.305 |
|
C1 |
N3 |
H6 |
121.698 |
| O2 |
C1 |
N3 |
124.775 |
|
O2 |
C1 |
H4 |
122.762 |
| N3 |
C1 |
H4 |
112.463 |
|
H5 |
N3 |
H6 |
118.997 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.298 |
|
|
|
| 2 |
O |
-0.408 |
|
|
|
| 3 |
N |
-0.668 |
|
|
|
| 4 |
H |
0.118 |
|
|
|
| 5 |
H |
0.334 |
|
|
|
| 6 |
H |
0.326 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.997 |
-0.769 |
0.000 |
4.071 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.130 |
0.108 |
0.000 |
| y |
0.108 |
-14.737 |
0.000 |
| z |
0.000 |
0.000 |
-18.698 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.412 |
0.108 |
0.000 |
| y |
0.108 |
3.677 |
0.000 |
| z |
0.000 |
0.000 |
-2.265 |
|
| Polar |
|---|
| 3z2-r2 | -4.530 |
|---|
| x2-y2 | -3.393 |
|---|
| xy | 0.108 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.440 |
0.297 |
0.000 |
| y |
0.297 |
3.272 |
0.000 |
| z |
0.000 |
0.000 |
1.221 |
<r2> (average value of r
2) Å
2
| <r2> |
41.978 |
| (<r2>)1/2 |
6.479 |
Jump to
S1C1
Energy calculated at PBEPBE/6-31G
| | hartrees |
|---|
| Energy at 0K | -169.649882 |
| Energy at 298.15K | -169.653870 |
| HF Energy | -169.649882 |
| Nuclear repulsion energy | 69.917896 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3651 |
3601 |
20.33 |
|
|
|
| 2 |
A |
3504 |
3456 |
16.38 |
|
|
|
| 3 |
A |
2923 |
2883 |
115.20 |
|
|
|
| 4 |
A |
1705 |
1682 |
292.15 |
|
|
|
| 5 |
A |
1616 |
1594 |
38.20 |
|
|
|
| 6 |
A |
1388 |
1369 |
13.62 |
|
|
|
| 7 |
A |
1265 |
1248 |
77.20 |
|
|
|
| 8 |
A |
1041 |
1026 |
1.61 |
|
|
|
| 9 |
A |
1007 |
993 |
0.93 |
|
|
|
| 10 |
A |
671 |
662 |
148.27 |
|
|
|
| 11 |
A |
547 |
540 |
10.16 |
|
|
|
| 12 |
A |
519 |
512 |
230.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9918.9 cm
-1
Scaled (by 0.9862) Zero Point Vibrational Energy (zpe) 9782.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/6-31G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.152 |
0.398 |
0.000 |
| O2 |
-1.224 |
-0.247 |
-0.000 |
| N3 |
1.099 |
-0.164 |
-0.000 |
| H4 |
-0.118 |
1.510 |
0.000 |
| H5 |
1.190 |
-1.178 |
0.001 |
| H6 |
1.941 |
0.405 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2516 | 1.3716 | 1.1120 | 2.0696 | 2.0925 |
O2 | 1.2516 | | 2.3251 | 2.0759 | 2.5872 | 3.2314 | N3 | 1.3716 | 2.3251 | | 2.0698 | 1.0177 | 1.0156 | H4 | 1.1120 | 2.0759 | 2.0698 | | 2.9888 | 2.3367 | H5 | 2.0696 | 2.5872 | 1.0177 | 2.9888 | | 1.7516 | H6 | 2.0925 | 3.2314 | 1.0156 | 2.3367 | 1.7516 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.300 |
|
C1 |
N3 |
H6 |
121.735 |
| O2 |
C1 |
N3 |
124.768 |
|
O2 |
C1 |
H4 |
122.757 |
| N3 |
C1 |
H4 |
112.475 |
|
H5 |
N3 |
H6 |
118.965 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.298 |
|
|
|
| 2 |
O |
-0.408 |
|
|
|
| 3 |
N |
-0.668 |
|
|
|
| 4 |
H |
0.118 |
|
|
|
| 5 |
H |
0.334 |
|
|
|
| 6 |
H |
0.326 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.009 |
0.706 |
0.004 |
4.071 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.625 |
-1.212 |
0.005 |
| y |
-1.212 |
-15.240 |
-0.002 |
| z |
0.005 |
-0.002 |
-18.698 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.655 |
-1.212 |
0.005 |
| y |
-1.212 |
2.921 |
-0.002 |
| z |
0.005 |
-0.002 |
-2.266 |
|
| Polar |
|---|
| 3z2-r2 | -4.531 |
|---|
| x2-y2 | -2.384 |
|---|
| xy | -1.212 |
|---|
| xz | 0.005 |
|---|
| yz | -0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.489 |
0.167 |
0.000 |
| y |
0.167 |
3.223 |
0.000 |
| z |
0.000 |
0.000 |
1.221 |
<r2> (average value of r
2) Å
2
| <r2> |
41.977 |
| (<r2>)1/2 |
6.479 |
