return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHFCH3 (2-Fluoropropane)

using model chemistry: MP3/TZVP

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP3/TZVP
 hartrees
Energy at 0K-217.896866
Energy at 298.15K-217.904792
HF Energy-217.203031
Nuclear repulsion energy132.923460
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/TZVP
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3188 2981 31.15      
2 A' 3181 2975 49.39      
3 A' 3113 2912 24.67      
4 A' 3098 2897 8.47      
5 A' 1539 1439 5.43      
6 A' 1524 1425 4.97      
7 A' 1461 1367 23.91      
8 A' 1428 1336 11.80      
9 A' 1234 1154 12.17      
10 A' 1201 1123 51.03      
11 A' 998 933 43.65      
12 A' 849 794 8.98      
13 A' 489 457 3.68      
14 A' 360 336 0.52      
15 A' 269 251 0.11      
16 A" 3185 2979 18.34      
17 A" 3173 2968 0.03      
18 A" 3094 2893 12.69      
19 A" 1514 1416 0.05      
20 A" 1512 1414 0.02      
21 A" 1456 1362 26.31      
22 A" 1410 1319 2.39      
23 A" 1194 1117 13.39      
24 A" 967 904 0.01      
25 A" 956 894 0.11      
26 A" 422 395 7.45      
27 A" 207 194 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 21510.3 cm-1
Scaled (by 0.9352) Zero Point Vibrational Energy (zpe) 20116.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/TZVP
ABC
0.29145 0.27174 0.16052

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/TZVP

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.277 0.244 0.000
F2 -0.865 1.044 0.000
H3 1.128 0.928 0.000
C4 0.277 -0.586 1.267
C5 0.277 -0.586 -1.267
H6 1.185 -1.187 1.319
H7 1.185 -1.187 -1.319
H8 0.235 0.058 2.144
H9 0.235 0.058 -2.144
H10 -0.585 -1.252 1.280
H11 -0.585 -1.252 -1.280

Atom - Atom Distances (Å)
  C1 F2 H3 C4 C5 H6 H7 H8 H9 H10 H11
C11.39461.09191.51451.51452.14742.14742.15232.15232.14952.1495
F21.39461.99702.35932.35933.30463.30462.60342.60342.64392.6439
H31.09191.99702.14952.14952.49262.49262.48002.48003.05403.0540
C41.51452.35932.14952.53421.09032.80591.08883.47161.09022.7702
C51.51452.35932.14952.53422.80591.09033.47161.08882.77021.0902
H62.14743.30462.49261.09032.80592.63771.77063.80061.77223.1450
H72.14743.30462.49262.80591.09032.63773.80061.77063.14501.7722
H82.15232.60342.48001.08883.47161.77063.80064.28781.77143.7565
H92.15232.60342.48003.47161.08883.80061.77064.28783.75651.7714
H102.14952.64393.05401.09022.77021.77223.14501.77143.75652.5592
H112.14952.64393.05402.77021.09023.14501.77223.75651.77142.5592

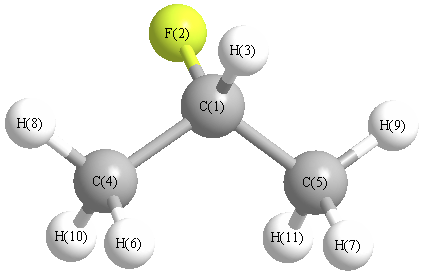
picture of 2-Fluoropropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H8 110.468 C1 C4 H10 110.159
C1 C5 H7 109.979 C1 C5 H9 110.468
C1 C5 H11 110.159 F2 C1 H3 106.221
F2 C1 C4 108.319 F2 C1 C5 108.319
H3 C1 C4 110.057 H3 C1 C5 110.057
C4 C1 C5 113.573 H7 C5 H9 108.689
H7 C5 H11 108.729 H8 C4 H10 108.772
H9 C5 H11 108.772
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability