Jump to
S2C1
Energy calculated at CCSD=FULL/aug-cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -151.122836 |
| Energy at 298.15K | -151.121304 |
| HF Energy | -150.530724 |
| Nuclear repulsion energy | 46.263231 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCSD=FULL/aug-cc-pVQZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.409 |
| C2 |
0.000 |
0.000 |
-0.053 |
| O3 |
0.000 |
0.000 |
1.097 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
| C1 | | 1.3558 | 2.5063 |
C2 | 1.3558 | | 1.1505 | O3 | 2.5063 | 1.1505 | |
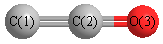 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD=FULL/aug-cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -151.089492 |
| Energy at 298.15K | -151.087958 |
| HF Energy | -150.469684 |
| Nuclear repulsion energy | 46.074925 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCSD=FULL/aug-cc-pVQZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.412 |
| C2 |
0.000 |
0.000 |
-0.058 |
| O3 |
0.000 |
0.000 |
1.102 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
| C1 | | 1.3541 | 2.5138 |
C2 | 1.3541 | | 1.1597 | O3 | 2.5138 | 1.1597 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
