return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2CN (Butanenitrile)

using model chemistry: CCSD=FULL/TZVP

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCSD=FULL/TZVP
 hartrees
Energy at 0K-210.941132
Energy at 298.15K-210.948409
HF Energy-210.063949
Nuclear repulsion energy154.260664
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/TZVP
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3146 2981 29.90      
2 A' 3090 2928 23.31      
3 A' 3081 2920 3.04      
4 A' 3061 2901 21.39      
5 A' 2365 2241 1.76      
6 A' 1535 1454 6.47      
7 A' 1520 1440 1.03      
8 A' 1505 1426 2.80      
9 A' 1454 1378 1.78      
10 A' 1434 1359 1.84      
11 A' 1336 1266 2.39      
12 A' 1143 1083 1.00      
13 A' 1077 1021 0.09      
14 A' 970 919 2.01      
15 A' 896 849 1.65      
16 A' 528 501 0.64      
17 A' 348 330 0.09      
18 A' 164 155 6.18      
19 A" 3147 2981 49.38      
20 A" 3131 2966 5.25      
21 A" 3114 2951 0.15      
22 A" 1526 1446 6.94      
23 A" 1351 1280 0.14      
24 A" 1287 1219 0.02      
25 A" 1154 1093 0.01      
26 A" 896 849 0.11      
27 A" 758 718 1.38      
28 A" 374 355 0.40      
29 A" 237 224 0.00      
30 A" 95 90 4.21      

Unscaled Zero Point Vibrational Energy (zpe) 22862.0 cm-1
Scaled (by 0.9475) Zero Point Vibrational Energy (zpe) 21661.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCSD=FULL/TZVP
ABC
0.79716 0.07558 0.07179

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCSD=FULL/TZVP

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -2.607 0.347 0.000
C2 -1.460 0.489 0.000
C3 0.000 0.636 0.000
C4 0.722 -0.717 0.000
C5 2.237 -0.542 0.000
H6 0.284 1.217 0.880
H7 0.284 1.217 -0.880
H8 0.408 -1.285 0.878
H9 0.408 -1.285 -0.878
H10 2.740 -1.509 0.000
H11 2.566 0.009 0.884
H12 2.566 0.009 -0.884

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.15542.62273.49394.92443.14433.14433.53833.53835.65935.25885.2588
C21.15541.46752.49283.83822.08442.08442.72182.72184.65124.15034.1503
C32.62271.46751.53342.52821.09181.09182.15152.15153.47992.78602.7860
C43.49392.49281.53341.52552.16912.16911.09171.09172.16822.17042.1704
C54.92443.83822.52821.52552.77172.77172.16082.16081.09051.09221.0922
H63.14432.08441.09182.16912.77171.75992.50533.06033.77342.58263.1276
H73.14432.08441.09182.16912.77171.75993.06032.50533.77343.12762.5826
H83.53832.72182.15151.09172.16082.50533.06031.75522.50162.51663.0719
H93.53832.72182.15151.09172.16083.06032.50531.75522.50163.07192.5166
H105.65934.65123.47992.16821.09053.77343.77342.50162.50161.76521.7652
H115.25884.15032.78602.17041.09222.58263.12762.51663.07191.76521.7680
H125.25884.15032.78602.17041.09223.12762.58263.07192.51661.76521.7680

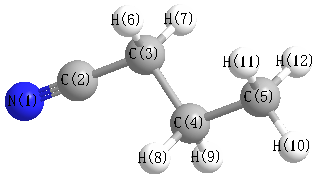
picture of Butanenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 178.651 C2 C3 C4 112.324
C2 C3 H6 108.174 C2 C3 H7 108.174
C3 C4 C5 111.487 C3 C4 H8 108.922
C3 C4 H9 108.922 C4 C3 H6 110.301
C4 C3 H7 110.301 C4 C5 H10 110.864
C4 C5 H11 110.936 C4 C5 H12 110.936
C5 C4 H8 110.195 C5 C4 H9 110.195
H6 C3 H7 107.407 H8 C4 H9 107.001
H10 C5 H11 107.950 H10 C5 H12 107.950
H11 C5 H12 108.077
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability