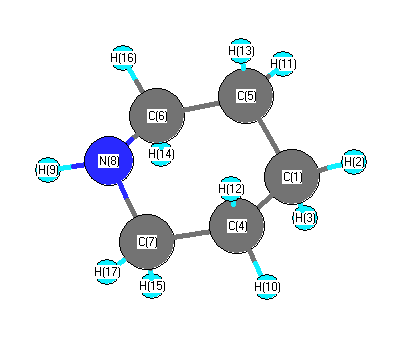
Vibrational Frequencies calculated at B3LYPultrafine/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3527 |
3409 |
0.01 |
|
|
|
| 2 |
A' |
3064 |
2962 |
85.01 |
|
|
|
| 3 |
A' |
3056 |
2954 |
3.85 |
|
|
|
| 4 |
A' |
3050 |
2949 |
69.56 |
|
|
|
| 5 |
A' |
3020 |
2920 |
17.57 |
|
|
|
| 6 |
A' |
3002 |
2902 |
24.99 |
|
|
|
| 7 |
A' |
2893 |
2797 |
156.25 |
|
|
|
| 8 |
A' |
1512 |
1462 |
2.29 |
|
|
|
| 9 |
A' |
1496 |
1446 |
6.22 |
|
|
|
| 10 |
A' |
1488 |
1439 |
7.14 |
|
|
|
| 11 |
A' |
1424 |
1376 |
2.26 |
|
|
|
| 12 |
A' |
1383 |
1337 |
0.20 |
|
|
|
| 13 |
A' |
1314 |
1270 |
0.90 |
|
|
|
| 14 |
A' |
1290 |
1247 |
2.97 |
|
|
|
| 15 |
A' |
1171 |
1132 |
4.55 |
|
|
|
| 16 |
A' |
1065 |
1030 |
9.19 |
|
|
|
| 17 |
A' |
1049 |
1014 |
1.63 |
|
|
|
| 18 |
A' |
908 |
877 |
3.05 |
|
|
|
| 19 |
A' |
871 |
842 |
9.72 |
|
|
|
| 20 |
A' |
818 |
791 |
0.27 |
|
|
|
| 21 |
A' |
759 |
734 |
79.46 |
|
|
|
| 22 |
A' |
547 |
528 |
21.42 |
|
|
|
| 23 |
A' |
435 |
421 |
7.10 |
|
|
|
| 24 |
A' |
397 |
384 |
1.45 |
|
|
|
| 25 |
A' |
242 |
234 |
1.24 |
|
|
|
| 26 |
A" |
3058 |
2957 |
16.28 |
|
|
|
| 27 |
A" |
3054 |
2952 |
92.27 |
|
|
|
| 28 |
A" |
3021 |
2920 |
24.82 |
|
|
|
| 29 |
A" |
2888 |
2792 |
31.10 |
|
|
|
| 30 |
A" |
1501 |
1451 |
0.17 |
|
|
|
| 31 |
A" |
1489 |
1440 |
1.22 |
|
|
|
| 32 |
A" |
1478 |
1429 |
4.39 |
|
|
|
| 33 |
A" |
1378 |
1332 |
0.76 |
|
|
|
| 34 |
A" |
1368 |
1322 |
3.80 |
|
|
|
| 35 |
A" |
1349 |
1304 |
24.02 |
|
|
|
| 36 |
A" |
1296 |
1253 |
2.06 |
|
|
|
| 37 |
A" |
1185 |
1146 |
5.71 |
|
|
|
| 38 |
A" |
1171 |
1132 |
3.28 |
|
|
|
| 39 |
A" |
1128 |
1090 |
19.53 |
|
|
|
| 40 |
A" |
1063 |
1028 |
1.98 |
|
|
|
| 41 |
A" |
983 |
950 |
0.81 |
|
|
|
| 42 |
A" |
878 |
849 |
0.23 |
|
|
|
| 43 |
A" |
823 |
795 |
0.72 |
|
|
|
| 44 |
A" |
453 |
438 |
0.78 |
|
|
|
| 45 |
A" |
248 |
240 |
0.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34796.0 cm
-1
Scaled (by 0.9667) Zero Point Vibrational Energy (zpe) 33637.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYPultrafine/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.165 |
|
|
|
| 2 |
H |
0.090 |
|
|
|
| 3 |
H |
0.076 |
|
|
|
| 4 |
C |
-0.154 |
|
|
|
| 5 |
C |
-0.154 |
|
|
|
| 6 |
C |
-0.085 |
|
|
|
| 7 |
C |
-0.085 |
|
|
|
| 8 |
N |
-0.223 |
|
|
|
| 9 |
H |
0.105 |
|
|
|
| 10 |
H |
0.086 |
|
|
|
| 11 |
H |
0.086 |
|
|
|
| 12 |
H |
0.076 |
|
|
|
| 13 |
H |
0.076 |
|
|
|
| 14 |
H |
0.051 |
|
|
|
| 15 |
H |
0.051 |
|
|
|
| 16 |
H |
0.084 |
|
|
|
| 17 |
H |
0.084 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.631 |
-0.455 |
0.000 |
0.778 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.589 |
-0.045 |
0.003 |
| y |
-0.045 |
-37.102 |
-0.001 |
| z |
0.003 |
-0.001 |
-38.566 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.755 |
-0.045 |
0.003 |
| y |
-0.045 |
2.976 |
-0.001 |
| z |
0.003 |
-0.001 |
0.779 |
|
| Polar |
|---|
| 3z2-r2 | 1.559 |
|---|
| x2-y2 | -4.487 |
|---|
| xy | -0.045 |
|---|
| xz | 0.003 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.953 |
0.294 |
0.000 |
| y |
0.294 |
10.021 |
0.001 |
| z |
0.000 |
0.001 |
10.471 |
<r2> (average value of r
2) Å
2
| <r2> |
158.473 |
| (<r2>)1/2 |
12.589 |