Jump to
S1C2
Energy calculated at PBEPBEultrafine/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -166.334894 |
| Energy at 298.15K | |
| HF Energy | -166.334894 |
| Nuclear repulsion energy | 48.552087 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3930 |
3885 |
48.97 |
|
|
|
| 2 |
A1 |
722 |
713 |
3.27 |
|
|
|
| 3 |
A1 |
511 |
505 |
226.16 |
|
|
|
| 4 |
A1 |
281 |
278 |
0.12 |
|
|
|
| 5 |
A2 |
201i |
199i |
0.00 |
|
|
|
| 6 |
B1 |
368 |
364 |
64.02 |
|
|
|
| 7 |
B2 |
3928 |
3883 |
183.89 |
|
|
|
| 8 |
B2 |
1498 |
1481 |
422.25 |
|
|
|
| 9 |
B2 |
367 |
363 |
260.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5701.5 cm
-1
Scaled (by 0.9886) Zero Point Vibrational Energy (zpe) 5636.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31+G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
-0.005 |
| O2 |
0.000 |
1.434 |
0.073 |
| O3 |
0.000 |
-1.434 |
0.073 |
| H4 |
0.000 |
2.144 |
-0.576 |
| H5 |
0.000 |
-2.144 |
-0.576 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4362 | 1.4362 | 2.2185 | 2.2185 |
O2 | 1.4362 | | 2.8681 | 0.9619 | 3.6363 | O3 | 1.4362 | 2.8681 | | 3.6363 | 0.9619 | H4 | 2.2185 | 0.9619 | 3.6363 | | 4.2878 | H5 | 2.2185 | 3.6363 | 0.9619 | 4.2878 | |
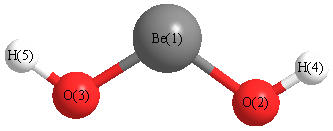 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
134.419 |
|
Be1 |
O3 |
H5 |
134.419 |
| O2 |
Be1 |
O3 |
173.717 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31+G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
0.975 |
|
|
|
| 2 |
O |
-0.852 |
|
|
|
| 3 |
O |
-0.852 |
|
|
|
| 4 |
H |
0.365 |
|
|
|
| 5 |
H |
0.365 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.508 |
2.508 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.469 |
0.000 |
0.000 |
| y |
0.000 |
-14.389 |
0.000 |
| z |
0.000 |
0.000 |
-15.352 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.598 |
0.000 |
0.000 |
| y |
0.000 |
2.021 |
0.000 |
| z |
0.000 |
0.000 |
0.577 |
|
| Polar |
|---|
| 3z2-r2 | 1.154 |
|---|
| x2-y2 | -3.080 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.632 |
0.000 |
0.000 |
| y |
0.000 |
4.587 |
0.000 |
| z |
0.000 |
0.000 |
2.581 |
<r2> (average value of r
2) Å
2
| <r2> |
52.674 |
| (<r2>)1/2 |
7.258 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -166.335602 |
| Energy at 298.15K | -166.336938 |
| HF Energy | -166.335602 |
| Nuclear repulsion energy | 48.510754 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3909 |
3865 |
23.56 |
|
|
|
| 2 |
A |
716 |
708 |
0.92 |
|
|
|
| 3 |
A |
502 |
496 |
121.61 |
|
|
|
| 4 |
A |
313 |
309 |
17.74 |
|
|
|
| 5 |
A |
192 |
190 |
166.62 |
|
|
|
| 6 |
B |
3908 |
3863 |
186.80 |
|
|
|
| 7 |
B |
1493 |
1476 |
410.00 |
|
|
|
| 8 |
B |
506 |
500 |
327.64 |
|
|
|
| 9 |
B |
303 |
299 |
99.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5920.8 cm
-1
Scaled (by 0.9886) Zero Point Vibrational Energy (zpe) 5853.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31+G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
-0.003 |
| O2 |
0.000 |
1.438 |
-0.050 |
| O3 |
0.000 |
-1.438 |
-0.050 |
| H4 |
0.542 |
2.089 |
0.408 |
| H5 |
-0.542 |
-2.089 |
0.408 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4386 | 1.4386 | 2.1971 | 2.1971 |
O2 | 1.4386 | | 2.8757 | 0.9633 | 3.5976 | O3 | 1.4386 | 2.8757 | | 3.5976 | 0.9633 | H4 | 2.1971 | 0.9633 | 3.5976 | | 4.3166 | H5 | 2.1971 | 3.5976 | 0.9633 | 4.3166 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
131.311 |
|
Be1 |
O3 |
H5 |
131.311 |
| O2 |
Be1 |
O3 |
176.267 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31+G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
0.960 |
|
|
|
| 2 |
O |
-0.840 |
|
|
|
| 3 |
O |
-0.840 |
|
|
|
| 4 |
H |
0.360 |
|
|
|
| 5 |
H |
0.360 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.733 |
1.733 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.742 |
4.889 |
0.000 |
| y |
4.889 |
-15.537 |
0.000 |
| z |
0.000 |
0.000 |
-16.421 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.237 |
4.889 |
0.000 |
| y |
4.889 |
0.545 |
0.000 |
| z |
0.000 |
0.000 |
-0.781 |
|
| Polar |
|---|
| 3z2-r2 | -1.563 |
|---|
| x2-y2 | -0.205 |
|---|
| xy | 4.889 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.613 |
-0.002 |
0.000 |
| y |
-0.002 |
4.594 |
0.000 |
| z |
0.000 |
0.000 |
2.631 |
<r2> (average value of r
2) Å
2
| <r2> |
52.700 |
| (<r2>)1/2 |
7.259 |
