return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CF3CHF2 (pentafluoroethane)

using model chemistry: MP3=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP3=FULL/cc-pVTZ
 hartrees
Energy at 0K-575.392463
Energy at 298.15K-575.396061
HF Energy-573.741972
Nuclear repulsion energy363.342621
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3184 2970 17.60      
2 A' 1525 1422 2.89      
3 A' 1389 1295 184.47      
4 A' 1280 1194 178.38      
5 A' 1207 1126 131.41      
6 A' 912 851 42.36      
7 A' 758 707 36.60      
8 A' 605 565 15.49      
9 A' 546 509 9.28      
10 A' 379 353 0.06      
11 A' 252 235 4.11      
12 A" 1441 1344 13.46      
13 A" 1317 1229 365.13      
14 A" 1239 1155 104.85      
15 A" 611 570 1.08      
16 A" 435 405 1.16      
17 A" 218 203 2.45      
18 A" 86 80 0.85      

Unscaled Zero Point Vibrational Energy (zpe) 8692.1 cm-1
Scaled (by 0.9326) Zero Point Vibrational Energy (zpe) 8106.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3=FULL/cc-pVTZ
ABC
0.12499 0.08223 0.06808

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.119 -0.599 0.000
C2 -0.597 0.743 0.000
F3 1.427 -0.441 0.000
F4 -0.231 -1.286 1.074
F5 -0.231 -1.286 -1.074
F6 -0.231 1.426 -1.087
F7 -0.231 1.426 1.087
H8 -1.668 0.596 0.000

Atom - Atom Distances (Å)
  C1 C2 F3 F4 F5 F6 F7 H8
C11.52081.31681.32231.32232.32452.32452.1504
C21.52082.34442.32472.32471.33491.33491.0818
F31.31682.34442.14832.14832.72312.72313.2644
F41.32232.32472.14832.14793.46772.71192.6005
F51.32232.32472.14832.14792.71193.46772.6005
F62.32451.33492.72313.46772.71192.17441.9840
F72.32451.33492.72312.71193.46772.17441.9840
H82.15041.08183.26442.60052.60051.98401.9840

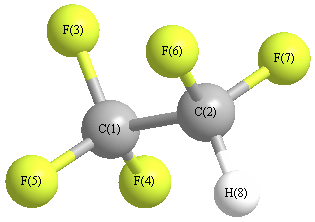
picture of pentafluoroethane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 F6 108.803 C1 C2 F7 108.803
C1 C2 H8 110.292 C2 C1 F3 111.216
C2 C1 F4 109.502 C2 C1 F5 109.502
F3 C1 F4 108.979 F3 C1 F5 108.979
F4 C1 F5 108.618 F6 C2 F7 109.064
F6 C2 H8 109.924 F7 C2 H8 109.924
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability