return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHNH2 (aminoethene)

using model chemistry: TPSSh/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at TPSSh/6-31G(2df,p)
 hartrees
Energy at 0K-133.963139
Energy at 298.15K-133.968585
HF Energy-133.963139
Nuclear repulsion energy71.414254
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3630 3503 6.67      
2 A 3528 3405 5.00      
3 A 3251 3137 14.71      
4 A 3164 3053 7.00      
5 A 3153 3043 14.45      
6 A 1720 1660 122.79      
7 A 1653 1596 20.03      
8 A 1459 1408 1.88      
9 A 1343 1296 8.33      
10 A 1292 1246 27.87      
11 A 1076 1038 9.17      
12 A 1015 979 20.06      
13 A 958 924 1.57      
14 A 810 782 53.71      
15 A 732 706 62.07      
16 A 631 609 217.35      
17 A 461 445 4.71      
18 A 360 347 44.90      

Unscaled Zero Point Vibrational Energy (zpe) 15117.8 cm-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 14588.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at TPSSh/6-31G(2df,p)
ABC
1.89584 0.33254 0.28565

See section I.F.4 to change rotational constant units
Geometric Data calculated at TPSSh/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.253 -0.196 0.018
C2 0.069 0.429 0.000
N3 -1.191 -0.168 -0.092
H4 1.338 -1.279 0.029
H5 2.175 0.371 -0.006
H6 0.022 1.515 -0.019
H7 -1.200 -1.156 0.129
H8 -1.925 0.322 0.404

Atom - Atom Distances (Å)
  C1 C2 N3 H4 H5 H6 H7 H8
C11.33872.44701.08641.08252.10832.63623.2425
C21.33871.39732.12742.10691.08792.03372.0369
N32.44701.39732.76513.41032.07581.01251.0115
H41.08642.12742.76511.85033.08882.54243.6531
H51.08252.10693.41031.85032.43853.70644.1204
H62.10831.08792.07583.08882.43852.94072.3221
H72.63622.03371.01252.54243.70642.94071.6683
H83.24252.03691.01153.65314.12042.32211.6683

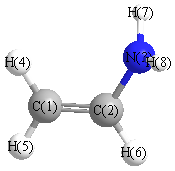
picture of aminoethene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 N3 126.846 C1 C2 H6 120.299
C2 C1 H4 122.282 C2 C1 H5 120.590
C2 N3 H7 114.166 C2 N3 H8 114.527
N3 C2 H6 112.700 H4 C1 H5 117.104
H7 N3 H8 111.027
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability