return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7NO (Propanamide)

using model chemistry: TPSSh/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at TPSSh/6-31G(2df,p)
 hartrees
Energy at 0K-248.557755
Energy at 298.15K-248.565784
HF Energy-248.557755
Nuclear repulsion energy178.481922
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3715 3585 26.00      
2 A 3578 3453 22.77      
3 A 3140 3030 18.54      
4 A 3119 3010 30.34      
5 A 3058 2951 19.84      
6 A 3049 2942 25.98      
7 A 3019 2913 22.18      
8 A 1786 1724 248.04      
9 A 1619 1562 87.64      
10 A 1515 1462 6.18      
11 A 1504 1451 5.72      
12 A 1476 1424 4.19      
13 A 1415 1366 4.56      
14 A 1394 1346 60.80      
15 A 1287 1242 20.20      
16 A 1279 1234 86.38      
17 A 1120 1081 2.08      
18 A 1095 1057 0.33      
19 A 1055 1018 2.60      
20 A 999 964 2.50      
21 A 809 781 7.21      
22 A 783 756 3.61      
23 A 644 621 4.22      
24 A 594 573 12.90      
25 A 498 480 3.98      
26 A 426 411 2.97      
27 A 249 241 10.15      
28 A 214 207 187.30      
29 A 196 190 3.33      
30 A 39 38 2.78      

Unscaled Zero Point Vibrational Energy (zpe) 22338.1 cm-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 21556.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at TPSSh/6-31G(2df,p)
ABC
0.32318 0.12656 0.09524

See section I.F.4 to change rotational constant units
Geometric Data calculated at TPSSh/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.971 0.035 -0.006
C2 -0.670 -0.764 0.009
C3 0.561 0.147 -0.002
O4 0.492 1.362 0.007
N5 1.750 -0.531 -0.023
H6 -2.833 -0.640 0.003
H7 -2.032 0.694 0.863
H8 -2.028 0.663 -0.899
H9 -0.615 -1.438 -0.855
H10 -0.620 -1.407 0.899
H11 1.804 -1.534 0.051
H12 2.618 -0.021 0.039

Atom - Atom Distances (Å)
  C1 C2 C3 O4 N5 H6 H7 H8 H9 H10 H11 H12
C11.52712.53432.79753.76411.09421.09271.09292.17422.17294.08854.5894
C21.52711.53092.42222.43142.16662.17022.16971.09771.09812.59113.3703
C32.53431.53091.21681.36913.48342.78722.78822.15022.14902.09162.0643
O42.79752.42221.21682.27293.88032.74702.76733.13163.11353.17972.5366
N53.76412.43141.36912.27294.58434.07294.05832.66702.68971.00781.0077
H61.09422.16663.48343.88034.58431.77751.77702.50802.50714.72245.4854
H71.09272.17022.78722.74704.07291.77751.76183.08282.53134.50984.7759
H81.09292.16972.78822.76734.05831.77701.76182.53233.08174.51844.7889
H92.17421.09772.15023.13162.66702.50803.08282.53231.75402.58523.6413
H102.17291.09812.14903.11352.68972.50712.53133.08171.75402.57123.6249
H114.08852.59112.09163.17971.00784.72244.50984.51842.58522.57121.7178
H124.58943.37032.06432.53661.00775.48544.77594.78893.64133.62491.7178

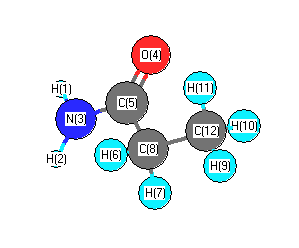
picture of Propanamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 N3 H2 33.982 H1 N3 C5 147.789
H2 N3 C5 113.827 N3 C5 O4 26.705
N3 C5 C8 17.968 O4 C5 C8 40.739
C5 C8 H6 95.523 C5 C8 H7 77.949
C5 C8 C12 9.030 H6 C8 H7 60.302
H6 C8 C12 103.589 H7 C8 C12 78.970
C8 C12 H9 31.363 C8 C12 H10 40.050
C8 C12 H11 70.588 H9 C12 H10 27.936
H9 C12 H11 40.401 H10 C12 H11 40.428
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability