Jump to
S1C2
S1C3
Energy calculated at TPSSh/6-311G**
| | hartrees |
|---|
| Energy at 0K | -1592.808703 |
| Energy at 298.15K | |
| HF Energy | -1592.808703 |
| Nuclear repulsion energy | 345.244301 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
491 |
491 |
0.00 |
53.00 |
0.08 |
0.14 |
| 2 |
A1 |
197 |
197 |
0.00 |
3.80 |
0.71 |
0.83 |
| 3 |
B1 |
411 |
411 |
0.00 |
37.67 |
0.75 |
0.86 |
| 4 |
B2 |
291 |
291 |
0.06 |
11.00 |
0.75 |
0.86 |
| 5 |
E |
398 |
398 |
0.28 |
7.66 |
0.75 |
0.86 |
| 5 |
E |
398 |
398 |
0.28 |
7.66 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1092.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1092.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-311G**
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.449 |
0.330 |
| S2 |
0.000 |
-1.449 |
0.330 |
| S3 |
-1.449 |
0.000 |
-0.330 |
| S4 |
1.449 |
0.000 |
-0.330 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8975 | 2.1525 | 2.1525 |
S2 | 2.8975 | | 2.1525 | 2.1525 | S3 | 2.1525 | 2.1525 | | 2.8975 | S4 | 2.1525 | 2.1525 | 2.8975 | |
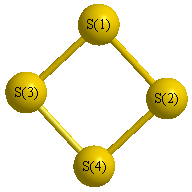 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.697 |
|
S1 |
S3 |
S4 |
47.697 |
| S2 |
S1 |
S3 |
47.697 |
|
S2 |
S4 |
S3 |
47.697 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
S |
0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.867 |
0.000 |
0.000 |
| y |
0.000 |
10.867 |
0.000 |
| z |
0.000 |
0.000 |
5.640 |
<r2> (average value of r
2) Å
2
| <r2> |
173.834 |
| (<r2>)1/2 |
13.185 |
Jump to
S1C1
S1C3
Energy calculated at TPSSh/6-311G**
| | hartrees |
|---|
| Energy at 0K | -1592.842650 |
| Energy at 298.15K | |
| HF Energy | -1592.842650 |
| Nuclear repulsion energy | 328.802262 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
682 |
682 |
1.38 |
36.71 |
0.25 |
0.40 |
| 2 |
A1 |
278 |
278 |
5.63 |
35.85 |
0.26 |
0.42 |
| 3 |
A1 |
61 |
61 |
8.52 |
2.25 |
0.63 |
0.77 |
| 4 |
A2 |
222 |
222 |
0.00 |
0.08 |
0.75 |
0.86 |
| 5 |
B2 |
647 |
647 |
139.27 |
1.79 |
0.75 |
0.86 |
| 6 |
B2 |
321 |
321 |
0.03 |
14.70 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1105.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1105.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-311G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.153 |
0.945 |
| S2 |
0.000 |
-1.153 |
0.945 |
| S3 |
0.000 |
1.488 |
-0.945 |
| S4 |
0.000 |
-1.488 |
-0.945 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.3069 | 1.9195 | 3.2482 |
S2 | 2.3069 | | 3.2482 | 1.9195 | S3 | 1.9195 | 3.2482 | | 2.9765 | S4 | 3.2482 | 1.9195 | 2.9765 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
100.046 |
|
S1 |
S3 |
S4 |
79.954 |
| S2 |
S1 |
S3 |
100.046 |
|
S2 |
S4 |
S3 |
79.954 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.083 |
|
|
|
| 2 |
S |
0.083 |
|
|
|
| 3 |
S |
-0.083 |
|
|
|
| 4 |
S |
-0.083 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.175 |
1.175 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.449 |
0.000 |
0.000 |
| y |
0.000 |
18.452 |
0.000 |
| z |
0.000 |
0.000 |
12.050 |
<r2> (average value of r
2) Å
2
| <r2> |
203.341 |
| (<r2>)1/2 |
14.260 |
Jump to
S1C1
S1C2
Energy calculated at TPSSh/6-311G**
| | hartrees |
|---|
| Energy at 0K | -1592.842376 |
| Energy at 298.15K | |
| HF Energy | -1592.842376 |
| Nuclear repulsion energy | 329.845446 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
705 |
705 |
0.00 |
40.50 |
0.26 |
0.42 |
| 2 |
Ag |
278 |
278 |
0.00 |
32.89 |
0.29 |
0.45 |
| 3 |
Au |
236 |
236 |
0.00 |
0.00 |
0.00 |
0.00 |
| 4 |
B1u |
659 |
659 |
142.74 |
0.00 |
0.00 |
0.00 |
| 5 |
B2u |
48i |
48i |
20.13 |
0.00 |
0.00 |
0.00 |
| 6 |
B3g |
320 |
320 |
0.00 |
15.04 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1074.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1074.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-311G**
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.956 |
1.300 |
| S2 |
0.000 |
0.956 |
-1.300 |
| S3 |
0.000 |
-0.956 |
1.300 |
| S4 |
0.000 |
-0.956 |
-1.300 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.6006 | 1.9118 | 3.2277 |
S2 | 2.6006 | | 3.2277 | 1.9118 | S3 | 1.9118 | 3.2277 | | 2.6006 | S4 | 3.2277 | 1.9118 | 2.6006 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
S |
0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.432 |
0.000 |
0.000 |
| y |
0.000 |
12.287 |
0.000 |
| z |
0.000 |
0.000 |
18.178 |
<r2> (average value of r
2) Å
2
| <r2> |
199.332 |
| (<r2>)1/2 |
14.119 |
