Jump to
S1C2
Vibrational Frequencies calculated at TPSSh/TZVP
Geometric Data calculated at TPSSh/TZVP
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at TPSSh/TZVP
| | hartrees |
|---|
| Energy at 0K | -208.037112 |
| Energy at 298.15K | -208.040452 |
| HF Energy | -208.037112 |
| Nuclear repulsion energy | 102.025722 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/TZVP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3779 |
3641 |
37.42 |
68.32 |
0.24 |
0.38 |
| 2 |
A |
3108 |
2995 |
3.58 |
84.75 |
0.37 |
0.54 |
| 3 |
A |
3018 |
2908 |
25.23 |
125.28 |
0.14 |
0.24 |
| 4 |
A |
2334 |
2249 |
0.12 |
71.96 |
0.20 |
0.34 |
| 5 |
A |
1492 |
1438 |
3.86 |
11.30 |
0.65 |
0.79 |
| 6 |
A |
1414 |
1363 |
39.27 |
6.87 |
0.74 |
0.85 |
| 7 |
A |
1366 |
1317 |
1.30 |
5.83 |
0.75 |
0.86 |
| 8 |
A |
1216 |
1171 |
17.85 |
3.61 |
0.55 |
0.71 |
| 9 |
A |
1050 |
1012 |
94.22 |
7.35 |
0.31 |
0.47 |
| 10 |
A |
985 |
949 |
30.71 |
0.41 |
0.69 |
0.81 |
| 11 |
A |
891 |
859 |
17.54 |
1.81 |
0.11 |
0.20 |
| 12 |
A |
578 |
557 |
2.63 |
2.48 |
0.22 |
0.36 |
| 13 |
A |
385 |
371 |
65.92 |
1.27 |
0.71 |
0.83 |
| 14 |
A |
304 |
293 |
91.87 |
4.07 |
0.74 |
0.85 |
| 15 |
A |
212 |
205 |
8.92 |
4.57 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11065.2 cm
-1
Scaled (by 0.9637) Zero Point Vibrational Energy (zpe) 10663.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/TZVP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.566 |
0.593 |
0.039 |
| C2 |
0.826 |
0.115 |
-0.007 |
| O3 |
-1.514 |
-0.451 |
-0.110 |
| H4 |
-0.703 |
1.152 |
0.972 |
| H5 |
-0.730 |
1.278 |
-0.794 |
| H6 |
-1.393 |
-1.082 |
0.613 |
| N7 |
1.911 |
-0.283 |
-0.015 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4726 | 1.4183 | 1.0969 | 1.0907 | 1.9543 | 2.6278 |
C2 | 1.4726 | | 2.4102 | 2.0907 | 2.0961 | 2.5967 | 1.1556 | O3 | 1.4183 | 2.4102 | | 2.0975 | 2.0189 | 0.9668 | 3.4305 | H4 | 1.0969 | 2.0907 | 2.0975 | | 1.7709 | 2.3654 | 3.1410 | H5 | 1.0907 | 2.0961 | 2.0189 | 1.7709 | | 2.8268 | 3.1651 | H6 | 1.9543 | 2.5967 | 0.9668 | 2.3654 | 2.8268 | | 3.4569 | N7 | 2.6278 | 1.1556 | 3.4305 | 3.1410 | 3.1651 | 3.4569 | |
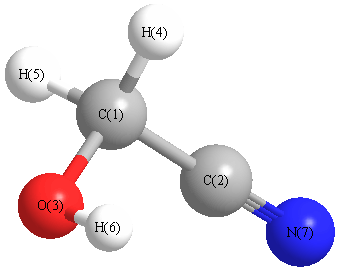 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
178.180 |
|
C1 |
O3 |
H6 |
108.565 |
| C2 |
C1 |
O3 |
112.956 |
|
C2 |
C1 |
H4 |
108.017 |
| C2 |
C1 |
H5 |
108.807 |
|
O3 |
C1 |
H4 |
112.384 |
| O3 |
C1 |
H5 |
106.425 |
|
H4 |
C1 |
H5 |
108.098 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/TZVP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.079 |
|
|
|
| 2 |
C |
-0.031 |
|
|
|
| 3 |
O |
-0.359 |
|
|
|
| 4 |
H |
0.139 |
|
|
|
| 5 |
H |
0.152 |
|
|
|
| 6 |
H |
0.278 |
|
|
|
| 7 |
N |
-0.100 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.346 |
1.201 |
1.360 |
2.966 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.047 |
-0.498 |
-0.034 |
| y |
-0.498 |
4.005 |
-0.085 |
| z |
-0.034 |
-0.085 |
3.441 |
<r2> (average value of r
2) Å
2
| <r2> |
79.060 |
| (<r2>)1/2 |
8.892 |
