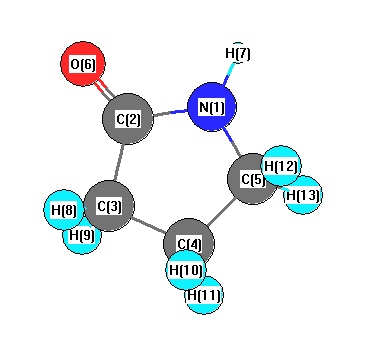
Vibrational Frequencies calculated at TPSSh/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3626 |
3491 |
27.80 |
|
|
|
| 2 |
A |
3139 |
3021 |
17.92 |
|
|
|
| 3 |
A |
3127 |
3009 |
23.40 |
|
|
|
| 4 |
A |
3095 |
2979 |
43.01 |
|
|
|
| 5 |
A |
3071 |
2956 |
21.64 |
|
|
|
| 6 |
A |
3062 |
2947 |
14.67 |
|
|
|
| 7 |
A |
3013 |
2900 |
69.98 |
|
|
|
| 8 |
A |
1782 |
1715 |
509.46 |
|
|
|
| 9 |
A |
1549 |
1491 |
3.48 |
|
|
|
| 10 |
A |
1514 |
1457 |
6.18 |
|
|
|
| 11 |
A |
1485 |
1429 |
3.57 |
|
|
|
| 12 |
A |
1448 |
1394 |
35.61 |
|
|
|
| 13 |
A |
1369 |
1317 |
7.35 |
|
|
|
| 14 |
A |
1341 |
1291 |
13.31 |
|
|
|
| 15 |
A |
1307 |
1258 |
25.25 |
|
|
|
| 16 |
A |
1269 |
1221 |
76.53 |
|
|
|
| 17 |
A |
1246 |
1199 |
21.29 |
|
|
|
| 18 |
A |
1212 |
1167 |
1.09 |
|
|
|
| 19 |
A |
1186 |
1142 |
4.05 |
|
|
|
| 20 |
A |
1093 |
1052 |
0.82 |
|
|
|
| 21 |
A |
1074 |
1034 |
16.11 |
|
|
|
| 22 |
A |
1003 |
965 |
12.10 |
|
|
|
| 23 |
A |
921 |
886 |
1.04 |
|
|
|
| 24 |
A |
905 |
871 |
1.21 |
|
|
|
| 25 |
A |
888 |
854 |
4.07 |
|
|
|
| 26 |
A |
811 |
781 |
3.62 |
|
|
|
| 27 |
A |
684 |
658 |
8.46 |
|
|
|
| 28 |
A |
626 |
603 |
18.85 |
|
|
|
| 29 |
A |
559 |
538 |
48.36 |
|
|
|
| 30 |
A |
488 |
470 |
61.30 |
|
|
|
| 31 |
A |
458 |
441 |
8.91 |
|
|
|
| 32 |
A |
191 |
184 |
4.41 |
|
|
|
| 33 |
A |
142 |
136 |
0.78 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24341.0 cm
-1
Scaled (by 0.9625) Zero Point Vibrational Energy (zpe) 23428.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31+G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.334 |
|
|
|
| 2 |
C |
0.155 |
|
|
|
| 3 |
C |
-0.096 |
|
|
|
| 4 |
C |
-0.302 |
|
|
|
| 5 |
C |
-0.245 |
|
|
|
| 6 |
O |
-0.517 |
|
|
|
| 7 |
H |
0.318 |
|
|
|
| 8 |
H |
0.185 |
|
|
|
| 9 |
H |
0.186 |
|
|
|
| 10 |
H |
0.162 |
|
|
|
| 11 |
H |
0.175 |
|
|
|
| 12 |
H |
0.155 |
|
|
|
| 13 |
H |
0.157 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.333 |
-0.607 |
0.386 |
4.392 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.815 |
0.443 |
-0.091 |
| y |
0.443 |
8.415 |
0.071 |
| z |
-0.091 |
0.071 |
6.414 |
<r2> (average value of r
2) Å
2
| <r2> |
147.118 |
| (<r2>)1/2 |
12.129 |