Jump to
S1C2
Energy calculated at TPSSh/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -323.773651 |
| Energy at 298.15K | |
| HF Energy | -323.773651 |
| Nuclear repulsion energy | 233.521369 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3099 |
3001 |
32.70 |
|
|
|
| 2 |
A' |
3036 |
2940 |
44.01 |
|
|
|
| 3 |
A' |
3025 |
2929 |
32.35 |
|
|
|
| 4 |
A' |
3022 |
2927 |
3.12 |
|
|
|
| 5 |
A' |
1734 |
1679 |
256.59 |
|
|
|
| 6 |
A' |
1526 |
1478 |
10.98 |
|
|
|
| 7 |
A' |
1514 |
1466 |
1.17 |
|
|
|
| 8 |
A' |
1502 |
1455 |
2.17 |
|
|
|
| 9 |
A' |
1423 |
1378 |
4.91 |
|
|
|
| 10 |
A' |
1404 |
1360 |
16.04 |
|
|
|
| 11 |
A' |
1331 |
1288 |
13.48 |
|
|
|
| 12 |
A' |
1154 |
1118 |
4.76 |
|
|
|
| 13 |
A' |
1054 |
1021 |
18.90 |
|
|
|
| 14 |
A' |
1032 |
999 |
8.27 |
|
|
|
| 15 |
A' |
920 |
891 |
65.82 |
|
|
|
| 16 |
A' |
803 |
777 |
424.11 |
|
|
|
| 17 |
A' |
632 |
612 |
91.31 |
|
|
|
| 18 |
A' |
368 |
356 |
1.05 |
|
|
|
| 19 |
A' |
340 |
329 |
3.26 |
|
|
|
| 20 |
A' |
147 |
142 |
0.38 |
|
|
|
| 21 |
A" |
3095 |
2997 |
78.11 |
|
|
|
| 22 |
A" |
3075 |
2978 |
1.96 |
|
|
|
| 23 |
A" |
3056 |
2959 |
2.50 |
|
|
|
| 24 |
A" |
1513 |
1465 |
6.35 |
|
|
|
| 25 |
A" |
1320 |
1279 |
0.03 |
|
|
|
| 26 |
A" |
1265 |
1225 |
0.25 |
|
|
|
| 27 |
A" |
1181 |
1143 |
0.81 |
|
|
|
| 28 |
A" |
887 |
858 |
0.84 |
|
|
|
| 29 |
A" |
751 |
727 |
2.10 |
|
|
|
| 30 |
A" |
231 |
224 |
0.01 |
|
|
|
| 31 |
A" |
211 |
204 |
0.78 |
|
|
|
| 32 |
A" |
94 |
91 |
1.29 |
|
|
|
| 33 |
A" |
64i |
61i |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22840.1 cm
-1
Scaled (by 0.9683) Zero Point Vibrational Energy (zpe) 22116.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.943 |
2.194 |
0.000 |
| C2 |
-1.513 |
0.726 |
0.000 |
| C3 |
0.000 |
0.588 |
0.000 |
| O4 |
0.313 |
-0.821 |
0.000 |
| N5 |
1.674 |
-1.038 |
0.000 |
| O6 |
1.953 |
-2.183 |
0.000 |
| H7 |
-3.031 |
2.276 |
0.000 |
| H8 |
-1.569 |
2.718 |
0.883 |
| H9 |
-1.569 |
2.718 |
-0.883 |
| H10 |
-1.914 |
0.215 |
-0.878 |
| H11 |
-1.914 |
0.215 |
0.878 |
| H12 |
0.439 |
1.051 |
0.887 |
| H13 |
0.439 |
1.051 |
-0.887 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5295 | 2.5210 | 3.7657 | 4.8508 | 5.8597 | 1.0910 | 1.0928 | 1.0928 | 2.1654 | 2.1654 | 2.7869 | 2.7869 |
C2 | 1.5295 | | 1.5189 | 2.3933 | 3.6426 | 4.5249 | 2.1695 | 2.1791 | 2.1791 | 1.0928 | 1.0928 | 2.1682 | 2.1682 | C3 | 2.5210 | 1.5189 | | 1.4430 | 2.3337 | 3.3898 | 3.4694 | 2.7890 | 2.7890 | 2.1389 | 2.1389 | 1.0930 | 1.0930 | O4 | 3.7657 | 2.3933 | 1.4430 | | 1.3781 | 2.1317 | 4.5579 | 4.1041 | 4.1041 | 2.6089 | 2.6089 | 2.0755 | 2.0755 | N5 | 4.8508 | 3.6426 | 2.3337 | 1.3781 | | 1.1783 | 5.7551 | 5.0402 | 5.0402 | 3.9010 | 3.9010 | 2.5842 | 2.5842 | O6 | 5.8597 | 4.5249 | 3.3898 | 2.1317 | 1.1783 | | 6.6875 | 6.0992 | 6.0992 | 4.6344 | 4.6344 | 3.6796 | 3.6796 | H7 | 1.0910 | 2.1695 | 3.4694 | 4.5579 | 5.7551 | 6.6875 | | 1.7641 | 1.7641 | 2.5033 | 2.5033 | 3.7851 | 3.7851 | H8 | 1.0928 | 2.1791 | 2.7890 | 4.1041 | 5.0402 | 6.0992 | 1.7641 | | 1.7666 | 3.0799 | 2.5264 | 2.6094 | 3.1533 | H9 | 1.0928 | 2.1791 | 2.7890 | 4.1041 | 5.0402 | 6.0992 | 1.7641 | 1.7666 | | 2.5264 | 3.0799 | 3.1533 | 2.6094 | H10 | 2.1654 | 1.0928 | 2.1389 | 2.6089 | 3.9010 | 4.6344 | 2.5033 | 3.0799 | 2.5264 | | 1.7564 | 3.0584 | 2.4975 | H11 | 2.1654 | 1.0928 | 2.1389 | 2.6089 | 3.9010 | 4.6344 | 2.5033 | 2.5264 | 3.0799 | 1.7564 | | 2.4975 | 3.0584 | H12 | 2.7869 | 2.1682 | 1.0930 | 2.0755 | 2.5842 | 3.6796 | 3.7851 | 2.6094 | 3.1533 | 3.0584 | 2.4975 | | 1.7743 | H13 | 2.7869 | 2.1682 | 1.0930 | 2.0755 | 2.5842 | 3.6796 | 3.7851 | 3.1533 | 2.6094 | 2.4975 | 3.0584 | 1.7743 | |
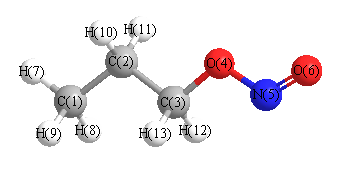 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.581 |
|
C1 |
C2 |
H10 |
110.224 |
| C1 |
C2 |
H11 |
110.224 |
|
C2 |
C1 |
H7 |
110.658 |
| C2 |
C1 |
H8 |
111.313 |
|
C2 |
C1 |
H9 |
111.313 |
| C2 |
C3 |
O4 |
107.779 |
|
C2 |
C3 |
H12 |
111.177 |
| C2 |
C3 |
H13 |
111.177 |
|
C3 |
C2 |
H10 |
108.865 |
| C3 |
C2 |
H11 |
108.865 |
|
C3 |
O4 |
N5 |
111.612 |
| O4 |
C3 |
H12 |
109.072 |
|
O4 |
C3 |
H13 |
109.072 |
| O4 |
N5 |
O6 |
112.766 |
|
H7 |
C1 |
H8 |
107.768 |
| H7 |
C1 |
H9 |
107.768 |
|
H8 |
C1 |
H9 |
107.861 |
| H10 |
C2 |
H11 |
106.954 |
|
H12 |
C3 |
H13 |
108.523 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.236 |
|
|
|
| 2 |
C |
-0.116 |
|
|
|
| 3 |
C |
0.072 |
|
|
|
| 4 |
O |
-0.202 |
|
|
|
| 5 |
N |
0.112 |
|
|
|
| 6 |
O |
-0.119 |
|
|
|
| 7 |
H |
0.083 |
|
|
|
| 8 |
H |
0.072 |
|
|
|
| 9 |
H |
0.072 |
|
|
|
| 10 |
H |
0.075 |
|
|
|
| 11 |
H |
0.075 |
|
|
|
| 12 |
H |
0.055 |
|
|
|
| 13 |
H |
0.055 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.285 |
2.320 |
0.000 |
2.652 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.085 |
-2.407 |
0.000 |
| y |
-2.407 |
9.394 |
0.000 |
| z |
0.000 |
0.000 |
6.173 |
<r2> (average value of r
2) Å
2
| <r2> |
245.623 |
| (<r2>)1/2 |
15.672 |
Jump to
S1C1
Energy calculated at TPSSh/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -323.775201 |
| Energy at 298.15K | -323.784116 |
| HF Energy | -323.775201 |
| Nuclear repulsion energy | 240.165176 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3103 |
3004 |
32.33 |
|
|
|
| 2 |
A |
3093 |
2995 |
45.18 |
|
|
|
| 3 |
A |
3078 |
2980 |
25.62 |
|
|
|
| 4 |
A |
3057 |
2960 |
15.28 |
|
|
|
| 5 |
A |
3027 |
2932 |
10.70 |
|
|
|
| 6 |
A |
3027 |
2931 |
50.81 |
|
|
|
| 7 |
A |
3021 |
2925 |
11.43 |
|
|
|
| 8 |
A |
1740 |
1685 |
272.68 |
|
|
|
| 9 |
A |
1517 |
1469 |
5.58 |
|
|
|
| 10 |
A |
1511 |
1463 |
6.69 |
|
|
|
| 11 |
A |
1496 |
1449 |
2.54 |
|
|
|
| 12 |
A |
1487 |
1440 |
2.03 |
|
|
|
| 13 |
A |
1421 |
1376 |
4.47 |
|
|
|
| 14 |
A |
1390 |
1346 |
13.45 |
|
|
|
| 15 |
A |
1380 |
1336 |
2.63 |
|
|
|
| 16 |
A |
1303 |
1262 |
1.16 |
|
|
|
| 17 |
A |
1282 |
1241 |
9.25 |
|
|
|
| 18 |
A |
1184 |
1146 |
7.70 |
|
|
|
| 19 |
A |
1113 |
1078 |
4.64 |
|
|
|
| 20 |
A |
1068 |
1034 |
18.17 |
|
|
|
| 21 |
A |
992 |
961 |
9.16 |
|
|
|
| 22 |
A |
923 |
893 |
45.56 |
|
|
|
| 23 |
A |
869 |
841 |
5.33 |
|
|
|
| 24 |
A |
805 |
779 |
247.53 |
|
|
|
| 25 |
A |
759 |
735 |
131.66 |
|
|
|
| 26 |
A |
580 |
562 |
68.96 |
|
|
|
| 27 |
A |
436 |
422 |
15.12 |
|
|
|
| 28 |
A |
364 |
352 |
2.21 |
|
|
|
| 29 |
A |
273 |
264 |
1.23 |
|
|
|
| 30 |
A |
235 |
228 |
0.70 |
|
|
|
| 31 |
A |
182 |
176 |
0.48 |
|
|
|
| 32 |
A |
108 |
104 |
0.91 |
|
|
|
| 33 |
A |
55 |
53 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22937.7 cm
-1
Scaled (by 0.9683) Zero Point Vibrational Energy (zpe) 22210.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.236 |
-0.927 |
0.163 |
| C2 |
-1.800 |
0.465 |
-0.296 |
| C3 |
-0.441 |
0.880 |
0.244 |
| O4 |
0.554 |
-0.025 |
-0.297 |
| N5 |
1.810 |
0.160 |
0.260 |
| O6 |
2.714 |
-0.504 |
-0.108 |
| H7 |
-3.222 |
-1.172 |
-0.236 |
| H8 |
-2.294 |
-0.980 |
1.253 |
| H9 |
-1.534 |
-1.689 |
-0.176 |
| H10 |
-1.772 |
0.511 |
-1.388 |
| H11 |
-2.527 |
1.213 |
0.033 |
| H12 |
-0.192 |
1.898 |
-0.061 |
| H13 |
-0.413 |
0.821 |
1.334 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5291 | 2.5483 | 2.9684 | 4.1909 | 4.9753 | 1.0911 | 1.0928 | 1.0904 | 2.1651 | 2.1633 | 3.4942 | 2.7840 |
C2 | 1.5291 | | 1.5199 | 2.4044 | 3.6651 | 4.6200 | 2.1691 | 2.1748 | 2.1737 | 1.0928 | 1.0938 | 2.1665 | 2.1694 | C3 | 2.5483 | 1.5199 | | 1.4495 | 2.3631 | 3.4625 | 3.4890 | 2.8121 | 2.8232 | 2.1373 | 2.1229 | 1.0919 | 1.0922 | O4 | 2.9684 | 2.4044 | 1.4495 | | 1.3864 | 2.2199 | 3.9467 | 3.3798 | 2.6730 | 2.6243 | 3.3367 | 2.0763 | 2.0766 | N5 | 4.1909 | 3.6651 | 2.3631 | 1.3864 | | 1.1804 | 5.2288 | 4.3731 | 3.8462 | 3.9583 | 4.4685 | 2.6708 | 2.5559 | O6 | 4.9753 | 4.6200 | 3.4625 | 2.2199 | 1.1804 | | 5.9741 | 5.2107 | 4.4104 | 4.7734 | 5.5163 | 3.7704 | 3.6897 | H7 | 1.0911 | 2.1691 | 3.4890 | 3.9467 | 5.2288 | 5.9741 | | 1.7651 | 1.7661 | 2.5023 | 2.4987 | 4.3168 | 3.7849 | H8 | 1.0928 | 2.1748 | 2.8121 | 3.3798 | 4.3731 | 5.2107 | 1.7651 | | 1.7669 | 3.0768 | 2.5197 | 3.7980 | 2.6048 | H9 | 1.0904 | 2.1737 | 2.8232 | 2.6730 | 3.8462 | 4.4104 | 1.7661 | 1.7669 | | 2.5231 | 3.0743 | 3.8318 | 3.1366 | H10 | 2.1651 | 1.0928 | 2.1373 | 2.6243 | 3.9583 | 4.7734 | 2.5023 | 3.0768 | 2.5231 | | 1.7553 | 2.4854 | 3.0575 | H11 | 2.1633 | 1.0938 | 2.1229 | 3.3367 | 4.4685 | 5.5163 | 2.4987 | 2.5197 | 3.0743 | 1.7553 | | 2.4350 | 2.5124 | H12 | 3.4942 | 2.1665 | 1.0919 | 2.0763 | 2.6708 | 3.7704 | 4.3168 | 3.7980 | 3.8318 | 2.4854 | 2.4350 | | 1.7766 | H13 | 2.7840 | 2.1694 | 1.0922 | 2.0766 | 2.5559 | 3.6897 | 3.7849 | 2.6048 | 3.1366 | 3.0575 | 2.5124 | 1.7766 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.391 |
|
C1 |
C2 |
H10 |
110.221 |
| C1 |
C2 |
H11 |
110.019 |
|
C2 |
C1 |
H7 |
110.644 |
| C2 |
C1 |
H8 |
110.994 |
|
C2 |
C1 |
H9 |
111.051 |
| C2 |
C3 |
O4 |
108.116 |
|
C2 |
C3 |
H12 |
111.032 |
| C2 |
C3 |
H13 |
111.252 |
|
C3 |
C2 |
H10 |
108.678 |
| C3 |
C2 |
H11 |
107.501 |
|
C3 |
O4 |
N5 |
112.862 |
| O4 |
C3 |
H12 |
108.759 |
|
O4 |
C3 |
H13 |
108.761 |
| O4 |
N5 |
O6 |
119.516 |
|
H7 |
C1 |
H8 |
107.851 |
| H7 |
C1 |
H9 |
108.112 |
|
H8 |
C1 |
H9 |
108.064 |
| H10 |
C2 |
H11 |
106.783 |
|
H12 |
C3 |
H13 |
108.861 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.228 |
|
|
|
| 2 |
C |
-0.099 |
|
|
|
| 3 |
C |
0.050 |
|
|
|
| 4 |
O |
-0.201 |
|
|
|
| 5 |
N |
0.112 |
|
|
|
| 6 |
O |
-0.120 |
|
|
|
| 7 |
H |
0.079 |
|
|
|
| 8 |
H |
0.069 |
|
|
|
| 9 |
H |
0.080 |
|
|
|
| 10 |
H |
0.072 |
|
|
|
| 11 |
H |
0.064 |
|
|
|
| 12 |
H |
0.076 |
|
|
|
| 13 |
H |
0.046 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.210 |
-0.790 |
0.717 |
2.454 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.181 |
-1.078 |
-0.050 |
| y |
-1.078 |
7.567 |
-0.016 |
| z |
-0.050 |
-0.016 |
6.590 |
<r2> (average value of r
2) Å
2
| <r2> |
198.110 |
| (<r2>)1/2 |
14.075 |
