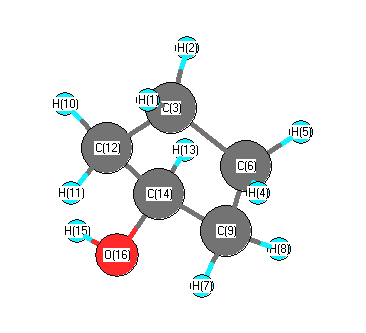
Vibrational Frequencies calculated at B3LYP/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3841 |
3714 |
27.10 |
|
|
|
| 2 |
A |
3095 |
2993 |
70.30 |
|
|
|
| 3 |
A |
3081 |
2979 |
16.34 |
|
|
|
| 4 |
A |
3067 |
2966 |
12.78 |
|
|
|
| 5 |
A |
3064 |
2963 |
40.96 |
|
|
|
| 6 |
A |
3050 |
2949 |
39.82 |
|
|
|
| 7 |
A |
3039 |
2939 |
26.43 |
|
|
|
| 8 |
A |
3025 |
2925 |
24.25 |
|
|
|
| 9 |
A |
3021 |
2921 |
28.54 |
|
|
|
| 10 |
A |
2960 |
2862 |
40.17 |
|
|
|
| 11 |
A |
1523 |
1473 |
3.09 |
|
|
|
| 12 |
A |
1501 |
1451 |
5.55 |
|
|
|
| 13 |
A |
1498 |
1449 |
2.43 |
|
|
|
| 14 |
A |
1491 |
1442 |
1.61 |
|
|
|
| 15 |
A |
1419 |
1372 |
4.54 |
|
|
|
| 16 |
A |
1374 |
1328 |
21.24 |
|
|
|
| 17 |
A |
1338 |
1294 |
0.09 |
|
|
|
| 18 |
A |
1331 |
1287 |
3.48 |
|
|
|
| 19 |
A |
1317 |
1273 |
4.67 |
|
|
|
| 20 |
A |
1300 |
1257 |
5.88 |
|
|
|
| 21 |
A |
1277 |
1235 |
1.21 |
|
|
|
| 22 |
A |
1261 |
1219 |
19.00 |
|
|
|
| 23 |
A |
1208 |
1168 |
5.15 |
|
|
|
| 24 |
A |
1197 |
1158 |
11.22 |
|
|
|
| 25 |
A |
1187 |
1148 |
8.84 |
|
|
|
| 26 |
A |
1089 |
1053 |
64.87 |
|
|
|
| 27 |
A |
1070 |
1035 |
36.75 |
|
|
|
| 28 |
A |
1035 |
1001 |
8.41 |
|
|
|
| 29 |
A |
992 |
960 |
0.36 |
|
|
|
| 30 |
A |
967 |
935 |
8.55 |
|
|
|
| 31 |
A |
947 |
916 |
6.07 |
|
|
|
| 32 |
A |
891 |
861 |
0.07 |
|
|
|
| 33 |
A |
875 |
846 |
0.11 |
|
|
|
| 34 |
A |
815 |
788 |
1.53 |
|
|
|
| 35 |
A |
774 |
748 |
1.01 |
|
|
|
| 36 |
A |
619 |
599 |
0.42 |
|
|
|
| 37 |
A |
538 |
521 |
5.67 |
|
|
|
| 38 |
A |
469 |
454 |
6.56 |
|
|
|
| 39 |
A |
360 |
348 |
10.46 |
|
|
|
| 40 |
A |
269 |
260 |
108.27 |
|
|
|
| 41 |
A |
184 |
178 |
1.10 |
|
|
|
| 42 |
A |
35 |
34 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31695.1 cm
-1
Scaled (by 0.967) Zero Point Vibrational Energy (zpe) 30649.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.136 |
|
|
|
| 2 |
H |
0.160 |
|
|
|
| 3 |
C |
-0.311 |
|
|
|
| 4 |
H |
0.149 |
|
|
|
| 5 |
H |
0.148 |
|
|
|
| 6 |
C |
-0.335 |
|
|
|
| 7 |
H |
0.157 |
|
|
|
| 8 |
H |
0.136 |
|
|
|
| 9 |
C |
-0.274 |
|
|
|
| 10 |
H |
0.126 |
|
|
|
| 11 |
H |
0.162 |
|
|
|
| 12 |
C |
-0.288 |
|
|
|
| 13 |
H |
0.143 |
|
|
|
| 14 |
C |
0.326 |
|
|
|
| 15 |
H |
0.224 |
|
|
|
| 16 |
O |
-0.659 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.692 |
-1.153 |
0.854 |
1.593 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.343 |
0.115 |
0.042 |
| y |
0.115 |
9.668 |
-0.026 |
| z |
0.042 |
-0.026 |
8.362 |
<r2> (average value of r
2) Å
2
| <r2> |
166.894 |
| (<r2>)1/2 |
12.919 |