Jump to
S1C2
S1C3
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -34.356134 |
| Energy at 298.15K | |
| HF Energy | -34.356134 |
| Nuclear repulsion energy | 24.917954 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3925 |
3790 |
47.10 |
|
|
|
| 2 |
A1 |
717 |
692 |
0.90 |
|
|
|
| 3 |
A1 |
634 |
612 |
142.84 |
|
|
|
| 4 |
A1 |
282 |
273 |
15.09 |
|
|
|
| 5 |
A2 |
238i |
230i |
0.00 |
|
|
|
| 6 |
B1 |
339 |
328 |
73.20 |
|
|
|
| 7 |
B2 |
3923 |
3789 |
104.23 |
|
|
|
| 8 |
B2 |
1504 |
1452 |
339.44 |
|
|
|
| 9 |
B2 |
510 |
492 |
310.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5797.7 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 5598.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
0.011 |
| O2 |
0.000 |
1.438 |
0.083 |
| O3 |
0.000 |
-1.438 |
0.083 |
| H4 |
0.000 |
2.029 |
-0.682 |
| H5 |
0.000 |
-2.029 |
-0.682 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4399 | 1.4399 | 2.1444 | 2.1444 |
O2 | 1.4399 | | 2.8761 | 0.9668 | 3.5508 | O3 | 1.4399 | 2.8761 | | 3.5508 | 0.9668 | H4 | 2.1444 | 0.9668 | 3.5508 | | 4.0588 | H5 | 2.1444 | 3.5508 | 0.9668 | 4.0588 | |
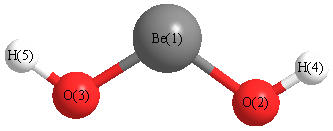 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
124.842 |
|
Be1 |
O3 |
H5 |
124.842 |
| O2 |
Be1 |
O3 |
174.269 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
0.020 |
|
|
|
| 2 |
O |
-0.378 |
|
|
|
| 3 |
O |
-0.378 |
|
|
|
| 4 |
H |
0.368 |
|
|
|
| 5 |
H |
0.368 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.805 |
2.805 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.945 |
0.000 |
0.000 |
| y |
0.000 |
-15.490 |
0.000 |
| z |
0.000 |
0.000 |
-14.111 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.145 |
0.000 |
0.000 |
| y |
0.000 |
0.038 |
0.000 |
| z |
0.000 |
0.000 |
2.107 |
|
| Polar |
|---|
| 3z2-r2 | 4.214 |
|---|
| x2-y2 | -1.456 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.204 |
0.000 |
0.000 |
| y |
0.000 |
3.651 |
0.000 |
| z |
0.000 |
0.000 |
2.487 |
<r2> (average value of r
2) Å
2
| <r2> |
43.757 |
| (<r2>)1/2 |
6.615 |
Jump to
S1C1
S1C3
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -34.357240 |
| Energy at 298.15K | -34.358738 |
| HF Energy | -34.357240 |
| Nuclear repulsion energy | 24.925757 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3906 |
3772 |
22.10 |
|
|
|
| 2 |
A |
714 |
690 |
0.02 |
|
|
|
| 3 |
A |
626 |
605 |
68.39 |
|
|
|
| 4 |
A |
304 |
294 |
59.42 |
|
|
|
| 5 |
A |
218 |
210 |
134.54 |
|
|
|
| 6 |
B |
3905 |
3771 |
115.51 |
|
|
|
| 7 |
B |
1505 |
1454 |
328.91 |
|
|
|
| 8 |
B |
626 |
605 |
335.42 |
|
|
|
| 9 |
B |
295 |
285 |
89.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6050.0 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 5842.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
-0.013 |
| O2 |
0.000 |
1.440 |
-0.057 |
| O3 |
0.000 |
-1.440 |
-0.057 |
| H4 |
0.605 |
1.974 |
0.478 |
| H5 |
-0.605 |
-1.974 |
0.478 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4407 | 1.4407 | 2.1221 | 2.1221 |
O2 | 1.4407 | | 2.8800 | 0.9681 | 3.5081 | O3 | 1.4407 | 2.8800 | | 3.5081 | 0.9681 | H4 | 2.1221 | 0.9681 | 3.5081 | | 4.1290 | H5 | 2.1221 | 3.5081 | 0.9681 | 4.1290 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
122.301 |
|
Be1 |
O3 |
H5 |
122.301 |
| O2 |
Be1 |
O3 |
176.498 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
0.015 |
|
|
|
| 2 |
O |
-0.376 |
|
|
|
| 3 |
O |
-0.376 |
|
|
|
| 4 |
H |
0.369 |
|
|
|
| 5 |
H |
0.369 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.935 |
1.935 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.887 |
4.931 |
0.000 |
| y |
4.931 |
-16.452 |
0.000 |
| z |
0.000 |
0.000 |
-15.569 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.124 |
4.931 |
0.000 |
| y |
4.931 |
-1.225 |
0.000 |
| z |
0.000 |
0.000 |
0.100 |
|
| Polar |
|---|
| 3z2-r2 | 0.201 |
|---|
| x2-y2 | 1.566 |
|---|
| xy | 4.931 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.388 |
0.160 |
0.000 |
| y |
0.160 |
3.621 |
0.000 |
| z |
0.000 |
0.000 |
2.360 |
<r2> (average value of r
2) Å
2
| <r2> |
43.670 |
| (<r2>)1/2 |
6.608 |
Jump to
S1C1
S1C2
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -34.347475 |
| Energy at 298.15K | |
| HF Energy | -34.347475 |
| Nuclear repulsion energy | 25.075214 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
4106 |
3965 |
0.00 |
|
|
|
| 2 |
Σg |
762 |
736 |
0.00 |
|
|
|
| 3 |
Σu |
4102 |
3961 |
438.16 |
|
|
|
| 4 |
Σu |
1592 |
1538 |
528.23 |
|
|
|
| 5 |
Πg |
432i |
418i |
0.00 |
|
|
|
| 5 |
Πg |
432i |
418i |
0.00 |
|
|
|
| 6 |
Πu |
332 |
320 |
12.47 |
|
|
|
| 6 |
Πu |
332 |
320 |
12.47 |
|
|
|
| 7 |
Πu |
453i |
437i |
423.47 |
|
|
|
| 7 |
Πu |
453i |
437i |
423.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4727.3 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 4565.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
0.000 |
| O2 |
0.000 |
0.000 |
1.411 |
| O3 |
0.000 |
0.000 |
-1.411 |
| H4 |
0.000 |
0.000 |
2.368 |
| H5 |
0.000 |
0.000 |
-2.368 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4113 | 1.4113 | 2.3676 | 2.3676 |
O2 | 1.4113 | | 2.8225 | 0.9563 | 3.7788 | O3 | 1.4113 | 2.8225 | | 3.7788 | 0.9563 | H4 | 2.3676 | 0.9563 | 3.7788 | | 4.7351 | H5 | 2.3676 | 3.7788 | 0.9563 | 4.7351 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
180.000 |
|
Be1 |
O3 |
H5 |
180.000 |
| O2 |
Be1 |
O3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
0.077 |
|
|
|
| 2 |
O |
-0.435 |
|
|
|
| 3 |
O |
-0.435 |
|
|
|
| 4 |
H |
0.397 |
|
|
|
| 5 |
H |
0.397 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.802 |
0.000 |
0.000 |
| y |
0.000 |
-16.802 |
0.000 |
| z |
0.000 |
0.000 |
-7.433 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.684 |
0.000 |
0.000 |
| y |
0.000 |
-4.684 |
0.000 |
| z |
0.000 |
0.000 |
9.369 |
|
| Polar |
|---|
| 3z2-r2 | 18.737 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.044 |
0.000 |
0.000 |
| y |
0.000 |
2.044 |
0.000 |
| z |
0.000 |
0.000 |
3.969 |
<r2> (average value of r
2) Å
2
| <r2> |
43.655 |
| (<r2>)1/2 |
6.607 |
